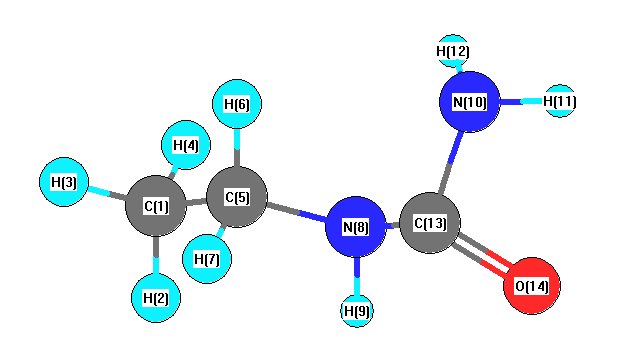
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3718 |
3579 |
23.41 |
|
|
|
| 2 |
A |
3646 |
3510 |
22.44 |
|
|
|
| 3 |
A |
3601 |
3467 |
14.49 |
|
|
|
| 4 |
A |
3128 |
3012 |
26.96 |
|
|
|
| 5 |
A |
3115 |
2999 |
37.06 |
|
|
|
| 6 |
A |
3044 |
2930 |
22.94 |
|
|
|
| 7 |
A |
3030 |
2917 |
32.68 |
|
|
|
| 8 |
A |
3000 |
2888 |
49.14 |
|
|
|
| 9 |
A |
1818 |
1751 |
509.54 |
|
|
|
| 10 |
A |
1667 |
1604 |
161.63 |
|
|
|
| 11 |
A |
1544 |
1486 |
2.51 |
|
|
|
| 12 |
A |
1518 |
1461 |
5.43 |
|
|
|
| 13 |
A |
1508 |
1451 |
9.35 |
|
|
|
| 14 |
A |
1479 |
1424 |
21.45 |
|
|
|
| 15 |
A |
1447 |
1393 |
263.34 |
|
|
|
| 16 |
A |
1422 |
1369 |
32.26 |
|
|
|
| 17 |
A |
1369 |
1318 |
19.90 |
|
|
|
| 18 |
A |
1314 |
1265 |
1.94 |
|
|
|
| 19 |
A |
1213 |
1168 |
19.83 |
|
|
|
| 20 |
A |
1173 |
1129 |
1.07 |
|
|
|
| 21 |
A |
1139 |
1096 |
27.99 |
|
|
|
| 22 |
A |
1076 |
1036 |
16.01 |
|
|
|
| 23 |
A |
1001 |
964 |
1.70 |
|
|
|
| 24 |
A |
905 |
871 |
2.53 |
|
|
|
| 25 |
A |
827 |
797 |
0.63 |
|
|
|
| 26 |
A |
786 |
757 |
57.10 |
|
|
|
| 27 |
A |
595 |
573 |
58.45 |
|
|
|
| 28 |
A |
560 |
539 |
17.01 |
|
|
|
| 29 |
A |
539 |
519 |
133.82 |
|
|
|
| 30 |
A |
488 |
470 |
26.89 |
|
|
|
| 31 |
A |
383 |
369 |
135.02 |
|
|
|
| 32 |
A |
361 |
348 |
0.71 |
|
|
|
| 33 |
A |
275 |
265 |
3.53 |
|
|
|
| 34 |
A |
195 |
188 |
3.29 |
|
|
|
| 35 |
A |
106 |
102 |
6.93 |
|
|
|
| 36 |
A |
44 |
42 |
0.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26516.1 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 25527.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.697 |
|
|
|
| 2 |
H |
0.232 |
|
|
|
| 3 |
H |
0.235 |
|
|
|
| 4 |
H |
0.235 |
|
|
|
| 5 |
C |
-0.283 |
|
|
|
| 6 |
H |
0.213 |
|
|
|
| 7 |
H |
0.232 |
|
|
|
| 8 |
N |
-0.604 |
|
|
|
| 9 |
H |
0.354 |
|
|
|
| 10 |
N |
-0.804 |
|
|
|
| 11 |
H |
0.358 |
|
|
|
| 12 |
H |
0.344 |
|
|
|
| 13 |
C |
0.598 |
|
|
|
| 14 |
O |
-0.412 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.111 |
2.691 |
-0.649 |
4.165 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.208 |
6.994 |
-1.555 |
| y |
6.994 |
-34.751 |
-1.732 |
| z |
-1.555 |
-1.732 |
-38.051 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.807 |
6.994 |
-1.555 |
| y |
6.994 |
3.378 |
-1.732 |
| z |
-1.555 |
-1.732 |
-1.571 |
|
| Polar |
|---|
| 3z2-r2 | -3.143 |
|---|
| x2-y2 | -3.457 |
|---|
| xy | 6.994 |
|---|
| xz | -1.555 |
|---|
| yz | -1.732 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
201.951 |
| (<r2>)1/2 |
14.211 |