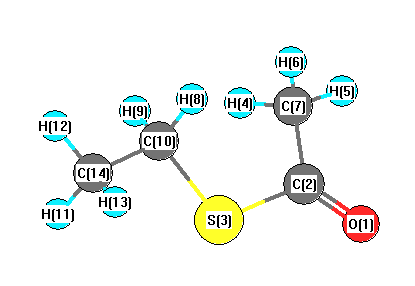
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3159 |
3042 |
8.68 |
|
|
|
| 2 |
A |
3135 |
3018 |
32.82 |
|
|
|
| 3 |
A |
3126 |
3009 |
5.81 |
|
|
|
| 4 |
A |
3119 |
3003 |
24.48 |
|
|
|
| 5 |
A |
3110 |
2994 |
0.18 |
|
|
|
| 6 |
A |
3064 |
2950 |
22.53 |
|
|
|
| 7 |
A |
3054 |
2940 |
0.98 |
|
|
|
| 8 |
A |
3048 |
2934 |
26.07 |
|
|
|
| 9 |
A |
1801 |
1733 |
330.22 |
|
|
|
| 10 |
A |
1516 |
1460 |
4.69 |
|
|
|
| 11 |
A |
1508 |
1452 |
12.97 |
|
|
|
| 12 |
A |
1494 |
1438 |
0.64 |
|
|
|
| 13 |
A |
1486 |
1430 |
16.58 |
|
|
|
| 14 |
A |
1472 |
1418 |
23.66 |
|
|
|
| 15 |
A |
1424 |
1371 |
7.51 |
|
|
|
| 16 |
A |
1389 |
1338 |
26.49 |
|
|
|
| 17 |
A |
1317 |
1268 |
27.42 |
|
|
|
| 18 |
A |
1287 |
1239 |
0.29 |
|
|
|
| 19 |
A |
1152 |
1109 |
163.01 |
|
|
|
| 20 |
A |
1085 |
1045 |
7.53 |
|
|
|
| 21 |
A |
1061 |
1021 |
0.75 |
|
|
|
| 22 |
A |
1027 |
989 |
2.28 |
|
|
|
| 23 |
A |
1004 |
967 |
4.22 |
|
|
|
| 24 |
A |
950 |
915 |
49.79 |
|
|
|
| 25 |
A |
802 |
772 |
7.93 |
|
|
|
| 26 |
A |
699 |
673 |
0.12 |
|
|
|
| 27 |
A |
640 |
617 |
69.32 |
|
|
|
| 28 |
A |
514 |
495 |
1.63 |
|
|
|
| 29 |
A |
448 |
432 |
3.20 |
|
|
|
| 30 |
A |
374 |
360 |
1.80 |
|
|
|
| 31 |
A |
302 |
291 |
1.25 |
|
|
|
| 32 |
A |
263 |
253 |
0.05 |
|
|
|
| 33 |
A |
157 |
152 |
1.10 |
|
|
|
| 34 |
A |
140 |
134 |
0.08 |
|
|
|
| 35 |
A |
86 |
83 |
0.43 |
|
|
|
| 36 |
A |
53 |
51 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25133.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 24195.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.295 |
|
|
|
| 2 |
C |
0.172 |
|
|
|
| 3 |
S |
0.149 |
|
|
|
| 4 |
H |
0.252 |
|
|
|
| 5 |
H |
0.262 |
|
|
|
| 6 |
H |
0.252 |
|
|
|
| 7 |
C |
-0.739 |
|
|
|
| 8 |
H |
0.256 |
|
|
|
| 9 |
H |
0.256 |
|
|
|
| 10 |
C |
-0.622 |
|
|
|
| 11 |
H |
0.243 |
|
|
|
| 12 |
H |
0.234 |
|
|
|
| 13 |
H |
0.243 |
|
|
|
| 14 |
C |
-0.662 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.568 |
3.507 |
0.000 |
4.346 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.650 |
3.799 |
0.001 |
| y |
3.799 |
-45.187 |
0.000 |
| z |
0.001 |
0.000 |
-44.333 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.891 |
3.799 |
0.001 |
| y |
3.799 |
0.305 |
0.000 |
| z |
0.001 |
0.000 |
1.586 |
|
| Polar |
|---|
| 3z2-r2 | 3.171 |
|---|
| x2-y2 | -1.464 |
|---|
| xy | 3.799 |
|---|
| xz | 0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
251.813 |
| (<r2>)1/2 |
15.869 |