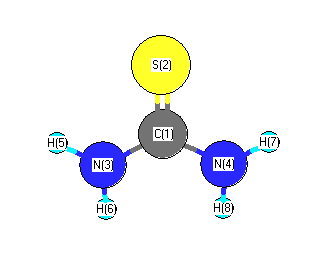
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3973 |
3593 |
41.86 |
|
|
|
| 2 |
A |
3844 |
3477 |
28.07 |
|
|
|
| 3 |
A |
1846 |
1670 |
104.46 |
|
|
|
| 4 |
A |
1533 |
1387 |
394.91 |
|
|
|
| 5 |
A |
1157 |
1046 |
96.35 |
|
|
|
| 6 |
A |
793 |
717 |
35.96 |
|
|
|
| 7 |
A |
520 |
470 |
0.17 |
|
|
|
| 8 |
A |
497 |
450 |
5.55 |
|
|
|
| 9 |
A |
232 |
210 |
104.60 |
|
|
|
| 10 |
B |
3972 |
3592 |
69.41 |
|
|
|
| 11 |
B |
3837 |
3470 |
92.33 |
|
|
|
| 12 |
B |
1819 |
1645 |
248.18 |
|
|
|
| 13 |
B |
1563 |
1414 |
179.04 |
|
|
|
| 14 |
B |
1165 |
1054 |
13.01 |
|
|
|
| 15 |
B |
704 |
636 |
2.97 |
|
|
|
| 16 |
B |
647 |
585 |
75.28 |
|
|
|
| 17 |
B |
430 |
389 |
3.86 |
|
|
|
| 18 |
B |
355 |
321 |
530.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14443.4 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 13062.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.444 |
0.672 |
|
0.436 |
| 2 |
S |
-0.342 |
-0.558 |
|
-0.502 |
| 3 |
N |
-0.849 |
-0.828 |
|
-0.726 |
| 4 |
N |
-0.849 |
-0.828 |
|
-0.710 |
| 5 |
H |
0.428 |
0.359 |
|
0.357 |
| 6 |
H |
0.371 |
0.412 |
|
0.400 |
| 7 |
H |
0.428 |
0.359 |
|
0.351 |
| 8 |
H |
0.371 |
0.412 |
|
0.394 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-6.211 |
6.211 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.009 |
0.003 |
-6.296 |
6.296 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.823 |
1.281 |
0.000 |
| y |
1.281 |
-25.437 |
0.000 |
| z |
0.000 |
0.000 |
-29.323 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.443 |
1.281 |
0.000 |
| y |
1.281 |
7.136 |
0.000 |
| z |
0.000 |
0.000 |
1.306 |
|
| Polar |
|---|
| 3z2-r2 | 2.613 |
|---|
| x2-y2 | -10.386 |
|---|
| xy | 1.281 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.944 |
-0.058 |
0.000 |
| y |
-0.058 |
5.527 |
0.000 |
| z |
0.000 |
0.000 |
8.538 |
<r2> (average value of r
2) Å
2
| <r2> |
101.225 |
| (<r2>)1/2 |
10.061 |