return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (Z)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-195.954305
Energy at 298.15K-195.964296
HF Energy-195.175926
Nuclear repulsion energy171.694248
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3196 3027 53.35      
2 A 3183 3015 7.48      
3 A 3174 3006 42.08      
4 A 3167 2999 29.45      
5 A 3162 2995 4.06      
6 A 3143 2977 12.78      
7 A 3142 2976 8.28      
8 A 3084 2921 24.58      
9 A 3077 2914 27.62      
10 A 3076 2913 25.53      
11 A 1730 1639 1.73      
12 A 1548 1466 13.95      
13 A 1538 1457 6.01      
14 A 1537 1456 4.52      
15 A 1525 1444 7.98      
16 A 1524 1443 4.45      
17 A 1463 1386 8.53      
18 A 1443 1367 5.67      
19 A 1434 1359 2.20      
20 A 1360 1288 2.13      
21 A 1323 1253 0.24      
22 A 1301 1232 0.11      
23 A 1187 1124 0.54      
24 A 1113 1055 5.00      
25 A 1077 1021 1.43      
26 A 1071 1014 0.04      
27 A 1062 1006 1.92      
28 A 975 923 0.78      
29 A 959 909 9.39      
30 A 882 835 1.01      
31 A 804 761 2.84      
32 A 726 688 43.88      
33 A 565 535 5.73      
34 A 478 452 1.91      
35 A 313 296 0.06      
36 A 257 243 0.02      
37 A 226 214 0.02      
38 A 105 99 0.28      
39 A 40 38 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 30485.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 28873.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.34406 0.08949 0.08123

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.095 -0.387 -0.431
H2 2.570 0.525 -0.804
H3 2.878 -1.030 -0.021
H4 1.641 -0.899 -1.282
C5 1.034 -0.057 0.624
H6 0.581 -0.980 0.993
H7 1.519 0.415 1.487
C8 -1.999 -0.717 -0.023
H9 -2.423 -1.056 -0.973
H10 -1.343 -1.504 0.349
H11 -2.834 -0.614 0.677
C12 -0.009 0.876 0.084
H13 0.337 1.889 -0.124
C14 -1.292 0.594 -0.202
H15 -1.900 1.400 -0.610

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 H11 C12 H13 C14 H15
C11.09311.09321.09231.53212.16132.15664.12774.59993.69865.05702.50752.89223.53314.3805
H21.09311.76781.76532.17633.07372.52234.79865.24014.55595.71732.75012.70383.90854.5592
H31.09321.76781.77182.18192.51082.49114.88715.38644.26395.76933.46123.87154.47835.3931
H41.09231.76531.77182.17072.51133.06723.85614.07933.45474.89342.78203.28873.46344.2754
C51.53212.17632.18192.17071.09231.09623.17143.93792.79683.90851.50102.19882.55303.5016
H62.16133.07372.51082.51131.09231.75262.78593.59172.09603.44932.14993.08892.72303.7942
H72.15662.52232.49113.06721.09621.75263.99284.87453.62944.54592.12542.48263.28414.1305
C84.12774.79864.88713.85613.17142.78593.99281.09411.08981.09442.55133.50161.50062.1995
H94.59995.24015.38644.07933.93793.59174.87451.09411.76501.75703.26784.12522.14442.5376
H103.69864.55594.26393.45472.79682.09603.62941.08981.76501.76672.74123.81582.16973.1087
H115.05705.71735.76934.89343.90853.44934.54591.09441.75701.76673.24804.11832.14722.5660
C122.50752.75013.46122.78201.50102.14992.12542.55133.26782.74123.24801.09061.34372.0814
H132.89222.70383.87153.28872.19883.08892.48263.50164.12523.81584.11831.09062.08232.3412
C143.53313.90854.47833.46342.55302.72303.28411.50062.14442.16972.14721.34372.08231.0895
H154.38054.55925.39314.27543.50163.79424.13052.19952.53763.10872.56602.08142.34121.0895

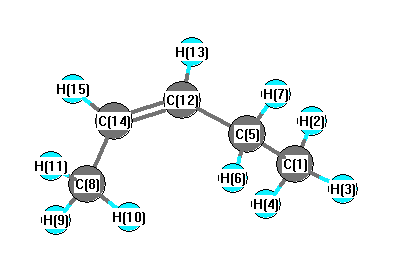
picture of 2-Pentene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.740 C1 C5 H7 109.153
C1 C5 C12 111.522 H2 C1 H3 107.917
H2 C1 H4 107.762 H2 C1 C5 110.890
H3 C1 H4 108.328 H3 C1 C5 111.328
H4 C1 C5 110.488 C5 C12 H13 115.172
C5 C12 C14 127.569 H6 C5 H7 106.415
H6 C5 C12 111.015 H7 C5 C12 108.843
C8 C14 C12 127.447 C8 C14 H15 115.325
H9 C8 H10 107.842 H9 C8 H11 106.796
H9 C8 C14 110.486 H10 C8 H11 107.972
H10 C8 C14 112.812 H11 C8 C14 110.695
C12 C14 H15 117.228 H13 C12 C14 117.232
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability