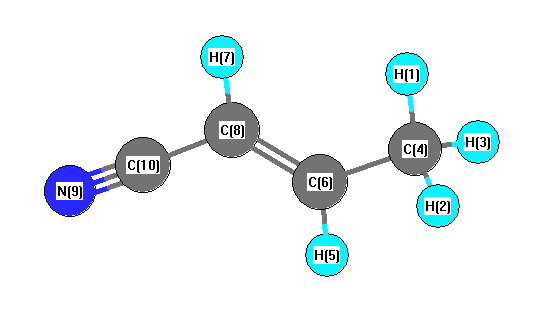
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3353 |
3033 |
6.50 |
|
|
|
| 2 |
A' |
3331 |
3012 |
10.92 |
|
|
|
| 3 |
A' |
3266 |
2954 |
22.53 |
|
|
|
| 4 |
A' |
3186 |
2881 |
25.02 |
|
|
|
| 5 |
A' |
2584 |
2337 |
48.34 |
|
|
|
| 6 |
A' |
1861 |
1684 |
27.82 |
|
|
|
| 7 |
A' |
1619 |
1464 |
16.64 |
|
|
|
| 8 |
A' |
1551 |
1403 |
1.57 |
|
|
|
| 9 |
A' |
1446 |
1308 |
3.13 |
|
|
|
| 10 |
A' |
1431 |
1294 |
0.58 |
|
|
|
| 11 |
A' |
1222 |
1106 |
0.14 |
|
|
|
| 12 |
A' |
1087 |
983 |
5.38 |
|
|
|
| 13 |
A' |
976 |
883 |
3.58 |
|
|
|
| 14 |
A' |
611 |
553 |
0.32 |
|
|
|
| 15 |
A' |
427 |
387 |
1.97 |
|
|
|
| 16 |
A' |
197 |
178 |
5.42 |
|
|
|
| 17 |
A" |
3236 |
2927 |
26.04 |
|
|
|
| 18 |
A" |
1612 |
1457 |
9.11 |
|
|
|
| 19 |
A" |
1188 |
1074 |
3.77 |
|
|
|
| 20 |
A" |
1088 |
984 |
46.98 |
|
|
|
| 21 |
A" |
892 |
807 |
5.12 |
|
|
|
| 22 |
A" |
553 |
500 |
5.99 |
|
|
|
| 23 |
A" |
203 |
183 |
1.17 |
|
|
|
| 24 |
A" |
192 |
173 |
0.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18555.8 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 16781.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.228 |
0.040 |
|
0.094 |
| 2 |
H |
0.235 |
0.040 |
|
0.094 |
| 3 |
H |
0.235 |
0.040 |
|
0.094 |
| 4 |
C |
-0.644 |
-0.043 |
|
-0.253 |
| 5 |
H |
0.243 |
0.116 |
|
0.121 |
| 6 |
C |
-0.126 |
0.025 |
|
0.115 |
| 7 |
H |
0.257 |
0.207 |
|
0.265 |
| 8 |
C |
-0.242 |
-0.391 |
|
-0.509 |
| 9 |
N |
-0.341 |
-0.535 |
|
-0.510 |
| 10 |
C |
0.157 |
0.502 |
|
0.488 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.857 |
-0.643 |
0.000 |
4.899 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
4.868 |
-0.621 |
0.000 |
4.907 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.598 |
0.831 |
0.000 |
| y |
0.831 |
-27.302 |
0.000 |
| z |
0.000 |
0.000 |
-31.353 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.270 |
0.831 |
0.000 |
| y |
0.831 |
8.173 |
0.000 |
| z |
0.000 |
0.000 |
2.097 |
|
| Polar |
|---|
| 3z2-r2 | 4.194 |
|---|
| x2-y2 | -12.295 |
|---|
| xy | 0.831 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.047 |
-1.690 |
0.000 |
| y |
-1.690 |
6.327 |
0.000 |
| z |
0.000 |
0.000 |
4.022 |
<r2> (average value of r
2) Å
2
| <r2> |
150.478 |
| (<r2>)1/2 |
12.267 |