return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5N ((E)-2-Butenenitrile)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-209.636959
Energy at 298.15K-209.641368
HF Energy-208.845982
Nuclear repulsion energy137.956170
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3216 3046 4.99      
2 A' 3201 3032 8.04      
3 A' 3178 3010 7.82      
4 A' 3078 2915 11.28      
5 A' 2194 2078 0.18      
6 A' 1713 1622 6.54      
7 A' 1528 1447 17.36      
8 A' 1451 1375 4.84      
9 A' 1349 1277 0.40      
10 A' 1335 1264 0.18      
11 A' 1156 1095 0.07      
12 A' 1052 996 5.72      
13 A' 926 877 6.75      
14 A' 558 528 0.01      
15 A' 396 376 1.16      
16 A' 176 167 3.83      
17 A" 3154 2987 9.44      
18 A" 1515 1434 11.45      
19 A" 1084 1026 0.28      
20 A" 985 933 50.10      
21 A" 793 751 0.80      
22 A" 480 454 4.15      
23 A" 188 178 1.45      
24 A" 173 164 0.27      

Unscaled Zero Point Vibrational Energy (zpe) 17438.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16516.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.26592 0.07617 0.07281

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 2.645 0.452 0.000
H2 2.651 -1.089 0.879
H3 2.651 -1.089 -0.879
C4 2.258 -0.569 0.000
H5 0.274 -1.562 0.000
C6 0.764 -0.590 0.000
H7 0.447 1.507 0.000
C8 0.000 0.516 0.000
N9 -2.604 0.415 0.000
C10 -1.429 0.455 0.000

Atom - Atom Distances (Å)
  H1 H2 H3 C4 H5 C6 H7 C8 N9 C10
H11.77331.77331.09143.11052.14942.43792.64555.24904.0737
H21.77331.75791.09372.57812.13993.51653.22095.53604.4497
H31.77331.75791.09372.57812.13993.51653.22095.53604.4497
C41.09141.09371.09372.21911.49412.75532.50574.96123.8271
H53.11052.57812.57812.21911.08893.07382.09613.49162.6398
C62.14942.13992.13991.49411.08892.12081.34463.51542.4298
H72.43793.51653.51652.75533.07382.12081.08693.24072.1508
C82.64553.22093.22092.50572.09611.34461.08692.60621.4304
N95.24905.53605.53604.96123.49163.51543.24072.60621.1758
C104.07374.44974.44973.82712.63982.42982.15081.43041.1758

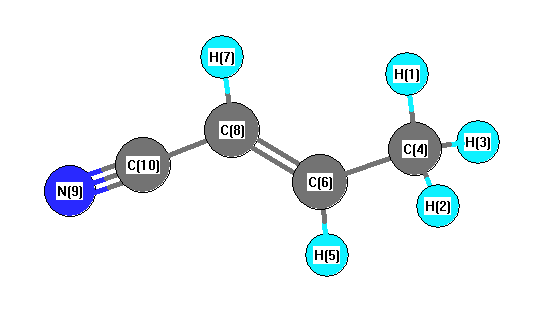
picture of (E)-2-Butenenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H2 108.494 H1 C4 H3 108.494
H1 C4 C6 111.522 H2 C4 H3 106.959
H2 C4 C6 110.613 H3 C4 C6 110.613
C4 C6 H5 117.579 C4 C6 C8 123.854
H5 C6 C8 118.567 C6 C8 H7 121.067
C6 C8 C10 122.203 H7 C8 C10 116.730
C8 C10 N9 179.508
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability