Jump to
S1C2
Energy calculated at mPW1PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -554.183942 |
| Energy at 298.15K | -554.189186 |
| Nuclear repulsion energy | 199.128001 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3258 |
3110 |
10.96 |
|
|
|
| 2 |
A |
3191 |
3046 |
6.30 |
|
|
|
| 3 |
A |
3166 |
3022 |
1.69 |
|
|
|
| 4 |
A |
1688 |
1611 |
6.15 |
|
|
|
| 5 |
A |
1432 |
1367 |
9.64 |
|
|
|
| 6 |
A |
1318 |
1258 |
0.52 |
|
|
|
| 7 |
A |
1069 |
1020 |
0.64 |
|
|
|
| 8 |
A |
1004 |
959 |
9.02 |
|
|
|
| 9 |
A |
921 |
879 |
22.28 |
|
|
|
| 10 |
A |
744 |
710 |
0.66 |
|
|
|
| 11 |
A |
632 |
603 |
13.64 |
|
|
|
| 12 |
A |
386 |
369 |
0.43 |
|
|
|
| 13 |
A |
189 |
180 |
0.00 |
|
|
|
| 14 |
A |
80 |
76 |
0.20 |
|
|
|
| 15 |
A |
3258 |
3110 |
6.54 |
|
|
|
| 16 |
A |
3187 |
3042 |
10.56 |
|
|
|
| 17 |
A |
3166 |
3021 |
4.49 |
|
|
|
| 18 |
A |
1670 |
1594 |
92.36 |
|
|
|
| 19 |
A |
1425 |
1360 |
3.86 |
|
|
|
| 20 |
A |
1306 |
1246 |
15.67 |
|
|
|
| 21 |
A |
1049 |
1002 |
11.98 |
|
|
|
| 22 |
A |
994 |
949 |
83.86 |
|
|
|
| 23 |
A |
921 |
879 |
60.15 |
|
|
|
| 24 |
A |
736 |
703 |
22.95 |
|
|
|
| 25 |
A |
601 |
573 |
21.93 |
|
|
|
| 26 |
A |
401 |
383 |
1.57 |
|
|
|
| 27 |
A |
70 |
67 |
3.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18931.2 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 18067.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.677 |
| C2 |
0.000 |
1.358 |
-0.448 |
| C3 |
0.000 |
-1.358 |
-0.448 |
| C4 |
-0.620 |
2.506 |
-0.200 |
| C5 |
0.620 |
-2.506 |
-0.200 |
| H6 |
0.613 |
1.223 |
-1.334 |
| H7 |
-0.613 |
-1.223 |
-1.334 |
| H8 |
-0.516 |
3.345 |
-0.879 |
| H9 |
-1.231 |
2.653 |
0.683 |
| H10 |
0.516 |
-3.345 |
-0.879 |
| H11 |
1.231 |
-2.653 |
0.683 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7636 | 1.7636 | 2.7267 | 2.7267 | 2.4324 | 2.4324 | 3.7247 | 2.9247 | 3.7247 | 2.9247 |
C2 | 1.7636 | | 2.7156 | 1.3282 | 3.9212 | 1.0856 | 2.7964 | 2.0973 | 2.1149 | 4.7501 | 4.3455 | C3 | 1.7636 | 2.7156 | | 3.9212 | 1.3282 | 2.7964 | 1.0856 | 4.7501 | 4.3455 | 2.0973 | 2.1149 | C4 | 2.7267 | 1.3282 | 3.9212 | | 5.1633 | 2.1099 | 3.8976 | 1.0837 | 1.0841 | 5.9984 | 5.5519 | C5 | 2.7267 | 3.9212 | 1.3282 | 5.1633 | | 3.8976 | 2.1099 | 5.9984 | 5.5519 | 1.0837 | 1.0841 | H6 | 2.4324 | 1.0856 | 2.7964 | 2.1099 | 3.8976 | | 2.7354 | 2.4459 | 3.0845 | 4.5910 | 4.4131 | H7 | 2.4324 | 2.7964 | 1.0856 | 3.8976 | 2.1099 | 2.7354 | | 4.5910 | 4.4131 | 2.4459 | 3.0845 | H8 | 3.7247 | 2.0973 | 4.7501 | 1.0837 | 5.9984 | 2.4459 | 4.5910 | | 1.8519 | 6.7681 | 6.4392 | H9 | 2.9247 | 2.1149 | 4.3455 | 1.0841 | 5.5519 | 3.0845 | 4.4131 | 1.8519 | | 6.4392 | 5.8495 | H10 | 3.7247 | 4.7501 | 2.0973 | 5.9984 | 1.0837 | 4.5910 | 2.4459 | 6.7681 | 6.4392 | | 1.8519 | H11 | 2.9247 | 4.3455 | 2.1149 | 5.5519 | 1.0841 | 4.4131 | 3.0845 | 6.4392 | 5.8495 | 1.8519 | |
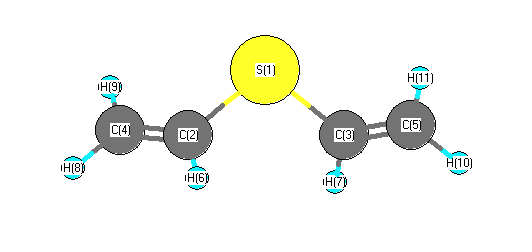 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.124 |
|
S1 |
C2 |
H6 |
115.153 |
| S1 |
C3 |
C5 |
123.124 |
|
S1 |
C3 |
H7 |
115.153 |
| C2 |
S1 |
C3 |
100.690 |
|
C2 |
C4 |
H8 |
120.473 |
| C2 |
C4 |
H9 |
122.166 |
|
C3 |
C5 |
H10 |
120.473 |
| C3 |
C5 |
H11 |
122.166 |
|
C4 |
C2 |
H6 |
121.550 |
| C5 |
C3 |
H7 |
121.550 |
|
H8 |
C4 |
H9 |
117.355 |
| H10 |
C5 |
H11 |
117.355 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.221 |
|
|
|
| 2 |
C |
-0.394 |
|
|
|
| 3 |
C |
-0.394 |
|
|
|
| 4 |
C |
-0.426 |
|
|
|
| 5 |
C |
-0.426 |
|
|
|
| 6 |
H |
0.249 |
|
|
|
| 7 |
H |
0.249 |
|
|
|
| 8 |
H |
0.228 |
|
|
|
| 9 |
H |
0.232 |
|
|
|
| 10 |
H |
0.228 |
|
|
|
| 11 |
H |
0.232 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.229 |
1.229 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
185.020 |
| (<r2>)1/2 |
13.602 |
Jump to
S1C1
Energy calculated at mPW1PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -554.186900 |
| Energy at 298.15K | -554.192385 |
| HF Energy | -554.186900 |
| Nuclear repulsion energy | 202.455508 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3259 |
3111 |
10.91 |
|
|
|
| 2 |
A |
3259 |
3110 |
5.92 |
|
|
|
| 3 |
A |
3197 |
3052 |
5.62 |
|
|
|
| 4 |
A |
3182 |
3037 |
7.25 |
|
|
|
| 5 |
A |
3168 |
3024 |
2.29 |
|
|
|
| 6 |
A |
3164 |
3020 |
3.91 |
|
|
|
| 7 |
A |
1681 |
1604 |
41.67 |
|
|
|
| 8 |
A |
1671 |
1595 |
33.68 |
|
|
|
| 9 |
A |
1432 |
1367 |
4.30 |
|
|
|
| 10 |
A |
1429 |
1364 |
7.47 |
|
|
|
| 11 |
A |
1319 |
1259 |
1.78 |
|
|
|
| 12 |
A |
1307 |
1248 |
8.94 |
|
|
|
| 13 |
A |
1071 |
1022 |
10.47 |
|
|
|
| 14 |
A |
1044 |
997 |
14.26 |
|
|
|
| 15 |
A |
1007 |
961 |
29.48 |
|
|
|
| 16 |
A |
992 |
946 |
37.25 |
|
|
|
| 17 |
A |
933 |
891 |
41.03 |
|
|
|
| 18 |
A |
893 |
853 |
49.36 |
|
|
|
| 19 |
A |
751 |
717 |
13.02 |
|
|
|
| 20 |
A |
703 |
671 |
5.02 |
|
|
|
| 21 |
A |
635 |
606 |
19.60 |
|
|
|
| 22 |
A |
610 |
582 |
17.31 |
|
|
|
| 23 |
A |
465 |
444 |
0.56 |
|
|
|
| 24 |
A |
371 |
354 |
0.48 |
|
|
|
| 25 |
A |
223 |
212 |
1.14 |
|
|
|
| 26 |
A |
150 |
143 |
4.94 |
|
|
|
| 27 |
A |
91 |
87 |
1.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19003.6 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 18137.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.014 |
-0.976 |
-0.003 |
| C2 |
1.082 |
0.371 |
0.371 |
| C3 |
-1.562 |
-0.196 |
-0.015 |
| C4 |
2.260 |
0.551 |
-0.215 |
| C5 |
-1.822 |
1.103 |
-0.133 |
| H6 |
0.744 |
1.005 |
1.186 |
| H7 |
-2.362 |
-0.920 |
0.094 |
| H8 |
2.932 |
1.329 |
0.129 |
| H9 |
2.597 |
-0.072 |
-1.036 |
| H10 |
-1.043 |
1.843 |
-0.275 |
| H11 |
-2.845 |
1.457 |
-0.098 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7591 | 1.7587 | 2.7241 | 2.7766 | 2.4228 | 2.3792 | 3.7208 | 2.9248 | 3.0233 | 3.7557 |
C2 | 1.7591 | | 2.7310 | 1.3283 | 3.0364 | 1.0864 | 3.6886 | 2.0975 | 2.1144 | 2.6645 | 4.1011 | C3 | 1.7587 | 2.7310 | | 3.8992 | 1.3294 | 2.8635 | 1.0853 | 4.7474 | 4.2837 | 2.1196 | 2.0941 | C4 | 2.7241 | 1.3283 | 3.8992 | | 4.1198 | 2.1140 | 4.8607 | 1.0841 | 1.0838 | 3.5477 | 5.1863 | C5 | 2.7766 | 3.0364 | 1.3294 | 4.1198 | | 2.8866 | 2.1062 | 4.7661 | 4.6601 | 1.0836 | 1.0835 | H6 | 2.4228 | 1.0864 | 2.8635 | 2.1140 | 2.8866 | | 3.8138 | 2.4519 | 3.0871 | 2.4559 | 3.8386 | H7 | 2.3792 | 3.6886 | 1.0853 | 4.8607 | 2.1062 | 3.8138 | | 5.7523 | 5.1565 | 3.0841 | 2.4333 | H8 | 3.7208 | 2.0975 | 4.7474 | 1.0841 | 4.7661 | 2.4519 | 5.7523 | | 1.8523 | 4.0284 | 5.7827 | H9 | 2.9248 | 2.1144 | 4.2837 | 1.0838 | 4.6601 | 3.0871 | 5.1565 | 1.8523 | | 4.1828 | 5.7297 | H10 | 3.0233 | 2.6645 | 2.1196 | 3.5477 | 1.0836 | 2.4559 | 3.0841 | 4.0284 | 4.1828 | | 1.8511 | H11 | 3.7557 | 4.1011 | 2.0941 | 5.1863 | 1.0835 | 3.8386 | 2.4333 | 5.7827 | 5.7297 | 1.8511 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.244 |
|
S1 |
C2 |
H6 |
114.665 |
| S1 |
C3 |
C5 |
127.541 |
|
S1 |
C3 |
H7 |
111.351 |
| C2 |
S1 |
C3 |
101.853 |
|
C2 |
C4 |
H8 |
120.464 |
| C2 |
C4 |
H9 |
122.138 |
|
C3 |
C5 |
H10 |
122.575 |
| C3 |
C5 |
H11 |
120.085 |
|
C4 |
C2 |
H6 |
121.885 |
| C5 |
C3 |
H7 |
121.107 |
|
H8 |
C4 |
H9 |
117.393 |
| H10 |
C5 |
H11 |
117.337 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.232 |
|
|
|
| 2 |
C |
-0.408 |
|
|
|
| 3 |
C |
-0.382 |
|
|
|
| 4 |
C |
-0.420 |
|
|
|
| 5 |
C |
-0.454 |
|
|
|
| 6 |
H |
0.255 |
|
|
|
| 7 |
H |
0.247 |
|
|
|
| 8 |
H |
0.229 |
|
|
|
| 9 |
H |
0.236 |
|
|
|
| 10 |
H |
0.235 |
|
|
|
| 11 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.039 |
1.044 |
0.203 |
1.064 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.700 |
1.745 |
-0.875 |
| y |
1.745 |
-37.458 |
1.201 |
| z |
-0.875 |
1.201 |
-40.618 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.338 |
1.745 |
-0.875 |
| y |
1.745 |
-0.799 |
1.201 |
| z |
-0.875 |
1.201 |
-5.539 |
|
| Polar |
|---|
| 3z2-r2 | -11.078 |
|---|
| x2-y2 | 4.759 |
|---|
| xy | 1.745 |
|---|
| xz | -0.875 |
|---|
| yz | 1.201 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.070 |
-0.060 |
-0.735 |
| y |
-0.060 |
9.469 |
0.194 |
| z |
-0.735 |
0.194 |
5.315 |
<r2> (average value of r
2) Å
2
| <r2> |
164.306 |
| (<r2>)1/2 |
12.818 |
