return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (Butane, 1-methoxy-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-272.227575
Energy at 298.15K-272.241038
HF Energy-271.227619
Nuclear repulsion energy248.924306
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3193 3024 28.99      
2 A' 3167 3000 32.96      
3 A' 3088 2924 40.50      
4 A' 3077 2914 23.63      
5 A' 3071 2909 19.62      
6 A' 3032 2871 70.68      
7 A' 3006 2847 39.17      
8 A' 1571 1488 2.86      
9 A' 1551 1469 6.59      
10 A' 1547 1465 13.93      
11 A' 1534 1453 0.80      
12 A' 1529 1448 0.25      
13 A' 1518 1438 0.01      
14 A' 1464 1387 24.80      
15 A' 1450 1373 6.89      
16 A' 1414 1339 0.76      
17 A' 1314 1244 3.84      
18 A' 1255 1188 35.01      
19 A' 1196 1132 124.93      
20 A' 1156 1095 19.79      
21 A' 1113 1054 0.84      
22 A' 1073 1016 1.76      
23 A' 1011 958 23.21      
24 A' 930 881 6.65      
25 A' 503 477 1.11      
26 A' 392 372 3.16      
27 A' 323 306 0.73      
28 A' 142 134 1.04      
29 A" 3167 2999 54.28      
30 A" 3146 2980 28.23      
31 A" 3114 2950 4.69      
32 A" 3096 2932 59.66      
33 A" 3049 2888 59.42      
34 A" 1541 1460 10.01      
35 A" 1529 1449 8.52      
36 A" 1359 1287 0.06      
37 A" 1347 1276 0.02      
38 A" 1283 1215 2.39      
39 A" 1228 1163 6.61      
40 A" 1200 1136 0.18      
41 A" 977 925 0.04      
42 A" 835 790 0.76      
43 A" 751 712 3.06      
44 A" 259 246 1.43      
45 A" 242 229 2.58      
46 A" 139 131 1.12      
47 A" 101 96 0.86      
48 A" 74 70 1.62      

Unscaled Zero Point Vibrational Energy (zpe) 37026.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 35068.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.54730 0.04065 0.03922

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.489 2.758 0.000
C2 1.431 1.233 0.000
C3 -1.544 -2.606 0.000
O4 -1.419 -1.202 0.000
C5 -0.063 -0.809 0.000
C6 0.000 0.705 0.000
H7 -2.610 -2.833 0.000
H8 2.520 3.121 0.000
H9 0.990 3.166 0.883
H10 0.990 3.166 -0.883
H11 1.962 0.846 0.878
H12 1.962 0.846 -0.878
H13 -1.082 -3.051 0.891
H14 -1.082 -3.051 -0.891
H15 -0.537 1.074 -0.880
H16 -0.537 1.074 0.880
H17 0.449 -1.213 -0.888
H18 0.449 -1.213 0.888

Atom - Atom Distances (Å)
  C1 C2 C3 O4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.52596.16234.91343.88982.53656.93211.09271.09311.09312.15612.15616.41426.41422.77802.77804.19974.1997
C21.52594.85743.74922.53031.52585.73242.17892.17062.17061.09621.09625.04575.04572.16232.16232.78142.7814
C36.16234.85741.40922.32913.65311.08917.02236.36596.36594.99824.99821.09811.09813.91523.91522.58942.5894
O44.91343.74921.40921.41202.37702.01875.84845.06665.06664.04944.04942.07932.07932.59472.59472.06872.0687
C53.88982.53032.32911.41201.51463.25294.70234.20604.20602.75852.75852.61872.61872.13142.13141.10211.1021
C62.53651.52583.65312.37701.51464.39563.49102.79662.79662.15392.15394.00814.00811.09521.09522.16032.1603
H76.93215.73241.08912.01873.25294.39567.85837.05177.05175.93325.93321.78221.78224.50864.50863.57353.5735
H81.09272.17897.02235.84844.70233.49107.85831.76661.76662.50102.50107.20067.20063.78293.78294.88424.8842
H91.09312.17066.36595.06664.20602.79667.05171.76661.76562.51533.07036.55316.78903.13372.59094.75464.4125
H101.09312.17066.36595.06664.20602.79667.05171.76661.76563.07032.51536.78906.55312.59093.13374.41254.7546
H112.15611.09624.99824.04942.75852.15395.93322.50102.51533.07031.75574.94455.25143.06392.50943.10592.5549
H122.15611.09624.99824.04942.75852.15395.93322.50103.07032.51531.75575.25144.94452.50943.06392.55493.1059
H136.41425.04571.09812.07932.61874.00811.78227.20066.55316.78904.94455.25141.78234.52134.16002.98122.3919
H146.41425.04571.09812.07932.61874.00811.78227.20066.78906.55315.25144.94451.78234.16004.52132.39192.9812
H152.77802.16233.91522.59472.13141.09524.50863.78293.13372.59093.06392.50944.52134.16001.76002.49023.0542
H162.77802.16233.91522.59472.13141.09524.50863.78292.59093.13372.50943.06394.16004.52131.76003.05422.4902
H174.19972.78142.58942.06871.10212.16033.57354.88424.75464.41253.10592.55492.98122.39192.49023.05421.7766
H184.19972.78142.58942.06871.10212.16033.57354.88424.41254.75462.55493.10592.39192.98123.05422.49021.7766

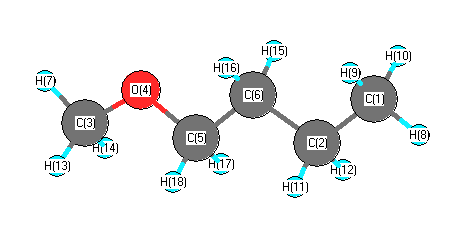
picture of Butane, 1-methoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C6 112.437 C1 C2 H11 109.533
C1 C2 H12 109.533 C2 C1 H8 111.556
C2 C1 H9 110.861 C2 C1 H10 110.861
C2 C6 C5 112.652 C2 C6 H15 110.089
C2 C6 H16 110.089 C3 O4 C5 111.290
O4 C3 H7 107.109 O4 C3 H13 111.451
O4 C3 H14 111.451 O4 C5 C6 108.569
O4 C5 H17 110.129 O4 C5 H18 110.129
C5 C6 H15 108.445 C5 C6 H16 108.445
C6 C2 H11 109.368 C6 C2 H12 109.368
C6 C5 H17 110.300 C6 C5 H18 110.300
H7 C3 H13 109.144 H7 C3 H14 109.144
H8 C1 H9 107.842 H8 C1 H10 107.842
H9 C1 H10 107.720 H11 C2 H12 106.420
H13 C3 H14 108.494 H15 C6 H16 106.935
H17 C5 H18 107.414
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability