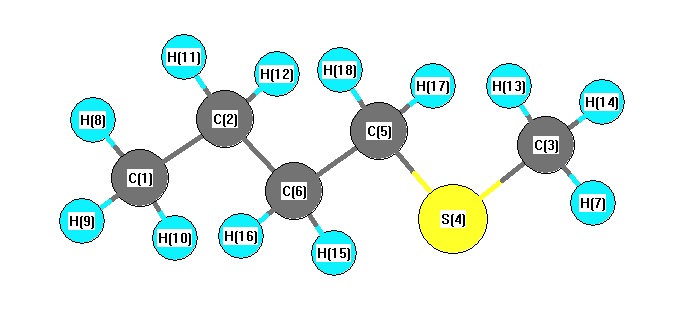
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3163 |
3019 |
9.46 |
|
|
|
| 2 |
A' |
3128 |
2985 |
43.59 |
|
|
|
| 3 |
A' |
3060 |
2920 |
22.60 |
|
|
|
| 4 |
A' |
3057 |
2917 |
106.65 |
|
|
|
| 5 |
A' |
3052 |
2913 |
25.31 |
|
|
|
| 6 |
A' |
3047 |
2908 |
3.76 |
|
|
|
| 7 |
A' |
3041 |
2902 |
9.54 |
|
|
|
| 8 |
A' |
1529 |
1459 |
9.14 |
|
|
|
| 9 |
A' |
1517 |
1448 |
1.44 |
|
|
|
| 10 |
A' |
1506 |
1437 |
0.05 |
|
|
|
| 11 |
A' |
1505 |
1436 |
4.91 |
|
|
|
| 12 |
A' |
1498 |
1430 |
9.92 |
|
|
|
| 13 |
A' |
1430 |
1365 |
4.31 |
|
|
|
| 14 |
A' |
1414 |
1349 |
4.13 |
|
|
|
| 15 |
A' |
1387 |
1324 |
2.80 |
|
|
|
| 16 |
A' |
1351 |
1289 |
8.56 |
|
|
|
| 17 |
A' |
1268 |
1210 |
16.02 |
|
|
|
| 18 |
A' |
1140 |
1088 |
3.34 |
|
|
|
| 19 |
A' |
1088 |
1039 |
0.98 |
|
|
|
| 20 |
A' |
1052 |
1005 |
0.83 |
|
|
|
| 21 |
A' |
998 |
952 |
8.35 |
|
|
|
| 22 |
A' |
915 |
873 |
1.24 |
|
|
|
| 23 |
A' |
784 |
748 |
1.96 |
|
|
|
| 24 |
A' |
736 |
702 |
2.41 |
|
|
|
| 25 |
A' |
424 |
404 |
0.45 |
|
|
|
| 26 |
A' |
317 |
303 |
0.58 |
|
|
|
| 27 |
A' |
237 |
226 |
0.00 |
|
|
|
| 28 |
A' |
116 |
111 |
0.95 |
|
|
|
| 29 |
A" |
3141 |
2998 |
13.65 |
|
|
|
| 30 |
A" |
3124 |
2982 |
83.72 |
|
|
|
| 31 |
A" |
3109 |
2967 |
34.34 |
|
|
|
| 32 |
A" |
3086 |
2946 |
0.97 |
|
|
|
| 33 |
A" |
3067 |
2927 |
4.04 |
|
|
|
| 34 |
A" |
1520 |
1451 |
9.83 |
|
|
|
| 35 |
A" |
1491 |
1423 |
11.29 |
|
|
|
| 36 |
A" |
1353 |
1291 |
0.29 |
|
|
|
| 37 |
A" |
1325 |
1265 |
0.50 |
|
|
|
| 38 |
A" |
1242 |
1185 |
0.51 |
|
|
|
| 39 |
A" |
1097 |
1047 |
2.23 |
|
|
|
| 40 |
A" |
995 |
950 |
8.77 |
|
|
|
| 41 |
A" |
937 |
894 |
1.32 |
|
|
|
| 42 |
A" |
802 |
765 |
0.25 |
|
|
|
| 43 |
A" |
744 |
710 |
5.75 |
|
|
|
| 44 |
A" |
249 |
238 |
0.06 |
|
|
|
| 45 |
A" |
183 |
175 |
0.53 |
|
|
|
| 46 |
A" |
130 |
124 |
0.03 |
|
|
|
| 47 |
A" |
93 |
88 |
0.71 |
|
|
|
| 48 |
A" |
57 |
54 |
0.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35751.3 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 34121.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.654 |
|
|
|
| 2 |
C |
-0.431 |
|
|
|
| 3 |
C |
-0.821 |
|
|
|
| 4 |
S |
0.135 |
|
|
|
| 5 |
C |
-0.586 |
|
|
|
| 6 |
C |
-0.453 |
|
|
|
| 7 |
H |
0.255 |
|
|
|
| 8 |
H |
0.225 |
|
|
|
| 9 |
H |
0.222 |
|
|
|
| 10 |
H |
0.222 |
|
|
|
| 11 |
H |
0.217 |
|
|
|
| 12 |
H |
0.217 |
|
|
|
| 13 |
H |
0.247 |
|
|
|
| 14 |
H |
0.247 |
|
|
|
| 15 |
H |
0.234 |
|
|
|
| 16 |
H |
0.234 |
|
|
|
| 17 |
H |
0.245 |
|
|
|
| 18 |
H |
0.245 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.224 |
1.253 |
0.000 |
1.752 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.194 |
3.267 |
0.000 |
| y |
3.267 |
-44.893 |
0.000 |
| z |
0.000 |
0.000 |
-48.344 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.575 |
3.267 |
0.000 |
| y |
3.267 |
3.376 |
0.000 |
| z |
0.000 |
0.000 |
-1.800 |
|
| Polar |
|---|
| 3z2-r2 | -3.600 |
|---|
| x2-y2 | -3.301 |
|---|
| xy | 3.267 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
369.508 |
| (<r2>)1/2 |
19.223 |