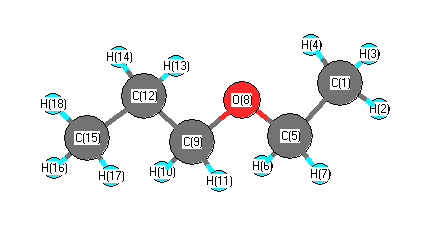
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3124 |
3007 |
33.75 |
|
|
|
| 2 |
A' |
3112 |
2996 |
39.81 |
|
|
|
| 3 |
A' |
3048 |
2935 |
28.58 |
|
|
|
| 4 |
A' |
3047 |
2933 |
25.61 |
|
|
|
| 5 |
A' |
3034 |
2921 |
33.52 |
|
|
|
| 6 |
A' |
2969 |
2858 |
104.67 |
|
|
|
| 7 |
A' |
2953 |
2843 |
21.07 |
|
|
|
| 8 |
A' |
1548 |
1490 |
3.10 |
|
|
|
| 9 |
A' |
1529 |
1472 |
7.42 |
|
|
|
| 10 |
A' |
1519 |
1462 |
6.17 |
|
|
|
| 11 |
A' |
1513 |
1457 |
2.66 |
|
|
|
| 12 |
A' |
1504 |
1448 |
1.17 |
|
|
|
| 13 |
A' |
1463 |
1408 |
3.46 |
|
|
|
| 14 |
A' |
1423 |
1370 |
2.10 |
|
|
|
| 15 |
A' |
1419 |
1366 |
45.55 |
|
|
|
| 16 |
A' |
1397 |
1345 |
15.54 |
|
|
|
| 17 |
A' |
1336 |
1286 |
6.71 |
|
|
|
| 18 |
A' |
1188 |
1144 |
45.03 |
|
|
|
| 19 |
A' |
1171 |
1127 |
201.92 |
|
|
|
| 20 |
A' |
1123 |
1081 |
6.46 |
|
|
|
| 21 |
A' |
1074 |
1034 |
6.38 |
|
|
|
| 22 |
A' |
1045 |
1006 |
10.23 |
|
|
|
| 23 |
A' |
918 |
884 |
7.55 |
|
|
|
| 24 |
A' |
905 |
871 |
5.44 |
|
|
|
| 25 |
A' |
494 |
476 |
3.71 |
|
|
|
| 26 |
A' |
407 |
392 |
0.91 |
|
|
|
| 27 |
A' |
306 |
294 |
0.68 |
|
|
|
| 28 |
A' |
139 |
133 |
0.34 |
|
|
|
| 29 |
A" |
3127 |
3010 |
35.98 |
|
|
|
| 30 |
A" |
3106 |
2990 |
75.38 |
|
|
|
| 31 |
A" |
3079 |
2964 |
2.39 |
|
|
|
| 32 |
A" |
2990 |
2878 |
49.52 |
|
|
|
| 33 |
A" |
2981 |
2869 |
84.16 |
|
|
|
| 34 |
A" |
1516 |
1459 |
9.73 |
|
|
|
| 35 |
A" |
1495 |
1439 |
8.15 |
|
|
|
| 36 |
A" |
1328 |
1278 |
0.78 |
|
|
|
| 37 |
A" |
1312 |
1263 |
2.27 |
|
|
|
| 38 |
A" |
1279 |
1231 |
1.91 |
|
|
|
| 39 |
A" |
1206 |
1161 |
7.88 |
|
|
|
| 40 |
A" |
1181 |
1137 |
0.65 |
|
|
|
| 41 |
A" |
909 |
875 |
1.85 |
|
|
|
| 42 |
A" |
826 |
796 |
0.71 |
|
|
|
| 43 |
A" |
768 |
740 |
1.95 |
|
|
|
| 44 |
A" |
256 |
246 |
0.90 |
|
|
|
| 45 |
A" |
236 |
228 |
0.01 |
|
|
|
| 46 |
A" |
153 |
148 |
3.54 |
|
|
|
| 47 |
A" |
93 |
90 |
1.25 |
|
|
|
| 48 |
A" |
68 |
65 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36306.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 34952.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.668 |
|
|
|
| 2 |
H |
0.215 |
|
|
|
| 3 |
H |
0.227 |
|
|
|
| 4 |
H |
0.227 |
|
|
|
| 5 |
C |
-0.213 |
|
|
|
| 6 |
H |
0.188 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
| 8 |
O |
-0.336 |
|
|
|
| 9 |
C |
-0.186 |
|
|
|
| 10 |
H |
0.186 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
| 12 |
C |
-0.462 |
|
|
|
| 13 |
H |
0.223 |
|
|
|
| 14 |
H |
0.223 |
|
|
|
| 15 |
C |
-0.662 |
|
|
|
| 16 |
H |
0.218 |
|
|
|
| 17 |
H |
0.218 |
|
|
|
| 18 |
H |
0.227 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.882 |
-0.501 |
0.000 |
1.015 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.884 |
1.952 |
0.000 |
| y |
1.952 |
-39.138 |
0.000 |
| z |
0.000 |
0.000 |
-39.448 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.409 |
1.952 |
0.000 |
| y |
1.952 |
0.029 |
0.000 |
| z |
0.000 |
0.000 |
-0.437 |
|
| Polar |
|---|
| 3z2-r2 | -0.875 |
|---|
| x2-y2 | 0.253 |
|---|
| xy | 1.952 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
285.654 |
| (<r2>)1/2 |
16.901 |