return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 (1,4-Cyclohexadiene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-232.784144
Energy at 298.15K-232.792590
HF Energy-231.873307
Nuclear repulsion energy217.843034
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3204 3035 0.00      
2 Ag 3045 2884 0.00      
3 Ag 1747 1655 0.00      
4 Ag 1507 1428 0.00      
5 Ag 1243 1177 0.00      
6 Ag 878 832 0.00      
7 Ag 540 512 0.00      
8 Au 1236 1170 0.00      
9 Au 957 907 0.00      
10 Au 365 345 0.00      
11 B1g 3181 3012 0.00      
12 B1g 1430 1355 0.00      
13 B1g 1382 1309 0.00      
14 B1g 1075 1018 0.00      
15 B1g 576 546 0.00      
16 B1u 3073 2910 39.28      
17 B1u 1003 950 28.15      
18 B1u 627 594 80.22      
19 B1u 64 61 0.64      
20 B2g 3074 2912 0.00      
21 B2g 992 940 0.00      
22 B2g 954 903 0.00      
23 B2g 338 320 0.00      
24 B2u 3203 3034 71.81      
25 B2u 1703 1613 3.45      
26 B2u 1411 1337 1.06      
27 B2u 1195 1131 0.10      
28 B2u 968 917 0.57      
29 B3g 1243 1177 0.00      
30 B3g 719 681 0.00      
31 B3u 3181 3012 14.56      
32 B3u 3047 2886 71.02      
33 B3u 1510 1430 9.21      
34 B3u 1453 1376 0.43      
35 B3u 970 919 0.73      
36 B3u 922 873 10.33      

Unscaled Zero Point Vibrational Energy (zpe) 27007.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 25578.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.17258 0.16280 0.08640

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.501 0.000 0.000
C2 -1.501 0.000 0.000
C3 -0.670 1.251 0.000
C4 0.670 1.251 0.000
C5 -0.670 -1.251 0.000
C6 0.670 -1.251 0.000
H7 1.201 -2.201 0.000
H8 -1.201 -2.201 0.000
H9 1.201 2.201 0.000
H10 -1.201 2.201 0.000
H11 2.172 0.000 0.871
H12 -2.172 0.000 0.871
H13 2.172 0.000 -0.871
H14 -2.172 0.000 -0.871

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C13.00292.50601.50182.50601.50182.22163.48542.22163.48541.09923.77521.09923.7752
C23.00291.50182.50601.50182.50603.48542.22163.48542.22163.77521.09923.77521.0992
C32.50601.50181.34032.50152.83793.92643.49252.09861.08863.22502.13973.22502.1397
C41.50182.50601.34032.83792.50153.49253.92641.08862.09862.13973.22502.13973.2250
C52.50601.50182.50152.83791.34032.09861.08863.92643.49253.22502.13973.22502.1397
C61.50182.50602.83792.50151.34031.08862.09863.49253.92642.13973.22502.13973.2250
H72.22163.48543.92643.49252.09861.08862.40184.40245.01502.55874.12072.55874.1207
H83.48542.22163.49253.92641.08862.09862.40185.01504.40244.12072.55874.12072.5587
H92.22163.48542.09861.08863.92643.49254.40245.01502.40182.55874.12072.55874.1207
H103.48542.22161.08862.09863.49253.92645.01504.40242.40184.12072.55874.12072.5587
H111.09923.77523.22502.13973.22502.13972.55874.12072.55874.12074.34391.74204.6801
H123.77521.09922.13973.22502.13973.22504.12072.55874.12072.55874.34394.68011.7420
H131.09923.77523.22502.13973.22502.13972.55874.12072.55874.12071.74204.68014.3439
H143.77521.09922.13973.22502.13973.22504.12072.55874.12072.55874.68011.74204.3439

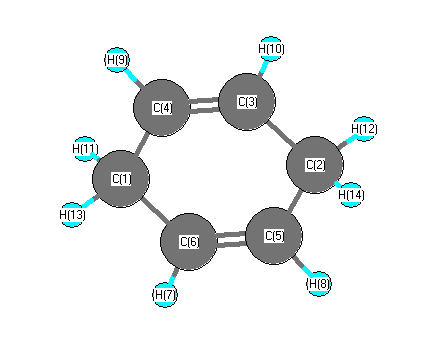
picture of 1,4-Cyclohexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 C3 123.610 C1 C4 H9 117.210
C1 C6 C5 123.610 C1 C6 H7 117.210
C2 C3 C4 123.610 C2 C3 H10 117.210
C2 C5 C6 123.610 C2 C5 H8 117.210
C3 C2 C5 112.780 C3 C2 H12 109.734
C3 C2 H14 109.734 C3 C4 H9 119.180
C4 C1 C6 112.780 C4 C1 H11 109.734
C4 C1 H13 109.734 C4 C3 H10 119.180
C5 C2 H12 109.734 C5 C2 H14 109.734
C5 C6 H7 119.180 C6 C1 H11 109.734
C6 C1 H13 109.734 C6 C5 H8 119.180
H11 C1 H13 104.824 H12 C2 H14 104.824
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability