return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2OH (Ethanol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-154.640175
Energy at 298.15K-154.646910
HF Energy-154.110074
Nuclear repulsion energy82.001235
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3858 3654 17.31 93.71 0.31 0.48
2 A' 3184 3016 24.89 58.74 0.75 0.86
3 A' 3092 2929 14.00 120.60 0.01 0.03
4 A' 3043 2882 69.13 111.93 0.12 0.21
5 A' 1576 1493 1.42 4.51 0.73 0.84
6 A' 1543 1461 4.01 16.99 0.75 0.86
7 A' 1500 1421 11.56 4.80 0.39 0.56
8 A' 1438 1362 5.53 0.43 0.53 0.70
9 A' 1308 1238 91.30 4.10 0.74 0.85
10 A' 1142 1081 26.30 6.12 0.63 0.77
11 A' 1076 1019 49.24 3.65 0.15 0.26
12 A' 932 883 9.90 5.30 0.36 0.53
13 A' 427 404 13.47 0.30 0.64 0.78
14 A" 3192 3023 27.19 46.70 0.75 0.86
15 A" 3085 2922 60.22 93.33 0.75 0.86
16 A" 1523 1442 8.07 11.00 0.75 0.86
17 A" 1333 1262 0.03 10.05 0.75 0.86
18 A" 1211 1147 5.15 1.65 0.75 0.86
19 A" 837 793 0.00 0.07 0.75 0.86
20 A" 307 291 96.78 2.73 0.75 0.86
21 A" 258 244 57.44 1.03 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 17931.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16983.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.16835 0.31626 0.27456

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.166 -0.404 0.000
C2 0.000 0.558 0.000
O3 -1.189 -0.221 0.000
H4 -1.940 0.377 0.000
H5 2.116 0.137 0.000
H6 1.125 -1.041 0.885
H7 1.125 -1.041 -0.885
H8 0.044 1.203 0.888
H9 0.044 1.203 -0.888

Atom - Atom Distances (Å)
  C1 C2 O3 H4 H5 H6 H7 H8 H9
C11.51182.36293.20301.09241.09101.09102.15162.1516
C21.51181.42161.94822.15732.14622.14621.09821.0982
O32.36291.42160.95983.32442.61032.61032.08262.0826
H43.20301.94820.95984.06283.49153.49152.32522.3252
H51.09242.15733.32444.06281.77501.77502.49322.4932
H61.09102.14622.61033.49151.77501.76942.49093.0571
H71.09102.14622.61033.49151.77501.76943.05712.4909
H82.15161.09822.08262.32522.49322.49093.05711.7753
H92.15161.09822.08262.32522.49323.05712.49091.7753

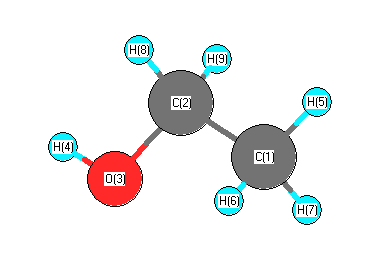
picture of Ethanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 107.277 C1 C2 H8 110.032
C1 C2 H9 110.032 C2 C1 H5 110.838
C2 C1 H6 110.031 C2 C1 H7 110.031
C2 O3 H4 108.227 O3 C2 H8 110.828
O3 C2 H9 110.828 H5 C1 H6 108.763
H5 C1 H7 108.763 H6 C1 H7 108.360
H8 C2 H9 107.856
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability