return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5COOH (benzoic acid)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-419.885974
Energy at 298.15K-419.892765
HF Energy-418.408733
Nuclear repulsion energy404.706556
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3778 3578 78.46      
2 A' 3249 3078 2.23      
3 A' 3240 3069 5.37      
4 A' 3229 3059 12.91      
5 A' 3220 3050 10.95      
6 A' 3208 3038 0.25      
7 A' 1830 1733 255.76      
8 A' 1664 1576 7.82      
9 A' 1642 1555 6.46      
10 A' 1538 1457 4.50      
11 A' 1491 1412 19.84      
12 A' 1466 1389 1.98      
13 A' 1412 1338 112.12      
14 A' 1350 1278 6.66      
15 A' 1245 1179 235.28      
16 A' 1213 1149 28.69      
17 A' 1198 1135 0.12      
18 A' 1135 1075 57.39      
19 A' 1106 1047 52.47      
20 A' 1054 998 12.65      
21 A' 1013 959 1.12      
22 A' 782 741 15.47      
23 A' 642 608 50.68      
24 A' 626 593 0.05      
25 A' 501 474 5.85      
26 A' 387 366 4.91      
27 A' 221 209 1.21      
28 A" 889 842 0.30      
29 A" 878 832 0.09      
30 A" 871 825 0.80      
31 A" 831 787 0.08      
32 A" 789 747 4.34      
33 A" 714 676 172.08      
34 A" 614 581 73.06      
35 A" 442 419 0.04      
36 A" 407 386 8.71      
37 A" 391 370 1.26      
38 A" 156 148 1.47      
39 A" 57 54 0.57      

Unscaled Zero Point Vibrational Energy (zpe) 25239.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23904.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.12900 0.04092 0.03107

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.222 0.000
C2 1.276 -0.355 0.000
C3 1.408 -1.743 0.000
C4 0.271 -2.557 0.000
C5 -1.001 -1.977 0.000
C6 -1.143 -0.588 0.000
C7 -0.085 1.707 0.000
O8 0.867 2.455 0.000
O9 -1.361 2.156 0.000
H10 2.146 0.295 0.000
H11 2.398 -2.191 0.000
H12 0.375 -3.638 0.000
H13 -1.886 -2.608 0.000
H14 -2.127 -0.132 0.000
H15 -1.307 3.122 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 O9 H10 H11 H12 H13 H14 H15
C11.40062.41752.79192.41621.40101.48752.39562.36452.14733.40213.87853.40072.15643.1803
C21.40061.39432.42052.79612.43042.47102.83973.64111.08562.15193.40483.88253.41064.3310
C32.41751.39431.39862.42092.80043.75954.23284.78202.16701.08642.15863.40563.88495.5708
C42.79192.42051.39861.39802.42354.27865.04724.98683.41272.15811.08662.15703.40985.8931
C52.41622.79612.42091.39801.39583.79604.80984.14813.88153.40582.15761.08642.16085.1073
C61.40102.43042.80042.42351.39582.52713.64732.75243.40553.88673.40702.15201.08453.7131
C71.48752.47103.75954.27863.79602.52711.21131.35222.64104.62225.36524.67542.74811.8688
O82.39562.83974.23285.04724.80983.64731.21132.24832.51084.89226.11335.76313.95742.2740
O92.36453.64114.78204.98684.14812.75241.35222.24833.97045.74686.04844.79232.41290.9672
H102.14731.08562.16703.41273.88153.40552.64102.51083.97042.49874.31324.96794.29454.4626
H113.40212.15191.08642.15813.40583.88674.62224.89225.74682.49872.48684.30364.97136.4769
H123.87853.40482.15861.08662.15763.40705.36526.11336.04844.31322.48682.48454.30736.9658
H133.40073.88253.40562.15701.08642.15204.67545.76314.79234.96794.30362.48452.48725.7584
H142.15643.41063.88493.40982.16081.08452.74813.95742.41294.29454.97134.30732.48723.3555
H153.18034.33105.57085.89315.10733.71311.86882.27400.96724.46266.47696.96585.75843.3555

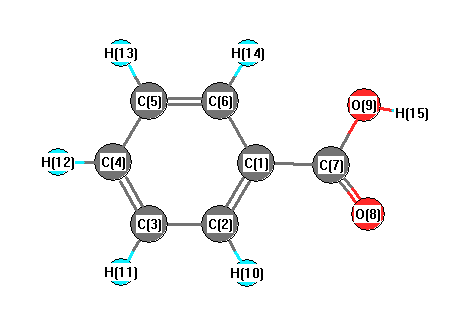
picture of benzoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.759 C1 C2 H10 118.928
C1 C6 C5 119.509 C1 C6 H14 119.818
C1 C7 O8 124.849 C1 C7 O9 112.668
C2 C1 C6 120.340 C2 C1 C7 117.621
C2 C3 C4 120.148 C2 C3 H11 119.816
C3 C2 H10 121.313 C3 C4 C5 119.915
C3 C4 H12 120.066 C4 C3 H11 120.036
C4 C5 C6 120.329 C4 C5 H13 119.982
C5 C4 H12 120.019 C5 C6 H14 120.673
C6 C1 C7 122.039 C6 C5 H13 119.689
C7 O9 H15 106.172 O8 C7 O9 122.484
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability