return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SOCH3 (Dimethyl sulfoxide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-552.381565
Energy at 298.15K-552.388562
HF Energy-551.589994
Nuclear repulsion energy183.993541
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3207 3037 4.13 99.06 0.73 0.85
2 A' 3200 3031 7.14 94.80 0.73 0.84
3 A' 3085 2922 7.35 218.46 0.00 0.00
4 A' 1516 1436 28.41 1.58 0.69 0.82
5 A' 1491 1412 5.04 18.29 0.71 0.83
6 A' 1401 1327 20.16 2.77 0.08 0.15
7 A' 1126 1066 147.80 4.27 0.52 0.68
8 A' 1063 1007 19.54 2.52 0.51 0.68
9 A' 999 946 12.67 3.75 0.71 0.83
10 A' 696 659 12.16 26.90 0.12 0.21
11 A' 379 359 9.51 2.17 0.46 0.63
12 A' 302 286 0.41 2.49 0.74 0.85
13 A' 242 229 0.62 0.12 0.56 0.72
14 A" 3205 3036 1.25 39.16 0.75 0.86
15 A" 3197 3028 0.02 7.73 0.75 0.86
16 A" 3083 2920 4.39 0.14 0.75 0.86
17 A" 1495 1416 0.22 23.04 0.75 0.86
18 A" 1475 1397 13.54 2.17 0.75 0.86
19 A" 1377 1304 9.37 0.31 0.75 0.86
20 A" 964 913 8.11 3.40 0.75 0.86
21 A" 929 880 2.14 0.50 0.75 0.86
22 A" 719 681 22.30 16.27 0.75 0.86
23 A" 316 300 10.31 4.07 0.75 0.86
24 A" 167 158 0.00 0.24 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 17816.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16874.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.23103 0.23099 0.14022

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.260 0.429 0.000
O2 -1.099 1.067 0.000
C3 0.260 -0.781 1.339
C4 0.260 -0.781 -1.339
H5 1.160 -1.398 1.305
H6 1.160 -1.398 -1.305
H7 0.232 -0.224 2.275
H8 0.232 -0.224 -2.275
H9 -0.641 -1.391 1.256
H10 -0.641 -1.391 -1.256

Atom - Atom Distances (Å)
  S1 O2 C3 C4 H5 H6 H7 H8 H9 H10
S11.50181.80561.80562.41922.41922.36722.36722.38812.3881
O21.50182.65682.65683.58943.58942.93482.93482.79762.7976
C31.80562.65682.67881.09122.86091.08973.65751.09132.8142
C41.80562.65682.67882.86091.09123.65751.08972.81421.0913
H52.41923.58941.09122.86092.61091.78363.88101.80193.1313
H62.41923.58942.86091.09122.61093.88101.78363.13131.8019
H72.36722.93481.08973.65751.78363.88104.55031.77833.8199
H82.36722.93483.65751.08973.88101.78364.55033.81991.7783
H92.38812.79761.09132.81421.80193.13131.77833.81992.5118
H102.38812.79762.81421.09133.13131.80193.81991.77832.5118

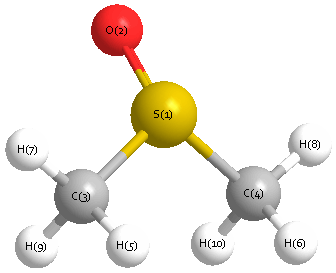
picture of Dimethyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 H5 110.836 S1 C3 H7 107.086
S1 C3 H9 108.525 S1 C4 H6 110.836
S1 C4 H8 107.086 S1 C4 H10 108.525
O2 S1 C3 106.532 O2 S1 C4 106.532
C3 S1 C4 95.774 H5 C3 H7 109.735
H5 C3 H9 111.305 H6 C4 H8 109.735
H6 C4 H10 111.305 H7 C3 H9 109.244
H8 C4 H10 109.244
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability