return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-247.910582
Energy at 298.15K-247.918906
HF Energy-247.043112
Nuclear repulsion energy181.624114
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3217 3047 0.52      
2 A' 3185 3017 16.10      
3 A' 3071 2908 49.79      
4 A' 3063 2901 52.10      
5 A' 3007 2848 96.87      
6 A' 1796 1701 423.13      
7 A' 1578 1495 21.70      
8 A' 1544 1462 16.57      
9 A' 1504 1424 7.25      
10 A' 1471 1393 25.34      
11 A' 1463 1385 44.62      
12 A' 1443 1366 44.45      
13 A' 1320 1250 36.03      
14 A' 1121 1061 115.58      
15 A' 1105 1046 3.60      
16 A' 897 849 2.38      
17 A' 668 632 6.72      
18 A' 395 374 1.25      
19 A' 325 308 10.61      
20 A" 3142 2976 16.69      
21 A" 3131 2965 51.35      
22 A" 1540 1458 19.78      
23 A" 1518 1437 4.07      
24 A" 1197 1134 2.84      
25 A" 1150 1089 0.12      
26 A" 1024 970 0.02      
27 A" 332 315 17.04      
28 A" 232 219 2.83      
29 A" 164 156 0.45      
30 A" 76 72 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 22837.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 21629.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.29973 0.13936 0.09868

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.698 -0.823 0.000
O2 0.213 -1.941 0.000
N3 0.000 0.348 0.000
C4 -1.448 0.332 0.000
C5 0.658 1.634 0.000
H6 1.790 -0.652 0.000
H7 -1.774 -0.706 0.000
H8 -1.837 0.836 0.889
H9 -1.837 0.836 -0.889
H10 1.739 1.493 0.000
H11 0.383 2.213 -0.888
H12 0.383 2.213 0.888

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.21861.36332.43662.45751.10562.47433.15773.15772.53953.17863.1786
O21.21862.29932.81513.60282.03672.33953.56513.56513.75804.25124.2512
N31.36332.29931.44781.44432.05052.06342.09892.09892.08212.10052.1005
C42.43662.81511.44782.47573.38391.08791.09421.09423.39192.77112.7711
C52.45753.60281.44432.47572.55123.37472.76642.76641.09061.09481.0948
H61.10562.03672.05053.38392.55123.56414.02044.02042.14573.31283.3128
H72.47432.33952.06341.08793.37473.56411.78151.78154.14453.73653.7365
H83.15773.56512.09891.09422.76644.02041.78151.77893.74363.15992.6127
H93.15773.56512.09891.09422.76644.02041.78151.77893.74362.61273.1599
H102.53953.75802.08213.39191.09062.14574.14453.74363.74361.77331.7733
H113.17864.25122.10052.77111.09483.31283.73653.15992.61271.77331.7757
H123.17864.25122.10052.77111.09483.31283.73652.61273.15991.77331.7757

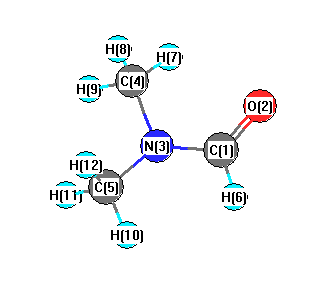
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 120.134 C1 N3 C5 122.128
O2 C1 N3 125.787 O2 C1 H6 122.329
N3 C1 H6 111.884 N3 C4 H7 108.087
N3 C4 H8 110.539 N3 C4 H9 110.539
N3 C5 H10 109.658 N3 C5 H11 110.885
N3 C5 H12 110.885 C4 N3 C5 117.738
H7 C4 H8 109.455 H7 C4 H9 109.455
H8 C4 H9 108.750 H10 C5 H11 108.477
H10 C5 H12 108.477 H11 C5 H12 108.388
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability