Jump to
S1C2
Energy calculated at CISD/6-311G*
| | hartrees |
|---|
| Energy at 0K | -229.488015 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 135.650522 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3248 |
3005 |
0.00 |
|
|
|
| 2 |
Ag |
3136 |
2902 |
0.00 |
|
|
|
| 3 |
Ag |
1600 |
1481 |
0.00 |
|
|
|
| 4 |
Ag |
1543 |
1428 |
0.00 |
|
|
|
| 5 |
Ag |
1350 |
1249 |
0.00 |
|
|
|
| 6 |
Ag |
1213 |
1122 |
0.00 |
|
|
|
| 7 |
Ag |
942 |
872 |
0.00 |
|
|
|
| 8 |
Ag |
525 |
485 |
0.00 |
|
|
|
| 9 |
Au |
3209 |
2969 |
97.91 |
|
|
|
| 10 |
Au |
1547 |
1431 |
12.88 |
|
|
|
| 11 |
Au |
1243 |
1150 |
3.59 |
|
|
|
| 12 |
Au |
223 |
207 |
3.48 |
|
|
|
| 13 |
Au |
21i |
20i |
12.82 |
|
|
|
| 14 |
Bg |
3209 |
2969 |
0.00 |
|
|
|
| 15 |
Bg |
1546 |
1431 |
0.00 |
|
|
|
| 16 |
Bg |
1248 |
1155 |
0.00 |
|
|
|
| 17 |
Bg |
282 |
261 |
0.00 |
|
|
|
| 18 |
Bu |
3248 |
3005 |
43.42 |
|
|
|
| 19 |
Bu |
3132 |
2898 |
89.61 |
|
|
|
| 20 |
Bu |
1594 |
1475 |
22.61 |
|
|
|
| 21 |
Bu |
1534 |
1419 |
6.83 |
|
|
|
| 22 |
Bu |
1249 |
1156 |
8.87 |
|
|
|
| 23 |
Bu |
1149 |
1063 |
145.19 |
|
|
|
| 24 |
Bu |
326 |
302 |
11.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19136.5 cm
-1
Scaled (by 0.9253) Zero Point Vibrational Energy (zpe) 17707.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CISD/6-311G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.427 |
0.564 |
0.000 |
| O2 |
0.427 |
-0.564 |
0.000 |
| C3 |
0.427 |
1.674 |
0.000 |
| C4 |
-0.427 |
-1.674 |
0.000 |
| H5 |
-0.229 |
2.538 |
0.000 |
| H6 |
1.053 |
1.679 |
0.891 |
| H7 |
1.053 |
1.679 |
-0.891 |
| H8 |
0.229 |
-2.538 |
0.000 |
| H9 |
-1.053 |
-1.679 |
0.891 |
| H10 |
-1.053 |
-1.679 |
-0.891 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 1.4152 | 1.4008 | 2.2375 | 1.9845 | 2.0564 | 2.0564 | 3.1709 | 2.4934 | 2.4934 |
O2 | 1.4152 | | 2.2375 | 1.4008 | 3.1709 | 2.4934 | 2.4934 | 1.9845 | 2.0564 | 2.0564 | C3 | 1.4008 | 2.2375 | | 3.4546 | 1.0855 | 1.0884 | 1.0884 | 4.2167 | 3.7719 | 3.7719 | C4 | 2.2375 | 1.4008 | 3.4546 | | 4.2167 | 3.7719 | 3.7719 | 1.0855 | 1.0884 | 1.0884 | H5 | 1.9845 | 3.1709 | 1.0855 | 4.2167 | | 1.7814 | 1.7814 | 5.0974 | 4.3890 | 4.3890 | H6 | 2.0564 | 2.4934 | 1.0884 | 3.7719 | 1.7814 | | 1.7817 | 4.3890 | 3.9642 | 4.3462 | H7 | 2.0564 | 2.4934 | 1.0884 | 3.7719 | 1.7814 | 1.7817 | | 4.3890 | 4.3462 | 3.9642 | H8 | 3.1709 | 1.9845 | 4.2167 | 1.0855 | 5.0974 | 4.3890 | 4.3890 | | 1.7814 | 1.7814 | H9 | 2.4934 | 2.0564 | 3.7719 | 1.0884 | 4.3890 | 3.9642 | 4.3462 | 1.7814 | | 1.7817 | H10 | 2.4934 | 2.0564 | 3.7719 | 1.0884 | 4.3890 | 4.3462 | 3.9642 | 1.7814 | 1.7817 | |
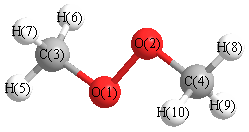 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
O2 |
C4 |
105.230 |
|
O1 |
C3 |
H5 |
105.208 |
| O1 |
C3 |
H6 |
110.786 |
|
O1 |
C3 |
H7 |
110.786 |
| O2 |
O1 |
C3 |
105.230 |
|
O2 |
C4 |
H8 |
105.208 |
| O2 |
C4 |
H9 |
110.786 |
|
O2 |
C4 |
H10 |
110.786 |
| H5 |
C3 |
H6 |
110.055 |
|
H5 |
C3 |
H7 |
110.055 |
| H6 |
C3 |
H7 |
109.866 |
|
H8 |
C4 |
H9 |
110.055 |
| H8 |
C4 |
H10 |
110.055 |
|
H9 |
C4 |
H10 |
109.866 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CISD/6-311G*
| | hartrees |
|---|
| Energy at 0K | -229.488174 |
| Energy at 298.15K | -229.495465 |
| Nuclear repulsion energy | 136.284747 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3243 |
3001 |
0.56 |
|
|
|
| 2 |
A |
3209 |
2969 |
72.30 |
|
|
|
| 3 |
A |
3134 |
2899 |
22.50 |
|
|
|
| 4 |
A |
1597 |
1477 |
0.31 |
|
|
|
| 5 |
A |
1551 |
1436 |
8.39 |
|
|
|
| 6 |
A |
1545 |
1430 |
0.82 |
|
|
|
| 7 |
A |
1321 |
1222 |
1.13 |
|
|
|
| 8 |
A |
1247 |
1154 |
1.53 |
|
|
|
| 9 |
A |
1196 |
1107 |
9.54 |
|
|
|
| 10 |
A |
938 |
868 |
5.02 |
|
|
|
| 11 |
A |
495 |
458 |
4.22 |
|
|
|
| 12 |
A |
244 |
225 |
1.57 |
|
|
|
| 13 |
A |
51 |
47 |
9.66 |
|
|
|
| 14 |
B |
3243 |
3001 |
45.57 |
|
|
|
| 15 |
B |
3207 |
2968 |
26.37 |
|
|
|
| 16 |
B |
3129 |
2895 |
58.09 |
|
|
|
| 17 |
B |
1590 |
1471 |
20.35 |
|
|
|
| 18 |
B |
1550 |
1434 |
4.11 |
|
|
|
| 19 |
B |
1535 |
1420 |
6.25 |
|
|
|
| 20 |
B |
1269 |
1174 |
4.09 |
|
|
|
| 21 |
B |
1242 |
1149 |
5.44 |
|
|
|
| 22 |
B |
1127 |
1043 |
100.12 |
|
|
|
| 23 |
B |
450 |
416 |
7.14 |
|
|
|
| 24 |
B |
251 |
233 |
3.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19180.7 cm
-1
Scaled (by 0.9253) Zero Point Vibrational Energy (zpe) 17747.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CISD/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.387 |
0.589 |
-0.367 |
| O2 |
-0.387 |
-0.589 |
-0.367 |
| C3 |
-0.387 |
1.570 |
0.274 |
| C4 |
0.387 |
-1.570 |
0.274 |
| H5 |
0.213 |
2.474 |
0.238 |
| H6 |
-1.325 |
1.725 |
-0.253 |
| H7 |
-0.586 |
1.297 |
1.310 |
| H8 |
-0.213 |
-2.474 |
0.238 |
| H9 |
1.325 |
-1.725 |
-0.253 |
| H10 |
0.586 |
-1.297 |
1.310 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 1.4090 | 1.4040 | 2.2517 | 1.9878 | 2.0582 | 2.0640 | 3.1795 | 2.5000 | 2.5317 |
O2 | 1.4090 | | 2.2517 | 1.4040 | 3.1795 | 2.5000 | 2.5317 | 1.9878 | 2.0582 | 2.0640 | C3 | 1.4040 | 2.2517 | | 3.2330 | 1.0860 | 1.0878 | 1.0892 | 4.0479 | 3.7505 | 3.1998 | C4 | 2.2517 | 1.4040 | 3.2330 | | 4.0479 | 3.7505 | 3.1998 | 1.0860 | 1.0878 | 1.0892 | H5 | 1.9878 | 3.1795 | 1.0860 | 4.0479 | | 1.7798 | 1.7812 | 4.9672 | 4.3724 | 3.9390 | H6 | 2.0582 | 2.5000 | 1.0878 | 3.7505 | 1.7798 | | 1.7809 | 4.3724 | 4.3515 | 3.9029 | H7 | 2.0640 | 2.5317 | 1.0892 | 3.1998 | 1.7812 | 1.7809 | | 3.9390 | 3.9029 | 2.8473 | H8 | 3.1795 | 1.9878 | 4.0479 | 1.0860 | 4.9672 | 4.3724 | 3.9390 | | 1.7798 | 1.7812 | H9 | 2.5000 | 2.0582 | 3.7505 | 1.0878 | 4.3724 | 4.3515 | 3.9029 | 1.7798 | | 1.7809 | H10 | 2.5317 | 2.0640 | 3.1998 | 1.0892 | 3.9390 | 3.9029 | 2.8473 | 1.7812 | 1.7809 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
O2 |
C4 |
106.352 |
|
O1 |
C3 |
H5 |
105.229 |
| O1 |
C3 |
H6 |
110.745 |
|
O1 |
C3 |
H7 |
111.134 |
| O2 |
O1 |
C3 |
106.352 |
|
O2 |
C4 |
H8 |
105.229 |
| O2 |
C4 |
H9 |
110.745 |
|
O2 |
C4 |
H10 |
111.134 |
| H5 |
C3 |
H6 |
109.920 |
|
H5 |
C3 |
H7 |
109.946 |
| H6 |
C3 |
H7 |
109.782 |
|
H8 |
C4 |
H9 |
109.920 |
| H8 |
C4 |
H10 |
109.946 |
|
H9 |
C4 |
H10 |
109.782 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
