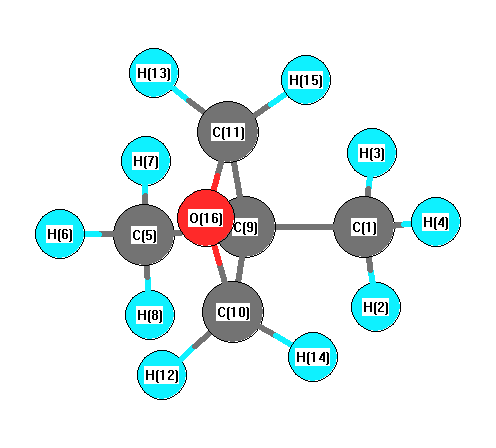
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3108 |
2992 |
38.42 |
|
|
|
| 2 |
A' |
3105 |
2989 |
36.36 |
|
|
|
| 3 |
A' |
3065 |
2951 |
91.96 |
|
|
|
| 4 |
A' |
3033 |
2920 |
24.15 |
|
|
|
| 5 |
A' |
3026 |
2913 |
28.01 |
|
|
|
| 6 |
A' |
3017 |
2904 |
22.22 |
|
|
|
| 7 |
A' |
1548 |
1491 |
1.72 |
|
|
|
| 8 |
A' |
1518 |
1461 |
5.74 |
|
|
|
| 9 |
A' |
1501 |
1445 |
4.74 |
|
|
|
| 10 |
A' |
1432 |
1378 |
4.70 |
|
|
|
| 11 |
A' |
1413 |
1361 |
11.83 |
|
|
|
| 12 |
A' |
1391 |
1340 |
0.20 |
|
|
|
| 13 |
A' |
1306 |
1257 |
0.54 |
|
|
|
| 14 |
A' |
1277 |
1229 |
9.12 |
|
|
|
| 15 |
A' |
1175 |
1131 |
0.95 |
|
|
|
| 16 |
A' |
1036 |
997 |
7.34 |
|
|
|
| 17 |
A' |
988 |
951 |
12.25 |
|
|
|
| 18 |
A' |
967 |
931 |
4.22 |
|
|
|
| 19 |
A' |
939 |
904 |
5.20 |
|
|
|
| 20 |
A' |
857 |
825 |
9.82 |
|
|
|
| 21 |
A' |
645 |
621 |
0.59 |
|
|
|
| 22 |
A' |
407 |
392 |
0.08 |
|
|
|
| 23 |
A' |
343 |
331 |
0.12 |
|
|
|
| 24 |
A' |
64 |
62 |
5.10 |
|
|
|
| 25 |
A" |
3099 |
2983 |
66.41 |
|
|
|
| 26 |
A" |
3094 |
2979 |
0.04 |
|
|
|
| 27 |
A" |
3064 |
2950 |
11.18 |
|
|
|
| 28 |
A" |
3007 |
2895 |
143.30 |
|
|
|
| 29 |
A" |
1531 |
1474 |
8.77 |
|
|
|
| 30 |
A" |
1515 |
1459 |
5.27 |
|
|
|
| 31 |
A" |
1500 |
1444 |
0.04 |
|
|
|
| 32 |
A" |
1315 |
1266 |
2.21 |
|
|
|
| 33 |
A" |
1228 |
1183 |
0.14 |
|
|
|
| 34 |
A" |
1171 |
1127 |
0.10 |
|
|
|
| 35 |
A" |
1064 |
1024 |
0.10 |
|
|
|
| 36 |
A" |
1056 |
1017 |
85.76 |
|
|
|
| 37 |
A" |
943 |
908 |
0.01 |
|
|
|
| 38 |
A" |
893 |
860 |
3.88 |
|
|
|
| 39 |
A" |
392 |
377 |
0.72 |
|
|
|
| 40 |
A" |
315 |
304 |
0.13 |
|
|
|
| 41 |
A" |
279 |
269 |
0.03 |
|
|
|
| 42 |
A" |
229 |
220 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31428.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 30256.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.636 |
|
|
|
| 2 |
H |
0.223 |
|
|
|
| 3 |
H |
0.223 |
|
|
|
| 4 |
H |
0.222 |
|
|
|
| 5 |
C |
-0.640 |
|
|
|
| 6 |
H |
0.230 |
|
|
|
| 7 |
H |
0.222 |
|
|
|
| 8 |
H |
0.222 |
|
|
|
| 9 |
C |
-0.237 |
|
|
|
| 10 |
C |
-0.139 |
|
|
|
| 11 |
C |
-0.139 |
|
|
|
| 12 |
H |
0.201 |
|
|
|
| 13 |
H |
0.201 |
|
|
|
| 14 |
H |
0.195 |
|
|
|
| 15 |
H |
0.195 |
|
|
|
| 16 |
O |
-0.344 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.072 |
2.009 |
0.000 |
2.010 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.736 |
0.053 |
0.000 |
| y |
0.053 |
-43.686 |
0.000 |
| z |
0.000 |
0.000 |
-36.021 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.118 |
0.053 |
0.000 |
| y |
0.053 |
-6.808 |
0.000 |
| z |
0.000 |
0.000 |
4.690 |
|
| Polar |
|---|
| 3z2-r2 | 9.380 |
|---|
| x2-y2 | 5.950 |
|---|
| xy | 0.053 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
158.555 |
| (<r2>)1/2 |
12.592 |