return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Ethenylcyclopropane)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-194.743114
Energy at 298.15K-194.751305
HF Energy-193.972825
Nuclear repulsion energy162.971026
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3284 3111 17.13      
2 A' 3282 3109 11.82      
3 A' 3198 3029 8.45      
4 A' 3186 3017 2.39      
5 A' 3182 3014 10.23      
6 A' 3166 2998 9.22      
7 A' 1706 1616 20.13      
8 A' 1542 1460 2.28      
9 A' 1491 1412 0.35      
10 A' 1377 1304 0.44      
11 A' 1332 1261 3.05      
12 A' 1258 1192 3.07      
13 A' 1245 1179 2.11      
14 A' 1076 1019 5.70      
15 A' 1025 970 5.17      
16 A' 989 937 38.65      
17 A' 844 800 1.36      
18 A' 788 746 0.30      
19 A' 450 427 1.18      
20 A' 271 256 1.05      
21 A" 3271 3098 0.08      
22 A" 3177 3009 15.17      
23 A" 1501 1421 3.76      
24 A" 1221 1156 0.85      
25 A" 1143 1083 3.59      
26 A" 1080 1023 6.57      
27 A" 1010 957 14.54      
28 A" 938 888 8.92      
29 A" 880 833 50.65      
30 A" 852 807 5.58      
31 A" 678 643 6.76      
32 A" 317 300 1.66      
33 A" 120 113 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 25438.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24093.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.50537 0.10191 0.09804

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.392 0.287 0.000
C2 0.257 -1.036 0.000
C3 -0.398 -2.206 0.000
C4 0.257 1.432 0.751
C5 0.257 1.432 -0.751
H6 -1.479 0.275 0.000
H7 1.348 -1.037 0.000
H8 0.133 -3.150 0.000
H9 -1.483 -2.247 0.000
H10 -0.394 2.129 1.265
H11 1.192 1.223 1.259
H12 -0.394 2.129 -1.265
H13 1.192 1.223 -1.259

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.47392.49271.51511.51511.08692.18713.47752.75862.23412.22912.23412.2291
C21.47391.34052.57952.57952.17531.09112.11802.11973.46972.74953.46972.7495
C32.49271.34053.77143.77142.70562.10091.08401.08574.51493.98294.51493.9829
C41.51512.57953.77141.50142.21742.80194.64504.13801.08371.08432.23012.2264
C51.51512.57953.77141.50142.21742.80194.64504.13802.23012.22641.08371.0843
H61.08692.17532.70562.21742.21743.11683.78562.52142.49273.10102.49273.1010
H72.18711.09112.10092.80192.80193.11682.43743.07863.82872.59173.82872.5917
H83.47752.11801.08404.64504.64503.78562.43741.85175.45404.67205.45404.6720
H92.75862.11971.08574.13804.13802.52143.07861.85174.68274.55784.68274.5578
H102.23413.46974.51491.08372.23012.49273.82875.45404.68271.82652.53023.1158
H112.22912.74953.98291.08432.22643.10102.59174.67204.55781.82653.11582.5185
H122.23413.46974.51492.23011.08372.49273.82875.45404.68272.53023.11581.8265
H132.22912.74953.98292.22641.08433.10102.59174.67204.55783.11582.51851.8265

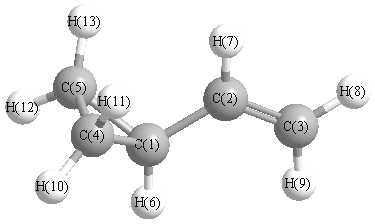
picture of Ethenylcyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.608 C1 C2 H7 116.205
C1 C4 C5 60.298 C1 C4 H10 117.595
C1 C4 H11 117.122 C1 C5 C4 60.298
C1 C5 H12 117.595 C1 C5 H13 117.122
C2 C1 C4 119.304 C2 C1 C5 119.304
C2 C1 H6 115.483 C2 C3 H8 121.398
C2 C3 H9 121.424 C3 C2 H7 119.186
C4 C1 H6 115.926 C4 C5 H12 118.338
C4 C5 H13 117.971 C5 C1 H6 115.926
C5 C4 H10 118.338 C5 C4 H11 117.971
H8 C3 H9 117.178 H10 C4 H11 114.810
H12 C5 H13 114.810
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability