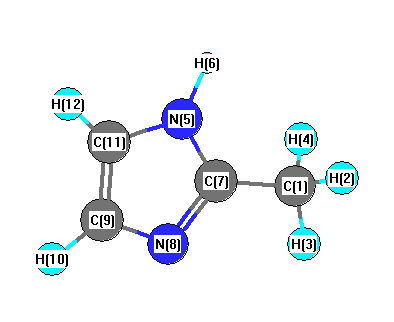
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3935 |
3559 |
63.60 |
|
|
|
| 2 |
A |
3444 |
3115 |
10.04 |
|
|
|
| 3 |
A |
3413 |
3087 |
9.83 |
|
|
|
| 4 |
A |
3300 |
2985 |
11.73 |
|
|
|
| 5 |
A |
3230 |
2922 |
36.41 |
|
|
|
| 6 |
A |
3185 |
2880 |
38.80 |
|
|
|
| 7 |
A |
1774 |
1605 |
65.11 |
|
|
|
| 8 |
A |
1667 |
1508 |
9.32 |
|
|
|
| 9 |
A |
1638 |
1482 |
8.78 |
|
|
|
| 10 |
A |
1618 |
1463 |
7.73 |
|
|
|
| 11 |
A |
1587 |
1435 |
49.13 |
|
|
|
| 12 |
A |
1553 |
1405 |
0.60 |
|
|
|
| 13 |
A |
1507 |
1363 |
2.23 |
|
|
|
| 14 |
A |
1380 |
1248 |
14.93 |
|
|
|
| 15 |
A |
1251 |
1132 |
15.53 |
|
|
|
| 16 |
A |
1233 |
1115 |
0.31 |
|
|
|
| 17 |
A |
1175 |
1063 |
31.32 |
|
|
|
| 18 |
A |
1168 |
1056 |
0.92 |
|
|
|
| 19 |
A |
1099 |
994 |
11.96 |
|
|
|
| 20 |
A |
1054 |
953 |
6.89 |
|
|
|
| 21 |
A |
1005 |
909 |
2.08 |
|
|
|
| 22 |
A |
980 |
886 |
7.26 |
|
|
|
| 23 |
A |
824 |
745 |
57.04 |
|
|
|
| 24 |
A |
757 |
684 |
11.38 |
|
|
|
| 25 |
A |
720 |
651 |
1.59 |
|
|
|
| 26 |
A |
693 |
626 |
9.11 |
|
|
|
| 27 |
A |
536 |
485 |
103.79 |
|
|
|
| 28 |
A |
373 |
337 |
5.59 |
|
|
|
| 29 |
A |
280 |
253 |
9.54 |
|
|
|
| 30 |
A |
90 |
82 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23234.0 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 21012.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.689 |
-0.300 |
|
-0.515 |
| 2 |
H |
0.225 |
0.070 |
|
0.124 |
| 3 |
H |
0.258 |
0.110 |
|
0.165 |
| 4 |
H |
0.224 |
0.070 |
|
0.124 |
| 5 |
N |
-0.649 |
-0.294 |
|
-0.338 |
| 6 |
H |
0.380 |
0.316 |
|
0.339 |
| 7 |
C |
0.483 |
0.544 |
|
0.655 |
| 8 |
N |
-0.424 |
-0.622 |
|
-0.628 |
| 9 |
C |
-0.173 |
0.157 |
|
0.091 |
| 10 |
H |
0.213 |
0.094 |
|
0.138 |
| 11 |
C |
-0.082 |
-0.354 |
|
-0.405 |
| 12 |
H |
0.234 |
0.208 |
|
0.249 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.570 |
3.704 |
-0.002 |
3.748 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.532 |
3.670 |
-0.001 |
3.709 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.579 |
0.630 |
-0.001 |
| y |
0.630 |
-34.252 |
-0.004 |
| z |
-0.001 |
-0.004 |
-38.382 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.739 |
0.630 |
-0.001 |
| y |
0.630 |
0.229 |
-0.004 |
| z |
-0.001 |
-0.004 |
-5.967 |
|
| Polar |
|---|
| 3z2-r2 | -11.934 |
|---|
| x2-y2 | 3.673 |
|---|
| xy | 0.630 |
|---|
| xz | -0.001 |
|---|
| yz | -0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.121 |
0.131 |
0.000 |
| y |
0.131 |
7.872 |
-0.000 |
| z |
0.000 |
-0.000 |
4.602 |
<r2> (average value of r
2) Å
2
| <r2> |
137.247 |
| (<r2>)1/2 |
11.715 |