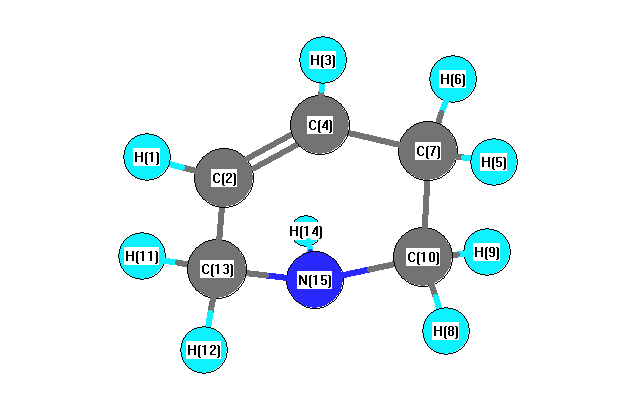
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3759 |
3400 |
0.46 |
|
|
|
| 2 |
A |
3324 |
3006 |
53.04 |
|
|
|
| 3 |
A |
3296 |
2981 |
21.39 |
|
|
|
| 4 |
A |
3233 |
2924 |
72.64 |
|
|
|
| 5 |
A |
3212 |
2905 |
50.73 |
|
|
|
| 6 |
A |
3200 |
2895 |
102.19 |
|
|
|
| 7 |
A |
3182 |
2878 |
25.06 |
|
|
|
| 8 |
A |
3168 |
2865 |
63.93 |
|
|
|
| 9 |
A |
3151 |
2850 |
52.38 |
|
|
|
| 10 |
A |
1865 |
1687 |
1.68 |
|
|
|
| 11 |
A |
1650 |
1492 |
10.88 |
|
|
|
| 12 |
A |
1635 |
1479 |
1.54 |
|
|
|
| 13 |
A |
1623 |
1468 |
3.12 |
|
|
|
| 14 |
A |
1616 |
1461 |
2.50 |
|
|
|
| 15 |
A |
1543 |
1395 |
0.57 |
|
|
|
| 16 |
A |
1534 |
1388 |
13.49 |
|
|
|
| 17 |
A |
1503 |
1359 |
4.94 |
|
|
|
| 18 |
A |
1479 |
1338 |
0.27 |
|
|
|
| 19 |
A |
1450 |
1312 |
1.17 |
|
|
|
| 20 |
A |
1373 |
1242 |
5.36 |
|
|
|
| 21 |
A |
1315 |
1189 |
17.33 |
|
|
|
| 22 |
A |
1303 |
1178 |
3.89 |
|
|
|
| 23 |
A |
1230 |
1113 |
20.82 |
|
|
|
| 24 |
A |
1216 |
1100 |
3.41 |
|
|
|
| 25 |
A |
1121 |
1014 |
2.44 |
|
|
|
| 26 |
A |
1111 |
1005 |
0.72 |
|
|
|
| 27 |
A |
1092 |
987 |
12.63 |
|
|
|
| 28 |
A |
1067 |
965 |
13.36 |
|
|
|
| 29 |
A |
977 |
884 |
16.06 |
|
|
|
| 30 |
A |
952 |
861 |
3.88 |
|
|
|
| 31 |
A |
916 |
829 |
48.61 |
|
|
|
| 32 |
A |
878 |
794 |
63.56 |
|
|
|
| 33 |
A |
823 |
745 |
40.15 |
|
|
|
| 34 |
A |
718 |
649 |
23.15 |
|
|
|
| 35 |
A |
565 |
511 |
0.99 |
|
|
|
| 36 |
A |
520 |
470 |
0.70 |
|
|
|
| 37 |
A |
433 |
391 |
1.07 |
|
|
|
| 38 |
A |
313 |
283 |
8.85 |
|
|
|
| 39 |
A |
186 |
168 |
1.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31765.5 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 28728.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.206 |
0.109 |
|
0.131 |
| 2 |
C |
-0.242 |
-0.333 |
|
-0.326 |
| 3 |
H |
0.209 |
0.116 |
|
0.131 |
| 4 |
C |
-0.203 |
-0.180 |
|
-0.212 |
| 5 |
H |
0.213 |
-0.046 |
|
-0.032 |
| 6 |
H |
0.203 |
-0.019 |
|
0.000 |
| 7 |
C |
-0.430 |
0.129 |
|
0.133 |
| 8 |
H |
0.214 |
-0.002 |
|
0.055 |
| 9 |
H |
0.216 |
-0.034 |
|
-0.000 |
| 10 |
C |
-0.277 |
0.339 |
|
0.196 |
| 11 |
H |
0.212 |
-0.064 |
|
-0.013 |
| 12 |
H |
0.222 |
-0.042 |
|
0.016 |
| 13 |
C |
-0.251 |
0.583 |
|
0.409 |
| 14 |
H |
0.311 |
0.382 |
|
0.406 |
| 15 |
N |
-0.603 |
-0.936 |
|
-0.894 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.519 |
0.850 |
-0.512 |
1.120 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.493 |
0.801 |
-0.503 |
1.066 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.894 |
-2.050 |
0.234 |
| y |
-2.050 |
-40.731 |
0.958 |
| z |
0.234 |
0.958 |
-37.280 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.112 |
-2.050 |
0.234 |
| y |
-2.050 |
-4.144 |
0.958 |
| z |
0.234 |
0.958 |
1.032 |
|
| Polar |
|---|
| 3z2-r2 | 2.064 |
|---|
| x2-y2 | 4.838 |
|---|
| xy | -2.050 |
|---|
| xz | 0.234 |
|---|
| yz | 0.958 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.303 |
-0.015 |
-0.237 |
| y |
-0.015 |
7.968 |
0.037 |
| z |
-0.237 |
0.037 |
6.544 |
<r2> (average value of r
2) Å
2
| <r2> |
146.296 |
| (<r2>)1/2 |
12.095 |