return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10OS (Diethyl sulfoxide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-630.784971
Energy at 298.15K-630.796602
HF Energy-629.673359
Nuclear repulsion energy332.494865
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3191 3022 10.60      
2 A 3187 3018 15.77      
3 A 3181 3012 19.86      
4 A 3173 3005 10.19      
5 A 3165 2998 4.27      
6 A 3161 2994 1.51      
7 A 3108 2944 4.99      
8 A 3104 2940 11.60      
9 A 3092 2928 16.68      
10 A 3087 2924 14.51      
11 A 1548 1466 8.43      
12 A 1546 1464 8.53      
13 A 1541 1460 6.06      
14 A 1536 1455 14.45      
15 A 1510 1431 14.01      
16 A 1500 1421 9.04      
17 A 1452 1375 8.28      
18 A 1445 1369 11.49      
19 A 1353 1282 2.09      
20 A 1328 1258 1.53      
21 A 1295 1226 1.50      
22 A 1280 1213 0.38      
23 A 1119 1060 11.37      
24 A 1104 1045 91.65      
25 A 1094 1036 11.12      
26 A 1087 1030 35.41      
27 A 1054 999 5.74      
28 A 1015 962 3.26      
29 A 999 946 8.11      
30 A 810 768 10.68      
31 A 775 734 12.23      
32 A 716 678 22.65      
33 A 637 604 9.76      
34 A 479 453 6.21      
35 A 378 358 1.29      
36 A 324 307 3.63      
37 A 301 285 2.01      
38 A 243 231 1.18      
39 A 230 217 7.55      
40 A 204 193 4.09      
41 A 131 124 2.48      
42 A 77 73 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 30780.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29152.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.11261 0.08380 0.06011

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.372 1.497 0.559
H2 2.033 2.337 0.339
H3 0.551 1.870 1.172
H4 1.931 0.759 1.138
C5 -2.240 0.639 -0.084
H6 -2.521 0.271 -1.074
H7 -3.153 0.716 0.509
H8 -1.835 1.646 -0.196
C9 -1.253 -0.299 0.596
H10 -1.724 -1.255 0.843
H11 -0.837 0.104 1.522
C12 0.878 0.858 -0.728
H13 0.161 1.473 -1.277
S14 0.145 -0.787 -0.454
O15 1.101 -1.519 0.453
H16 1.716 0.649 -1.399

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 H13 S14 O15 H16
C11.09171.09111.09153.76744.39614.59253.29773.18114.15162.78401.51902.19962.78273.03032.1610
H21.09171.76381.77174.61755.19725.43683.96564.22135.22223.82492.15892.62043.73513.96942.4439
H31.09111.76381.77213.29814.12753.93562.75872.87953.87882.27352.17752.51143.14083.50853.0757
H41.09151.77171.77214.34724.99475.12294.09173.39864.18292.87052.14413.07772.84742.51972.5480
C53.76744.61753.29814.34721.09311.09191.09071.52172.17072.19803.19132.80712.80334.01364.1686
H64.39615.19724.12754.99471.09311.76171.76982.17262.57613.09853.46712.94582.93494.31934.2664
H74.59255.43683.93565.12291.09191.76171.76062.15622.45782.60084.21953.83983.75074.80655.2303
H83.29773.96562.75874.09171.09071.76981.76062.17893.08292.51462.87472.27593.14704.36563.8793
C93.18114.22132.87953.39861.52172.17262.15622.17891.09331.09222.76312.94041.81532.65573.7006
H104.15165.22223.87884.18292.17072.57612.45783.08291.09331.75863.70143.93472.32222.86384.5254
H112.78403.82492.27352.87052.19803.09852.60082.51461.09221.75862.92823.27172.37942.74533.9178
C121.51902.15892.17752.14413.19133.46714.21952.87472.76313.70142.92821.09291.82122.66371.0936
H132.19962.62042.51143.07772.80712.94583.83982.27592.94043.93473.27171.09292.40473.58181.7644
S142.78273.73513.14082.84742.80332.93493.75073.14701.81532.32222.37941.82122.40471.50722.3282
O153.03033.96943.50852.51974.01364.31934.80654.36562.65572.86382.74532.66373.58181.50722.9168
H162.16102.44393.07572.54804.16864.26645.23033.87933.70064.52543.91781.09361.76442.32822.9168

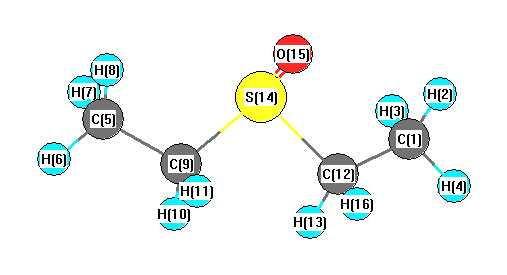
picture of Diethyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 H13 113.732 C1 C12 S14 112.522
C1 C12 H16 110.561 H2 C1 H3 107.808
H2 C1 H4 108.484 H2 C1 C12 110.507
H3 C1 H4 108.569 H3 C1 C12 112.035
H4 C1 C12 109.348 C5 C9 H10 111.158
C5 C9 H11 113.448 C5 C9 S14 114.007
H6 C5 H7 107.460 H6 C5 H8 108.271
H6 C5 C9 111.321 H7 C5 H8 107.538
H7 C5 C9 110.089 H8 C5 C9 111.979
C9 S14 C12 98.898 C9 S14 O15 105.754
H10 C9 H11 107.153 H10 C9 S14 103.136
H11 C9 S14 107.215 C12 S14 O15 105.931
H13 C12 S14 108.623 H13 C12 H16 107.596
S14 C12 H16 103.176
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability