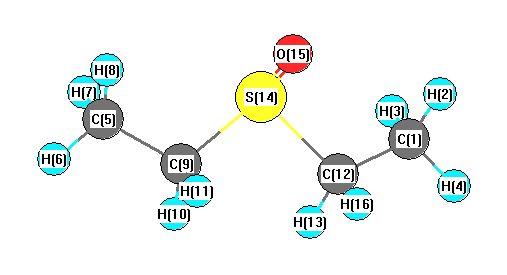
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3156 |
3012 |
18.19 |
|
|
|
| 2 |
A |
3151 |
3007 |
16.84 |
|
|
|
| 3 |
A |
3143 |
2999 |
23.54 |
|
|
|
| 4 |
A |
3137 |
2994 |
24.41 |
|
|
|
| 5 |
A |
3132 |
2989 |
5.10 |
|
|
|
| 6 |
A |
3128 |
2985 |
3.60 |
|
|
|
| 7 |
A |
3091 |
2950 |
6.08 |
|
|
|
| 8 |
A |
3086 |
2945 |
15.73 |
|
|
|
| 9 |
A |
3068 |
2928 |
23.15 |
|
|
|
| 10 |
A |
3063 |
2924 |
20.20 |
|
|
|
| 11 |
A |
1528 |
1458 |
6.38 |
|
|
|
| 12 |
A |
1523 |
1454 |
11.29 |
|
|
|
| 13 |
A |
1520 |
1451 |
4.10 |
|
|
|
| 14 |
A |
1517 |
1448 |
16.27 |
|
|
|
| 15 |
A |
1489 |
1421 |
11.27 |
|
|
|
| 16 |
A |
1478 |
1411 |
7.55 |
|
|
|
| 17 |
A |
1430 |
1364 |
9.92 |
|
|
|
| 18 |
A |
1423 |
1358 |
8.33 |
|
|
|
| 19 |
A |
1331 |
1270 |
1.02 |
|
|
|
| 20 |
A |
1304 |
1244 |
2.40 |
|
|
|
| 21 |
A |
1276 |
1218 |
1.37 |
|
|
|
| 22 |
A |
1266 |
1209 |
0.64 |
|
|
|
| 23 |
A |
1102 |
1052 |
10.87 |
|
|
|
| 24 |
A |
1088 |
1038 |
34.40 |
|
|
|
| 25 |
A |
1078 |
1029 |
35.20 |
|
|
|
| 26 |
A |
1071 |
1022 |
59.72 |
|
|
|
| 27 |
A |
1034 |
987 |
4.49 |
|
|
|
| 28 |
A |
1000 |
955 |
1.96 |
|
|
|
| 29 |
A |
987 |
942 |
11.02 |
|
|
|
| 30 |
A |
799 |
763 |
10.98 |
|
|
|
| 31 |
A |
766 |
731 |
10.72 |
|
|
|
| 32 |
A |
685 |
654 |
20.50 |
|
|
|
| 33 |
A |
605 |
577 |
6.85 |
|
|
|
| 34 |
A |
473 |
451 |
5.68 |
|
|
|
| 35 |
A |
368 |
351 |
1.81 |
|
|
|
| 36 |
A |
310 |
296 |
4.02 |
|
|
|
| 37 |
A |
276 |
263 |
1.66 |
|
|
|
| 38 |
A |
217 |
208 |
6.30 |
|
|
|
| 39 |
A |
212 |
203 |
2.55 |
|
|
|
| 40 |
A |
197 |
188 |
4.17 |
|
|
|
| 41 |
A |
122 |
116 |
2.07 |
|
|
|
| 42 |
A |
55 |
52 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30341.2 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 28957.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.671 |
|
|
|
| 2 |
H |
0.237 |
|
|
|
| 3 |
H |
0.225 |
|
|
|
| 4 |
H |
0.270 |
|
|
|
| 5 |
C |
-0.678 |
|
|
|
| 6 |
H |
0.242 |
|
|
|
| 7 |
H |
0.245 |
|
|
|
| 8 |
H |
0.234 |
|
|
|
| 9 |
C |
-0.639 |
|
|
|
| 10 |
H |
0.273 |
|
|
|
| 11 |
H |
0.271 |
|
|
|
| 12 |
C |
-0.648 |
|
|
|
| 13 |
H |
0.256 |
|
|
|
| 14 |
S |
0.703 |
|
|
|
| 15 |
O |
-0.590 |
|
|
|
| 16 |
H |
0.272 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.289 |
3.514 |
-0.917 |
4.293 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.939 |
4.770 |
-2.714 |
| y |
4.770 |
-50.264 |
1.020 |
| z |
-2.714 |
1.020 |
-45.405 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.895 |
4.770 |
-2.714 |
| y |
4.770 |
-5.592 |
1.020 |
| z |
-2.714 |
1.020 |
1.697 |
|
| Polar |
|---|
| 3z2-r2 | 3.394 |
|---|
| x2-y2 | 6.325 |
|---|
| xy | 4.770 |
|---|
| xz | -2.714 |
|---|
| yz | 1.020 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
222.052 |
| (<r2>)1/2 |
14.901 |