Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-311G*
Geometric Data calculated at PBE1PBE/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -194.158830 |
| Energy at 298.15K | |
| HF Energy | -194.158830 |
| Nuclear repulsion energy | 133.185620 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3862 |
3702 |
14.14 |
110.71 |
0.32 |
0.48 |
| 2 |
A |
3148 |
3018 |
28.19 |
41.04 |
0.75 |
0.86 |
| 3 |
A |
3121 |
2992 |
56.01 |
59.09 |
0.59 |
0.74 |
| 4 |
A |
3092 |
2964 |
22.97 |
96.49 |
0.72 |
0.84 |
| 5 |
A |
3056 |
2929 |
43.59 |
149.38 |
0.05 |
0.10 |
| 6 |
A |
3052 |
2925 |
39.57 |
92.75 |
0.06 |
0.11 |
| 7 |
A |
3018 |
2893 |
58.50 |
97.77 |
0.75 |
0.86 |
| 8 |
A |
2986 |
2862 |
75.08 |
113.83 |
0.12 |
0.22 |
| 9 |
A |
1538 |
1474 |
3.15 |
5.29 |
0.68 |
0.81 |
| 10 |
A |
1525 |
1462 |
9.72 |
6.89 |
0.75 |
0.86 |
| 11 |
A |
1509 |
1447 |
9.93 |
15.57 |
0.75 |
0.86 |
| 12 |
A |
1487 |
1425 |
4.17 |
12.85 |
0.72 |
0.84 |
| 13 |
A |
1474 |
1413 |
1.96 |
2.03 |
0.20 |
0.34 |
| 14 |
A |
1423 |
1364 |
6.51 |
1.69 |
0.67 |
0.81 |
| 15 |
A |
1387 |
1329 |
0.71 |
1.42 |
0.71 |
0.83 |
| 16 |
A |
1344 |
1289 |
29.60 |
12.25 |
0.74 |
0.85 |
| 17 |
A |
1286 |
1232 |
1.53 |
7.79 |
0.75 |
0.85 |
| 18 |
A |
1264 |
1211 |
44.81 |
3.61 |
0.53 |
0.69 |
| 19 |
A |
1172 |
1123 |
7.97 |
0.56 |
0.20 |
0.33 |
| 20 |
A |
1143 |
1096 |
18.08 |
4.23 |
0.60 |
0.75 |
| 21 |
A |
1091 |
1046 |
39.01 |
2.64 |
0.71 |
0.83 |
| 22 |
A |
1007 |
965 |
41.88 |
3.87 |
0.75 |
0.86 |
| 23 |
A |
933 |
894 |
2.77 |
0.18 |
0.67 |
0.80 |
| 24 |
A |
887 |
850 |
2.73 |
8.59 |
0.17 |
0.30 |
| 25 |
A |
778 |
746 |
0.70 |
0.49 |
0.69 |
0.82 |
| 26 |
A |
482 |
462 |
9.39 |
0.22 |
0.44 |
0.61 |
| 27 |
A |
334 |
320 |
19.26 |
0.58 |
0.58 |
0.73 |
| 28 |
A |
276 |
265 |
123.40 |
3.64 |
0.73 |
0.84 |
| 29 |
A |
229 |
219 |
2.36 |
0.06 |
0.48 |
0.65 |
| 30 |
A |
152 |
146 |
6.53 |
0.15 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 24027.1 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 23030.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.528 |
-0.513 |
0.127 |
| C2 |
-0.630 |
0.642 |
-0.285 |
| C3 |
0.766 |
0.541 |
0.290 |
| O4 |
1.368 |
-0.634 |
-0.218 |
| H5 |
-2.518 |
-0.433 |
-0.332 |
| H6 |
-1.668 |
-0.539 |
1.213 |
| H7 |
-1.090 |
-1.467 |
-0.174 |
| H8 |
-0.546 |
0.682 |
-1.377 |
| H9 |
-1.065 |
1.598 |
0.032 |
| H10 |
0.713 |
0.512 |
1.390 |
| H11 |
1.349 |
1.432 |
0.010 |
| H12 |
2.232 |
-0.735 |
0.187 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5208 | 2.5307 | 2.9195 | 1.0934 | 1.0953 | 1.0916 | 2.1577 | 2.1636 | 2.7693 | 3.4753 | 3.7673 |
C2 | 1.5208 | | 1.5132 | 2.3717 | 2.1733 | 2.1721 | 2.1615 | 1.0960 | 1.0965 | 2.1505 | 2.1508 | 3.2106 | C3 | 2.5307 | 1.5132 | | 1.4142 | 3.4815 | 2.8191 | 2.7736 | 2.1263 | 2.1301 | 1.1021 | 1.1006 | 1.9456 | O4 | 2.9195 | 2.3717 | 1.4142 | | 3.8927 | 3.3580 | 2.5958 | 2.5965 | 3.3110 | 2.0802 | 2.0782 | 0.9590 | H5 | 1.0934 | 2.1733 | 3.4815 | 3.8927 | | 1.7661 | 1.7694 | 2.4945 | 2.5234 | 3.7806 | 4.3066 | 4.7873 | H6 | 1.0953 | 2.1721 | 2.8191 | 3.3580 | 1.7661 | | 1.7662 | 3.0755 | 2.5152 | 2.6086 | 3.7997 | 4.0377 | H7 | 1.0916 | 2.1615 | 2.7736 | 2.5958 | 1.7694 | 1.7662 | | 2.5219 | 3.0714 | 3.1003 | 3.7926 | 3.4208 | H8 | 2.1577 | 1.0960 | 2.1263 | 2.5965 | 2.4945 | 3.0755 | 2.5219 | | 1.7581 | 3.0448 | 2.4651 | 3.4888 | H9 | 2.1636 | 1.0965 | 2.1301 | 3.3110 | 2.5234 | 2.5152 | 3.0714 | 1.7581 | | 2.4869 | 2.4198 | 4.0415 | H10 | 2.7693 | 2.1505 | 1.1021 | 2.0802 | 3.7806 | 2.6086 | 3.1003 | 3.0448 | 2.4869 | | 1.7769 | 2.3043 | H11 | 3.4753 | 2.1508 | 1.1006 | 2.0782 | 4.3066 | 3.7997 | 3.7926 | 2.4651 | 2.4198 | 1.7769 | | 2.3461 | H12 | 3.7673 | 3.2106 | 1.9456 | 0.9590 | 4.7873 | 4.0377 | 3.4208 | 3.4888 | 4.0415 | 2.3043 | 2.3461 | |
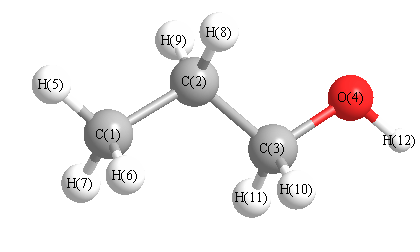 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.047 |
|
C1 |
C2 |
H8 |
110.028 |
| C1 |
C2 |
H9 |
110.459 |
|
C2 |
C1 |
H5 |
111.424 |
| C2 |
C1 |
H6 |
111.211 |
|
C2 |
C1 |
H7 |
110.587 |
| C2 |
C3 |
O4 |
108.182 |
|
C2 |
C3 |
H10 |
109.622 |
| C2 |
C3 |
H11 |
109.731 |
|
C3 |
C2 |
H8 |
108.091 |
| C3 |
C2 |
H9 |
108.360 |
|
C3 |
O4 |
H12 |
108.615 |
| O4 |
C3 |
H10 |
110.919 |
|
O4 |
C3 |
H11 |
110.841 |
| H5 |
C1 |
H6 |
107.590 |
|
H5 |
C1 |
H7 |
108.150 |
| H6 |
C1 |
H7 |
107.726 |
|
H8 |
C2 |
H9 |
106.619 |
| H10 |
C3 |
H11 |
107.541 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.662 |
|
|
|
| 2 |
C |
-0.466 |
|
|
|
| 3 |
C |
-0.201 |
|
|
|
| 4 |
O |
-0.575 |
|
|
|
| 5 |
H |
0.222 |
|
|
|
| 6 |
H |
0.212 |
|
|
|
| 7 |
H |
0.244 |
|
|
|
| 8 |
H |
0.228 |
|
|
|
| 9 |
H |
0.214 |
|
|
|
| 10 |
H |
0.190 |
|
|
|
| 11 |
H |
0.199 |
|
|
|
| 12 |
H |
0.395 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.893 |
0.891 |
1.051 |
1.642 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.565 |
-0.371 |
2.074 |
| y |
-0.371 |
-27.212 |
-0.622 |
| z |
2.074 |
-0.622 |
-26.894 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.488 |
-0.371 |
2.074 |
| y |
-0.371 |
-2.482 |
-0.622 |
| z |
2.074 |
-0.622 |
-2.005 |
|
| Polar |
|---|
| 3z2-r2 | -4.010 |
|---|
| x2-y2 | 4.647 |
|---|
| xy | -0.371 |
|---|
| xz | 2.074 |
|---|
| yz | -0.622 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.317 |
0.059 |
0.147 |
| y |
0.059 |
5.644 |
-0.000 |
| z |
0.147 |
-0.000 |
5.353 |
<r2> (average value of r
2) Å
2
| <r2> |
94.380 |
| (<r2>)1/2 |
9.715 |
