return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Butanol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.037672
Energy at 298.15K-233.048886
HF Energy-232.192560
Nuclear repulsion energy185.375931
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3855 3651 17.21      
2 A' 3167 2999 33.14      
3 A' 3088 2924 42.36      
4 A' 3077 2914 22.95      
5 A' 3071 2908 23.25      
6 A' 3031 2870 53.65      
7 A' 1570 1487 2.69      
8 A' 1548 1466 9.06      
9 A' 1535 1454 1.53      
10 A' 1529 1448 0.22      
11 A' 1499 1420 3.97      
12 A' 1450 1373 5.84      
13 A' 1420 1345 3.15      
14 A' 1334 1264 36.23      
15 A' 1273 1206 40.05      
16 A' 1143 1082 2.95      
17 A' 1107 1048 22.39      
18 A' 1102 1044 58.47      
19 A' 1041 986 0.51      
20 A' 930 881 14.55      
21 A' 449 426 15.01      
22 A' 404 383 0.09      
23 A' 189 179 3.09      
24 A" 3167 2999 56.29      
25 A" 3147 2980 32.73      
26 A" 3114 2949 4.98      
27 A" 3075 2912 47.69      
28 A" 1542 1460 10.14      
29 A" 1358 1287 0.18      
30 A" 1351 1280 0.85      
31 A" 1279 1212 0.18      
32 A" 1216 1152 2.46      
33 A" 976 925 0.15      
34 A" 832 788 1.54      
35 A" 752 712 2.37      
36 A" 277 263 137.52      
37 A" 255 241 4.64      
38 A" 120 113 0.80      
39 A" 107 102 7.79      

Unscaled Zero Point Vibrational Energy (zpe) 30689.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29065.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.62505 0.06645 0.06288

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.351 -0.345 0.000
C2 0.000 0.337 0.000
C3 -1.162 -0.651 0.000
C4 -2.520 0.046 0.000
O5 2.348 0.669 0.000
H6 1.442 -0.986 0.889
H7 1.442 -0.986 -0.889
H8 -0.057 0.987 0.880
H9 -0.057 0.987 -0.880
H10 -1.085 -1.303 0.878
H11 -1.085 -1.303 -0.878
H12 -3.341 -0.675 0.000
H13 -2.631 0.681 0.883
H14 -2.631 0.681 -0.883
H15 3.208 0.243 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.51352.53173.89041.42151.09941.09942.12882.12882.76122.76124.70364.20604.20601.9479
C21.51351.52542.53652.37092.14962.14961.09521.09522.15382.15383.49092.79652.79653.2095
C32.53171.52541.52603.74962.77222.77222.16252.16251.09621.09622.17902.17062.17064.4607
C43.89042.53651.52604.90694.18964.18962.77882.77882.15582.15581.09271.09311.09315.7311
O51.42152.37093.74964.90692.08482.08482.58072.58074.05504.05505.84505.05645.05640.9600
H61.09942.14962.77224.18962.08481.77732.47813.04442.54753.10014.87524.40144.74452.3275
H71.09942.14962.77224.18962.08481.77733.04442.47813.10012.54754.87524.74454.40142.3275
H82.12881.09522.16252.77882.58072.47813.04441.75992.50973.06413.78372.59183.13443.4629
H92.12881.09522.16252.77882.58073.04442.47811.75993.06412.50973.78373.13442.59183.4629
H102.76122.15381.09622.15584.05502.54753.10012.50973.06411.75582.50072.51513.07014.6469
H112.76122.15381.09622.15584.05503.10012.54753.06412.50971.75582.50073.07012.51514.6469
H124.70363.49092.17901.09275.84504.87524.87523.78373.78372.50072.50071.76671.76676.6130
H134.20602.79652.17061.09315.05644.40144.74452.59183.13442.51513.07011.76671.76545.9218
H144.20602.79652.17061.09315.05644.74454.40143.13442.59183.07012.51511.76671.76545.9218
H151.94793.20954.46075.73110.96002.32752.32753.46293.46294.64694.64696.61305.92185.9218

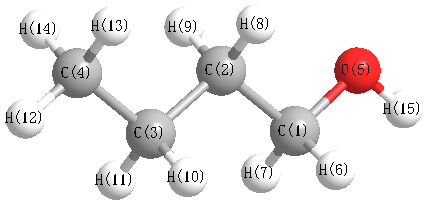
picture of 1-Butanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.833 C1 C2 H8 108.311
C1 C2 H9 108.311 C1 O5 H15 108.193
C2 C1 O5 107.721 C2 C1 H6 109.689
C2 C1 H7 109.689 C2 C3 C4 112.453
C2 C3 H10 109.386 C2 C3 H11 109.386
C3 C2 H8 110.131 C3 C2 H9 110.131
C3 C4 H12 111.559 C3 C4 H13 110.857
C3 C4 H14 110.857 C4 C3 H10 109.506
C4 C3 H11 109.506 O5 C1 H6 110.941
O5 C1 H7 110.941 H6 C1 H7 107.859
H8 C2 H9 106.924 H10 C3 H11 106.422
H12 C4 H13 107.851 H12 C4 H14 107.851
H13 C4 H14 107.706
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability