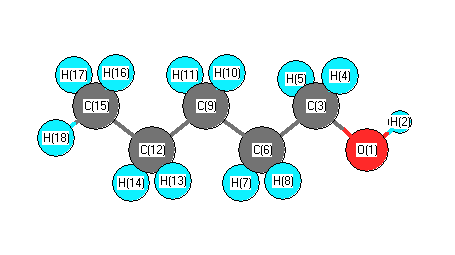
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3825 |
3683 |
9.58 |
|
|
|
| 2 |
A' |
3105 |
2989 |
47.06 |
|
|
|
| 3 |
A' |
3039 |
2926 |
64.54 |
|
|
|
| 4 |
A' |
3031 |
2918 |
41.19 |
|
|
|
| 5 |
A' |
3024 |
2912 |
25.83 |
|
|
|
| 6 |
A' |
3009 |
2897 |
19.61 |
|
|
|
| 7 |
A' |
2969 |
2858 |
61.63 |
|
|
|
| 8 |
A' |
1538 |
1481 |
3.44 |
|
|
|
| 9 |
A' |
1521 |
1464 |
9.48 |
|
|
|
| 10 |
A' |
1511 |
1454 |
1.17 |
|
|
|
| 11 |
A' |
1498 |
1443 |
0.73 |
|
|
|
| 12 |
A' |
1495 |
1440 |
0.00 |
|
|
|
| 13 |
A' |
1474 |
1419 |
3.91 |
|
|
|
| 14 |
A' |
1419 |
1366 |
5.26 |
|
|
|
| 15 |
A' |
1412 |
1360 |
0.26 |
|
|
|
| 16 |
A' |
1364 |
1313 |
12.29 |
|
|
|
| 17 |
A' |
1303 |
1255 |
38.04 |
|
|
|
| 18 |
A' |
1245 |
1198 |
25.93 |
|
|
|
| 19 |
A' |
1133 |
1090 |
3.33 |
|
|
|
| 20 |
A' |
1086 |
1045 |
89.87 |
|
|
|
| 21 |
A' |
1085 |
1045 |
0.23 |
|
|
|
| 22 |
A' |
1053 |
1013 |
2.88 |
|
|
|
| 23 |
A' |
1015 |
977 |
14.62 |
|
|
|
| 24 |
A' |
907 |
873 |
4.17 |
|
|
|
| 25 |
A' |
497 |
478 |
13.15 |
|
|
|
| 26 |
A' |
364 |
350 |
0.07 |
|
|
|
| 27 |
A' |
314 |
302 |
5.17 |
|
|
|
| 28 |
A' |
134 |
129 |
1.72 |
|
|
|
| 29 |
A" |
3100 |
2984 |
92.40 |
|
|
|
| 30 |
A" |
3085 |
2970 |
57.29 |
|
|
|
| 31 |
A" |
3059 |
2945 |
10.24 |
|
|
|
| 32 |
A" |
3033 |
2920 |
3.54 |
|
|
|
| 33 |
A" |
2996 |
2885 |
58.23 |
|
|
|
| 34 |
A" |
1513 |
1456 |
9.72 |
|
|
|
| 35 |
A" |
1347 |
1297 |
0.67 |
|
|
|
| 36 |
A" |
1334 |
1284 |
0.76 |
|
|
|
| 37 |
A" |
1309 |
1260 |
0.01 |
|
|
|
| 38 |
A" |
1244 |
1197 |
0.53 |
|
|
|
| 39 |
A" |
1193 |
1148 |
1.23 |
|
|
|
| 40 |
A" |
996 |
959 |
1.58 |
|
|
|
| 41 |
A" |
866 |
834 |
0.02 |
|
|
|
| 42 |
A" |
776 |
747 |
0.32 |
|
|
|
| 43 |
A" |
736 |
708 |
4.15 |
|
|
|
| 44 |
A" |
301 |
289 |
138.99 |
|
|
|
| 45 |
A" |
246 |
237 |
0.36 |
|
|
|
| 46 |
A" |
151 |
146 |
1.42 |
|
|
|
| 47 |
A" |
98 |
94 |
4.48 |
|
|
|
| 48 |
A" |
76 |
73 |
1.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36412.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 35054.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.577 |
|
|
|
| 2 |
H |
0.389 |
|
|
|
| 3 |
C |
-0.172 |
|
|
|
| 4 |
H |
0.187 |
|
|
|
| 5 |
H |
0.187 |
|
|
|
| 6 |
C |
-0.454 |
|
|
|
| 7 |
H |
0.224 |
|
|
|
| 8 |
H |
0.224 |
|
|
|
| 9 |
C |
-0.434 |
|
|
|
| 10 |
H |
0.209 |
|
|
|
| 11 |
H |
0.209 |
|
|
|
| 12 |
C |
-0.428 |
|
|
|
| 13 |
H |
0.215 |
|
|
|
| 14 |
H |
0.215 |
|
|
|
| 15 |
C |
-0.650 |
|
|
|
| 16 |
H |
0.216 |
|
|
|
| 17 |
H |
0.216 |
|
|
|
| 18 |
H |
0.222 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.396 |
0.703 |
0.000 |
1.563 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.569 |
-4.387 |
0.000 |
| y |
-4.387 |
-43.317 |
0.000 |
| z |
0.000 |
0.000 |
-39.599 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.889 |
-4.387 |
0.000 |
| y |
-4.387 |
-6.233 |
0.000 |
| z |
0.000 |
0.000 |
-0.656 |
|
| Polar |
|---|
| 3z2-r2 | -1.312 |
|---|
| x2-y2 | 8.748 |
|---|
| xy | -4.387 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
301.085 |
| (<r2>)1/2 |
17.352 |