return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (1-Pentanol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-272.236696
Energy at 298.15K-272.250104
HF Energy-271.233901
Nuclear repulsion energy244.923678
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3855 3651 17.35      
2 A' 3165 2998 33.70      
3 A' 3088 2925 44.30      
4 A' 3076 2913 45.10      
5 A' 3075 2912 11.11      
6 A' 3057 2895 19.03      
7 A' 3030 2869 50.32      
8 A' 1569 1486 3.07      
9 A' 1549 1467 10.88      
10 A' 1538 1457 0.95      
11 A' 1528 1448 1.12      
12 A' 1526 1446 0.01      
13 A' 1499 1420 4.32      
14 A' 1449 1372 6.39      
15 A' 1432 1356 0.17      
16 A' 1379 1306 10.20      
17 A' 1314 1245 46.71      
18 A' 1259 1193 19.44      
19 A' 1150 1089 3.45      
20 A' 1110 1052 2.66      
21 A' 1108 1049 74.78      
22 A' 1075 1018 6.28      
23 A' 1037 983 13.10      
24 A' 925 876 4.29      
25 A' 507 480 13.54      
26 A' 372 353 0.11      
27 A' 322 305 5.53      
28 A' 137 130 1.86      
29 A" 3163 2996 59.42      
30 A" 3148 2982 48.39      
31 A" 3125 2960 10.61      
32 A" 3100 2936 4.20      
33 A" 3074 2911 46.89      
34 A" 1541 1460 10.11      
35 A" 1368 1295 0.93      
36 A" 1353 1282 0.69      
37 A" 1332 1262 0.00      
38 A" 1265 1198 0.37      
39 A" 1216 1152 1.51      
40 A" 1010 957 1.46      
41 A" 882 835 0.06      
42 A" 786 745 0.43      
43 A" 737 698 3.70      
44 A" 276 261 138.79      
45 A" 252 239 1.86      
46 A" 156 147 2.95      
47 A" 98 92 3.75      
48 A" 77 73 3.36      

Unscaled Zero Point Vibrational Energy (zpe) 37044.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 35084.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.52799 0.03830 0.03694

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 1.299 -2.792 0.000
H2 2.184 -3.164 0.000
C3 1.412 -1.375 0.000
H4 1.956 -1.024 0.889
H5 1.956 -1.024 -0.889
C6 0.011 -0.801 0.000
H7 -0.517 -1.183 -0.880
H8 -0.517 -1.183 0.880
C9 0.000 0.724 0.000
H10 0.543 1.096 0.879
H11 0.543 1.096 -0.879
C12 -1.410 1.310 0.000
H13 -1.951 0.938 0.877
H14 -1.951 0.938 -0.877
C15 -1.414 2.835 0.000
H16 -0.902 3.227 0.883
H17 -0.902 3.227 -0.883
H18 -2.431 3.235 0.000

Atom - Atom Distances (Å)
  O1 H2 C3 H4 H5 C6 H7 H8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
O10.96001.42112.08492.08492.37082.58092.58093.74834.05704.05704.91515.02425.02426.24716.46946.46947.0876
H20.96001.94782.32792.32793.20953.46313.46314.45914.64854.64855.73785.88985.88986.99527.15157.15157.8891
C31.42111.94781.09951.09951.51362.12882.12882.53002.76302.76303.89484.17494.17495.07115.22655.22656.0018
H42.08492.32791.09951.77722.14943.04432.47792.76962.54773.10074.19064.37184.71505.20015.12245.42016.1784
H52.08492.32791.09951.77722.14942.47793.04432.76963.10072.54774.19064.71504.37185.20015.42015.12246.1784
C62.37083.20951.51362.14942.14941.09511.09511.52522.15702.15702.54452.76502.76503.90594.22394.22394.7175
H72.58093.46312.12883.04432.47791.09511.76002.16283.06722.51302.78983.10512.56004.21014.76494.42674.8942
H82.58093.46312.12882.47793.04431.09511.76002.16282.51303.06722.78982.56003.10514.21014.42674.76494.8942
C93.74834.45912.53002.76962.76961.52522.16282.16281.09761.09761.52632.14992.14992.54112.80332.80333.4949
H104.05704.64852.76302.54773.10072.15703.06722.51301.09761.75732.15152.49883.05412.76172.57473.11973.7671
H114.05704.64852.76303.10072.54772.15702.51303.06721.09761.75732.15153.05412.49882.76173.11972.57473.7671
C124.91515.73783.89484.19064.19062.54452.78982.78981.52632.15152.15151.09601.09601.52582.17132.17132.1794
H135.02425.88984.17494.37184.71502.76503.10512.56002.14992.49883.05411.09601.75472.15852.51843.07262.5052
H145.02425.88984.17494.71504.37182.76502.56003.10512.14993.05412.49881.09601.75472.15853.07262.51842.5052
C156.24716.99525.07115.20015.20013.90594.21014.21012.54112.76172.76171.52582.15852.15851.09341.09341.0928
H166.46947.15155.22655.12245.42014.22394.76494.42672.80332.57473.11972.17132.51843.07261.09341.76601.7661
H176.46947.15155.22655.42015.12244.22394.42674.76492.80333.11972.57472.17133.07262.51841.09341.76601.7661
H187.08767.88916.00186.17846.17844.71754.89424.89423.49493.76713.76712.17942.50522.50521.09281.76611.7661

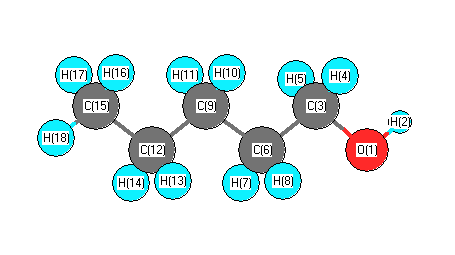
picture of 1-Pentanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C3 H4 110.969 O1 C3 H5 110.969
O1 C3 C6 107.732 H2 O1 C3 108.209
C3 C6 H7 108.314 C3 C6 H8 108.314
C3 C6 C9 112.727 H4 C3 H5 107.838
H4 C3 C6 109.665 H5 C3 C6 109.665
C6 C9 H10 109.571 C6 C9 H11 109.571
C6 C9 C12 112.989 H7 C6 H8 106.945
H7 C6 C9 110.174 H8 C6 C9 110.174
C9 C12 H13 109.036 C9 C12 H14 109.036
C9 C12 C15 112.727 H10 C9 H11 106.357
H10 C9 C12 109.067 H11 C9 C12 109.067
C12 C15 H16 110.914 C12 C15 H17 110.914
C12 C15 H18 111.599 H13 C12 H14 106.353
H13 C12 C15 109.742 H14 C12 C15 109.742
H16 C15 H17 107.716 H16 C15 H18 107.766
H17 C15 H18 107.766
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability