return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,4-Hexadiene, (E)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.948109
Energy at 298.15K-233.957731
HF Energy-233.027077
Nuclear repulsion energy214.877511
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3285 3111 13.71      
2 A 3192 3023 10.09      
3 A 3178 3010 43.13      
4 A 3176 3008 21.28      
5 A 3168 3000 7.27      
6 A 3162 2995 2.67      
7 A 3144 2978 18.17      
8 A 3101 2937 13.89      
9 A 3071 2908 26.20      
10 A 3058 2896 25.02      
11 A 1739 1647 0.95      
12 A 1703 1613 8.66      
13 A 1535 1454 12.82      
14 A 1519 1438 8.03      
15 A 1512 1432 12.66      
16 A 1460 1383 4.76      
17 A 1451 1375 4.95      
18 A 1389 1316 0.64      
19 A 1353 1281 0.71      
20 A 1338 1267 1.49      
21 A 1335 1264 0.93      
22 A 1246 1180 1.05      
23 A 1154 1093 3.24      
24 A 1136 1076 4.86      
25 A 1082 1025 0.42      
26 A 1067 1011 4.53      
27 A 1027 973 18.88      
28 A 992 940 42.15      
29 A 978 926 6.53      
30 A 942 892 1.64      
31 A 902 855 16.11      
32 A 899 851 22.98      
33 A 776 735 0.26      
34 A 606 574 5.79      
35 A 545 516 8.99      
36 A 472 447 0.36      
37 A 324 307 0.55      
38 A 286 270 1.52      
39 A 201 191 1.09      
40 A 156 148 0.58      
41 A 129 122 0.06      
42 A 75 71 0.29      

Unscaled Zero Point Vibrational Energy (zpe) 31430.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29767.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.27060 0.05839 0.05123

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.733 0.461 0.036
H2 2.945 1.315 -0.613
H3 2.677 0.821 1.066
H4 3.586 -0.219 -0.041
C5 1.460 -0.221 -0.367
H6 1.413 -0.613 -1.384
C7 0.394 -0.380 0.430
H8 0.427 0.015 1.446
C9 -0.878 -1.060 0.021
H10 -0.739 -1.556 -0.948
H11 -1.109 -1.862 0.735
C12 -2.068 -0.136 -0.060
H13 -3.043 -0.618 -0.139
C14 -2.006 1.201 -0.060
H15 -2.905 1.803 -0.130
H16 -1.058 1.724 0.005

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 H10 H11 C12 H13 C14 H15 H16
C11.09421.09211.09421.49922.21612.51672.73933.91804.13424.54314.83905.87784.79665.79773.9954
H21.09421.77091.75862.15112.58053.23643.50364.54574.68315.32445.24886.31014.98305.89074.0710
H31.09211.77091.77062.14953.10732.65772.42064.15584.62394.65224.97046.02004.83145.79293.9866
H41.09421.75861.77062.15112.58423.23113.50014.54294.61715.03405.65526.64155.76936.79965.0344
C51.49922.15112.14952.15111.09091.34042.09992.51392.63753.24113.54254.52563.75824.81703.2029
H62.21612.58053.10732.58421.09092.09333.06202.72442.38983.52253.75524.62644.09015.10453.6735
C72.51673.23642.65773.23111.34042.09331.09121.49872.13652.13212.52213.49112.91503.99522.5917
H82.73933.50362.42063.50012.09993.06201.09122.21123.09222.52742.91813.86643.09694.09632.6837
C93.91804.54574.15584.54292.51392.72441.49872.21121.09761.09811.50962.21542.52853.51162.7907
H104.13424.68314.62394.61712.63752.38982.13653.09221.09761.75042.13852.61563.16134.07953.4307
H114.54315.32444.65225.03403.24113.52252.13212.52741.09811.75042.12932.46053.28984.17253.6608
C124.83905.24884.97045.65523.54253.75522.52212.91811.50962.13852.12931.08981.33852.11272.1181
H135.87786.31016.02006.64154.52564.62643.49113.86642.21542.61562.46051.08982.09522.42443.0735
C144.79664.98304.83145.76933.75824.09012.91503.09692.52853.16133.28981.33852.09521.08451.0847
H155.79775.89075.79296.79964.81705.10453.99524.09633.51164.07954.17252.11272.42441.08451.8541
H163.99544.07103.98665.03443.20293.67352.59172.68372.79073.43073.66082.11813.07351.08471.8541

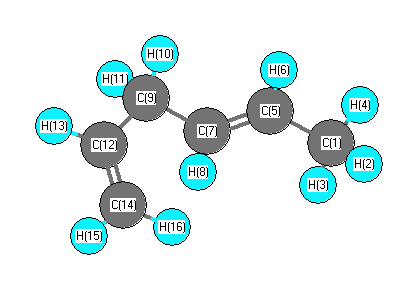
picture of 1,4-Hexadiene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 116.772 C1 C5 C7 124.728
H2 C1 H3 108.195 H2 C1 H4 106.948
H2 C1 C5 111.125 H3 C1 H4 108.159
H3 C1 C5 111.127 H4 C1 C5 111.122
C5 C7 H8 119.097 C5 C7 C9 124.528
H6 C5 C7 118.500 C7 C9 H10 109.789
C7 C9 H11 109.412 C7 C9 C12 113.939
H8 C7 C9 116.370 C9 C12 H13 115.986
C9 C12 C14 125.084 H10 C9 H11 105.727
H10 C9 C12 109.194 H11 C9 C12 108.449
C12 C14 H15 121.012 C12 C14 H16 121.527
H13 C12 C14 118.922 H15 C14 H16 117.460
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability