Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -79.562787 |
| Energy at 298.15K | -79.568733 |
| HF Energy | -79.242649 |
| Nuclear repulsion energy | 42.256520 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3087 |
2924 |
0.00 |
226.91 |
0.01 |
0.02 |
| 2 |
A1g |
1465 |
1387 |
0.00 |
3.09 |
0.61 |
0.76 |
| 3 |
A1g |
1040 |
985 |
0.00 |
10.55 |
0.27 |
0.43 |
| 4 |
A1u |
324 |
307 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3087 |
2924 |
53.37 |
0.00 |
0.01 |
0.02 |
| 6 |
A2u |
1444 |
1368 |
5.73 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3161 |
2994 |
0.00 |
135.85 |
0.75 |
0.86 |
| 7 |
Eg |
3161 |
2994 |
0.00 |
135.85 |
0.75 |
0.86 |
| 8 |
Eg |
1547 |
1465 |
0.00 |
25.87 |
0.75 |
0.86 |
| 8 |
Eg |
1547 |
1465 |
0.00 |
25.87 |
0.75 |
0.86 |
| 9 |
Eg |
1250 |
1184 |
0.00 |
1.19 |
0.75 |
0.86 |
| 9 |
Eg |
1250 |
1184 |
0.00 |
1.19 |
0.75 |
0.86 |
| 10 |
Eu |
3183 |
3014 |
59.82 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3183 |
3014 |
59.82 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1549 |
1467 |
14.01 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1549 |
1467 |
14.01 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
838 |
794 |
4.96 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
838 |
794 |
4.96 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16751.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15864.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.763 |
| C2 |
0.000 |
0.000 |
-0.763 |
| H3 |
0.000 |
1.018 |
1.159 |
| H4 |
-0.881 |
-0.509 |
1.159 |
| H5 |
0.881 |
-0.509 |
1.159 |
| H6 |
0.000 |
-1.018 |
-1.159 |
| H7 |
-0.881 |
0.509 |
-1.159 |
| H8 |
0.881 |
0.509 |
-1.159 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5262 | 1.0920 | 1.0920 | 1.0920 | 2.1749 | 2.1749 | 2.1749 |
C2 | 1.5262 | | 2.1749 | 2.1749 | 2.1749 | 1.0920 | 1.0920 | 1.0920 | H3 | 1.0920 | 2.1749 | | 1.7628 | 1.7628 | 3.0848 | 2.5315 | 2.5315 | H4 | 1.0920 | 2.1749 | 1.7628 | | 1.7628 | 2.5315 | 2.5315 | 3.0848 | H5 | 1.0920 | 2.1749 | 1.7628 | 1.7628 | | 2.5315 | 3.0848 | 2.5315 | H6 | 2.1749 | 1.0920 | 3.0848 | 2.5315 | 2.5315 | | 1.7628 | 1.7628 | H7 | 2.1749 | 1.0920 | 2.5315 | 2.5315 | 3.0848 | 1.7628 | | 1.7628 | H8 | 2.1749 | 1.0920 | 2.5315 | 3.0848 | 2.5315 | 1.7628 | 1.7628 | |
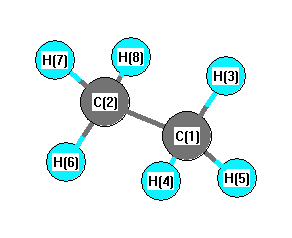 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.252 |
|
C1 |
C2 |
H7 |
111.252 |
| C1 |
C2 |
H8 |
111.252 |
|
C2 |
C1 |
H3 |
111.252 |
| C2 |
C1 |
H4 |
111.252 |
|
C2 |
C1 |
H5 |
111.252 |
| H3 |
C1 |
H4 |
107.634 |
|
H3 |
C1 |
H5 |
107.634 |
| H4 |
C1 |
H5 |
107.634 |
|
H6 |
C2 |
H7 |
107.634 |
| H6 |
C2 |
H8 |
107.634 |
|
H7 |
C2 |
H8 |
107.634 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -79.558026 |
| Energy at 298.15K | |
| HF Energy | -79.237916 |
| Nuclear repulsion energy | 42.091512 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3100 |
2936 |
0.00 |
|
|
|
| 2 |
A1' |
1490 |
1411 |
0.00 |
|
|
|
| 3 |
A1' |
1037 |
982 |
0.00 |
|
|
|
| 4 |
A1" |
314i |
297i |
0.00 |
|
|
|
| 5 |
A2" |
3094 |
2930 |
55.61 |
|
|
|
| 6 |
A2" |
1446 |
1369 |
5.40 |
|
|
|
| 7 |
E' |
3188 |
3019 |
50.91 |
|
|
|
| 7 |
E' |
3188 |
3019 |
50.91 |
|
|
|
| 8 |
E' |
1556 |
1474 |
15.26 |
|
|
|
| 8 |
E' |
1556 |
1474 |
15.26 |
|
|
|
| 9 |
E' |
925 |
876 |
4.62 |
|
|
|
| 9 |
E' |
925 |
876 |
4.62 |
|
|
|
| 10 |
E" |
3168 |
3000 |
0.00 |
|
|
|
| 10 |
E" |
3168 |
3000 |
0.00 |
|
|
|
| 11 |
E" |
1544 |
1462 |
0.00 |
|
|
|
| 11 |
E" |
1544 |
1462 |
0.00 |
|
|
|
| 12 |
E" |
1198 |
1134 |
0.00 |
|
|
|
| 12 |
E" |
1198 |
1134 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16504.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15631.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.770 |
| C2 |
0.000 |
0.000 |
-0.770 |
| H3 |
0.000 |
1.013 |
1.174 |
| H4 |
-0.877 |
-0.507 |
1.174 |
| H5 |
0.877 |
-0.507 |
1.174 |
| H6 |
0.000 |
1.013 |
-1.174 |
| H7 |
0.877 |
-0.507 |
-1.174 |
| H8 |
-0.877 |
-0.507 |
-1.174 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5397 | 1.0910 | 1.0910 | 1.0910 | 2.1925 | 2.1925 | 2.1925 |
C2 | 1.5397 | | 2.1925 | 2.1925 | 2.1925 | 1.0910 | 1.0910 | 1.0910 | H3 | 1.0910 | 2.1925 | | 1.7549 | 1.7549 | 2.3490 | 2.9321 | 2.9321 | H4 | 1.0910 | 2.1925 | 1.7549 | | 1.7549 | 2.9321 | 2.9321 | 2.3490 | H5 | 1.0910 | 2.1925 | 1.7549 | 1.7549 | | 2.9321 | 2.3490 | 2.9321 | H6 | 2.1925 | 1.0910 | 2.3490 | 2.9321 | 2.9321 | | 1.7549 | 1.7549 | H7 | 2.1925 | 1.0910 | 2.9321 | 2.9321 | 2.3490 | 1.7549 | | 1.7549 | H8 | 2.1925 | 1.0910 | 2.9321 | 2.3490 | 2.9321 | 1.7549 | 1.7549 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.771 |
|
C1 |
C2 |
H7 |
111.771 |
| C1 |
C2 |
H8 |
111.771 |
|
C2 |
C1 |
H3 |
111.771 |
| C2 |
C1 |
H4 |
111.771 |
|
C2 |
C1 |
H5 |
111.771 |
| H3 |
C1 |
H4 |
107.076 |
|
H3 |
C1 |
H5 |
107.076 |
| H4 |
C1 |
H5 |
107.076 |
|
H6 |
C2 |
H7 |
107.076 |
| H6 |
C2 |
H8 |
107.076 |
|
H7 |
C2 |
H8 |
107.076 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
