Jump to
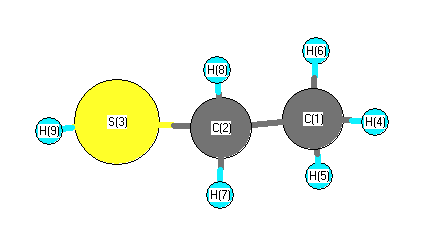
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -477.336600 |
| Energy at 298.15K | -477.342913 |
| HF Energy | -476.771784 |
| Nuclear repulsion energy | 107.728267 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3179 |
3010 |
19.20 |
|
|
|
| 2 |
A' |
3110 |
2946 |
24.42 |
|
|
|
| 3 |
A' |
3089 |
2925 |
17.51 |
|
|
|
| 4 |
A' |
2686 |
2544 |
11.42 |
|
|
|
| 5 |
A' |
1544 |
1463 |
3.54 |
|
|
|
| 6 |
A' |
1529 |
1448 |
3.97 |
|
|
|
| 7 |
A' |
1454 |
1377 |
7.76 |
|
|
|
| 8 |
A' |
1359 |
1287 |
41.53 |
|
|
|
| 9 |
A' |
1145 |
1084 |
2.83 |
|
|
|
| 10 |
A' |
1030 |
976 |
2.39 |
|
|
|
| 11 |
A' |
887 |
840 |
2.40 |
|
|
|
| 12 |
A' |
715 |
678 |
0.39 |
|
|
|
| 13 |
A' |
315 |
299 |
2.30 |
|
|
|
| 14 |
A" |
3188 |
3020 |
25.43 |
|
|
|
| 15 |
A" |
3165 |
2997 |
0.65 |
|
|
|
| 16 |
A" |
1534 |
1453 |
11.83 |
|
|
|
| 17 |
A" |
1306 |
1237 |
1.91 |
|
|
|
| 18 |
A" |
1093 |
1035 |
2.51 |
|
|
|
| 19 |
A" |
814 |
771 |
6.35 |
|
|
|
| 20 |
A" |
276 |
262 |
0.98 |
|
|
|
| 21 |
A" |
166 |
157 |
26.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16792.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15903.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.515 |
0.672 |
0.000 |
| C2 |
0.000 |
0.828 |
0.000 |
| S3 |
-0.757 |
-0.828 |
0.000 |
| H4 |
1.995 |
1.653 |
0.000 |
| H5 |
1.852 |
0.128 |
0.885 |
| H6 |
1.852 |
0.128 |
-0.885 |
| H7 |
-0.320 |
1.378 |
0.887 |
| H8 |
-0.320 |
1.378 |
-0.887 |
| H9 |
-2.038 |
-0.420 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5226 | 2.7217 | 1.0921 | 1.0917 | 1.0917 | 2.1562 | 2.1562 | 3.7162 |
C2 | 1.5226 | | 1.8201 | 2.1587 | 2.1683 | 2.1683 | 1.0912 | 1.0912 | 2.3894 | S3 | 2.7217 | 1.8201 | | 3.7045 | 2.9157 | 2.9157 | 2.4165 | 2.4165 | 1.3440 | H4 | 1.0921 | 2.1587 | 3.7045 | | 1.7682 | 1.7682 | 2.4940 | 2.4940 | 4.5340 | H5 | 1.0917 | 2.1683 | 2.9157 | 1.7682 | | 1.7690 | 2.5055 | 3.0683 | 4.0263 | H6 | 1.0917 | 2.1683 | 2.9157 | 1.7682 | 1.7690 | | 3.0683 | 2.5055 | 4.0263 | H7 | 2.1562 | 1.0912 | 2.4165 | 2.4940 | 2.5055 | 3.0683 | | 1.7734 | 2.6398 | H8 | 2.1562 | 1.0912 | 2.4165 | 2.4940 | 3.0683 | 2.5055 | 1.7734 | | 2.6398 | H9 | 3.7162 | 2.3894 | 1.3440 | 4.5340 | 4.0263 | 4.0263 | 2.6398 | 2.6398 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.694 |
|
C1 |
C2 |
H7 |
110.069 |
| C1 |
C2 |
H8 |
110.069 |
|
C2 |
C1 |
H4 |
110.215 |
| C2 |
C1 |
H5 |
111.001 |
|
C2 |
C1 |
H6 |
111.001 |
| C2 |
S3 |
H9 |
96.928 |
|
S3 |
C2 |
H7 |
109.654 |
| S3 |
C2 |
H8 |
109.654 |
|
H4 |
C1 |
H5 |
108.140 |
| H4 |
C1 |
H6 |
108.140 |
|
H5 |
C1 |
H6 |
108.241 |
| H7 |
C2 |
H8 |
108.691 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
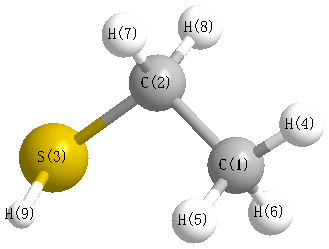
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -477.337469 |
| Energy at 298.15K | -477.343881 |
| HF Energy | -476.772289 |
| Nuclear repulsion energy | 107.494945 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3189 |
3021 |
23.65 |
|
|
|
| 2 |
A |
3170 |
3002 |
16.08 |
|
|
|
| 3 |
A |
3161 |
2993 |
11.04 |
|
|
|
| 4 |
A |
3112 |
2947 |
16.74 |
|
|
|
| 5 |
A |
3081 |
2918 |
22.08 |
|
|
|
| 6 |
A |
2682 |
2540 |
10.52 |
|
|
|
| 7 |
A |
1538 |
1457 |
5.18 |
|
|
|
| 8 |
A |
1532 |
1451 |
12.52 |
|
|
|
| 9 |
A |
1516 |
1436 |
2.10 |
|
|
|
| 10 |
A |
1453 |
1376 |
7.10 |
|
|
|
| 11 |
A |
1361 |
1289 |
19.75 |
|
|
|
| 12 |
A |
1318 |
1248 |
3.84 |
|
|
|
| 13 |
A |
1163 |
1101 |
13.38 |
|
|
|
| 14 |
A |
1105 |
1046 |
0.40 |
|
|
|
| 15 |
A |
1022 |
968 |
6.80 |
|
|
|
| 16 |
A |
899 |
851 |
9.16 |
|
|
|
| 17 |
A |
762 |
721 |
3.10 |
|
|
|
| 18 |
A |
700 |
663 |
2.64 |
|
|
|
| 19 |
A |
342 |
324 |
1.82 |
|
|
|
| 20 |
A |
286 |
271 |
1.37 |
|
|
|
| 21 |
A |
217 |
206 |
22.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16803.5 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15914.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.628 |
-0.351 |
-0.053 |
| C2 |
0.490 |
0.648 |
0.090 |
| S3 |
-1.157 |
-0.101 |
-0.079 |
| H4 |
2.595 |
0.155 |
0.016 |
| H5 |
1.588 |
-1.107 |
0.734 |
| H6 |
1.572 |
-0.865 |
-1.014 |
| H7 |
0.552 |
1.178 |
1.042 |
| H8 |
0.539 |
1.398 |
-0.702 |
| H9 |
-1.046 |
-0.927 |
0.976 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5209 | 2.7962 | 1.0936 | 1.0924 | 1.0909 | 2.1672 | 2.1603 | 2.9222 |
C2 | 1.5209 | | 1.8167 | 2.1632 | 2.1676 | 2.1628 | 1.0915 | 1.0922 | 2.3706 | S3 | 2.7962 | 1.8167 | | 3.7617 | 3.0341 | 2.9838 | 2.4107 | 2.3472 | 1.3443 | H4 | 1.0936 | 2.1632 | 3.7617 | | 1.7675 | 1.7743 | 2.5050 | 2.5077 | 3.9174 | H5 | 1.0924 | 2.1676 | 3.0341 | 1.7675 | | 1.7646 | 2.5280 | 3.0722 | 2.6505 | H6 | 1.0909 | 2.1628 | 2.9838 | 1.7743 | 1.7646 | | 3.0728 | 2.5075 | 3.2885 | H7 | 2.1672 | 1.0915 | 2.4107 | 2.5050 | 2.5280 | 3.0728 | | 1.7575 | 2.6429 | H8 | 2.1603 | 1.0922 | 2.3472 | 2.5077 | 3.0722 | 2.5075 | 1.7575 | | 3.2754 | H9 | 2.9222 | 2.3706 | 1.3443 | 3.9174 | 2.6505 | 3.2885 | 2.6429 | 3.2754 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.520 |
|
C1 |
C2 |
H7 |
111.045 |
| C1 |
C2 |
H8 |
110.451 |
|
C2 |
C1 |
H4 |
110.603 |
| C2 |
C1 |
H5 |
111.024 |
|
C2 |
C1 |
H6 |
110.726 |
| C2 |
S3 |
H9 |
96.021 |
|
S3 |
C2 |
H7 |
109.441 |
| S3 |
C2 |
H8 |
104.841 |
|
H4 |
C1 |
H5 |
107.911 |
| H4 |
C1 |
H6 |
108.625 |
|
H5 |
C1 |
H6 |
107.845 |
| H7 |
C2 |
H8 |
107.188 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
