Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -477.826575 |
| Energy at 298.15K | |
| HF Energy | -477.645772 |
| Nuclear repulsion energy | 107.233211 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3138 |
3138 |
29.26 |
|
|
|
| 2 |
A' |
3089 |
3089 |
26.19 |
|
|
|
| 3 |
A' |
3067 |
3067 |
22.43 |
|
|
|
| 4 |
A' |
2652 |
2652 |
15.86 |
|
|
|
| 5 |
A' |
1539 |
1539 |
3.06 |
|
|
|
| 6 |
A' |
1523 |
1523 |
3.33 |
|
|
|
| 7 |
A' |
1448 |
1448 |
5.57 |
|
|
|
| 8 |
A' |
1343 |
1343 |
48.50 |
|
|
|
| 9 |
A' |
1134 |
1134 |
2.60 |
|
|
|
| 10 |
A' |
1012 |
1012 |
3.11 |
|
|
|
| 11 |
A' |
876 |
876 |
2.37 |
|
|
|
| 12 |
A' |
682 |
682 |
1.44 |
|
|
|
| 13 |
A' |
309 |
309 |
2.37 |
|
|
|
| 14 |
A" |
3153 |
3153 |
37.07 |
|
|
|
| 15 |
A" |
3130 |
3130 |
0.12 |
|
|
|
| 16 |
A" |
1530 |
1530 |
10.81 |
|
|
|
| 17 |
A" |
1299 |
1299 |
1.60 |
|
|
|
| 18 |
A" |
1077 |
1077 |
2.40 |
|
|
|
| 19 |
A" |
808 |
808 |
6.20 |
|
|
|
| 20 |
A" |
265 |
265 |
1.06 |
|
|
|
| 21 |
A" |
171 |
171 |
23.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16621.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16621.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.517 |
0.692 |
0.000 |
| C2 |
0.000 |
0.832 |
0.000 |
| S3 |
-0.757 |
-0.840 |
0.000 |
| H4 |
1.985 |
1.679 |
0.000 |
| H5 |
1.865 |
0.155 |
0.884 |
| H6 |
1.865 |
0.155 |
-0.884 |
| H7 |
-0.331 |
1.375 |
0.885 |
| H8 |
-0.331 |
1.375 |
-0.885 |
| H9 |
-2.042 |
-0.437 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5234 | 2.7424 | 1.0917 | 1.0912 | 1.0912 | 2.1599 | 2.1599 | 3.7343 |
C2 | 1.5234 | | 1.8357 | 2.1583 | 2.1717 | 2.1717 | 1.0899 | 1.0899 | 2.4044 | S3 | 2.7424 | 1.8357 | | 3.7237 | 2.9402 | 2.9402 | 2.4232 | 2.4232 | 1.3473 | H4 | 1.0917 | 2.1583 | 3.7237 | | 1.7656 | 1.7656 | 2.4984 | 2.4984 | 4.5494 | H5 | 1.0912 | 2.1717 | 2.9402 | 1.7656 | | 1.7671 | 2.5121 | 3.0723 | 4.0493 | H6 | 1.0912 | 2.1717 | 2.9402 | 1.7656 | 1.7671 | | 3.0723 | 2.5121 | 4.0493 | H7 | 2.1599 | 1.0899 | 2.4232 | 2.4984 | 2.5121 | 3.0723 | | 1.7703 | 2.6445 | H8 | 2.1599 | 1.0899 | 2.4232 | 2.4984 | 3.0723 | 2.5121 | 1.7703 | | 2.6445 | H9 | 3.7343 | 2.4044 | 1.3473 | 4.5494 | 4.0493 | 4.0493 | 2.6445 | 2.6445 | |
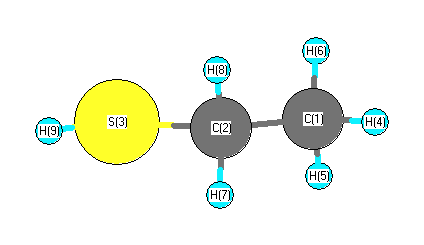 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.099 |
|
C1 |
C2 |
H7 |
110.379 |
| C1 |
C2 |
H8 |
110.379 |
|
C2 |
C1 |
H4 |
110.143 |
| C2 |
C1 |
H5 |
111.251 |
|
C2 |
C1 |
H6 |
111.251 |
| C2 |
S3 |
H9 |
96.921 |
|
S3 |
C2 |
H7 |
109.173 |
| S3 |
C2 |
H8 |
109.173 |
|
H4 |
C1 |
H5 |
107.964 |
| H4 |
C1 |
H6 |
107.964 |
|
H5 |
C1 |
H6 |
108.142 |
| H7 |
C2 |
H8 |
108.613 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -477.827432 |
| Energy at 298.15K | -477.833787 |
| HF Energy | -477.646484 |
| Nuclear repulsion energy | 106.981196 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3154 |
3154 |
34.88 |
|
|
|
| 2 |
A |
3133 |
3133 |
10.53 |
|
|
|
| 3 |
A |
3123 |
3123 |
27.60 |
|
|
|
| 4 |
A |
3091 |
3091 |
16.30 |
|
|
|
| 5 |
A |
3058 |
3058 |
29.04 |
|
|
|
| 6 |
A |
2650 |
2650 |
14.83 |
|
|
|
| 7 |
A |
1533 |
1533 |
4.21 |
|
|
|
| 8 |
A |
1526 |
1526 |
11.43 |
|
|
|
| 9 |
A |
1511 |
1511 |
1.85 |
|
|
|
| 10 |
A |
1445 |
1445 |
4.92 |
|
|
|
| 11 |
A |
1346 |
1346 |
22.71 |
|
|
|
| 12 |
A |
1311 |
1311 |
4.70 |
|
|
|
| 13 |
A |
1150 |
1150 |
13.68 |
|
|
|
| 14 |
A |
1090 |
1090 |
0.43 |
|
|
|
| 15 |
A |
1003 |
1003 |
7.41 |
|
|
|
| 16 |
A |
891 |
891 |
8.53 |
|
|
|
| 17 |
A |
752 |
752 |
2.66 |
|
|
|
| 18 |
A |
668 |
668 |
4.25 |
|
|
|
| 19 |
A |
333 |
333 |
1.50 |
|
|
|
| 20 |
A |
273 |
273 |
1.56 |
|
|
|
| 21 |
A |
217 |
217 |
20.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16628.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16628.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.641 |
-0.351 |
-0.053 |
| C2 |
0.498 |
0.645 |
0.090 |
| S3 |
-1.166 |
-0.101 |
-0.081 |
| H4 |
2.605 |
0.160 |
0.014 |
| H5 |
1.609 |
-1.105 |
0.736 |
| H6 |
1.590 |
-0.868 |
-1.011 |
| H7 |
0.553 |
1.178 |
1.040 |
| H8 |
0.540 |
1.395 |
-0.701 |
| H9 |
-1.079 |
-0.911 |
0.993 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5224 | 2.8178 | 1.0933 | 1.0919 | 1.0903 | 2.1713 | 2.1636 | 2.9672 |
C2 | 1.5224 | | 1.8318 | 2.1627 | 2.1711 | 2.1670 | 1.0901 | 1.0909 | 2.3927 | S3 | 2.8178 | 1.8318 | | 3.7809 | 3.0618 | 3.0085 | 2.4176 | 2.3517 | 1.3478 | H4 | 1.0933 | 2.1627 | 3.7809 | | 1.7650 | 1.7715 | 2.5096 | 2.5100 | 3.9595 | H5 | 1.0919 | 2.1711 | 3.0618 | 1.7650 | | 1.7634 | 2.5338 | 3.0756 | 2.7072 | H6 | 1.0903 | 2.1670 | 3.0085 | 1.7715 | 1.7634 | | 3.0772 | 2.5143 | 3.3383 | H7 | 2.1713 | 1.0901 | 2.4176 | 2.5096 | 2.5338 | 3.0772 | | 1.7543 | 2.6512 | H8 | 2.1636 | 1.0909 | 2.3517 | 2.5100 | 3.0756 | 2.5143 | 1.7543 | | 3.2873 | H9 | 2.9672 | 2.3927 | 1.3478 | 3.9595 | 2.7072 | 3.3383 | 2.6512 | 3.2873 | |
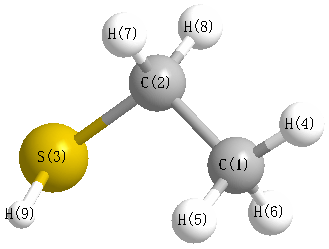 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.983 |
|
C1 |
C2 |
H7 |
111.358 |
| C1 |
C2 |
H8 |
110.690 |
|
C2 |
C1 |
H4 |
110.476 |
| C2 |
C1 |
H5 |
111.235 |
|
C2 |
C1 |
H6 |
110.995 |
| C2 |
S3 |
H9 |
96.427 |
|
S3 |
C2 |
H7 |
109.007 |
| S3 |
C2 |
H8 |
104.264 |
|
H4 |
C1 |
H5 |
107.747 |
| H4 |
C1 |
H6 |
108.438 |
|
H5 |
C1 |
H6 |
107.825 |
| H7 |
C2 |
H8 |
107.100 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
