Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/6-311G*
| | hartrees |
|---|
| Energy at 0K | -477.826571 |
| Energy at 298.15K | |
| HF Energy | -477.645769 |
| Nuclear repulsion energy | 107.232376 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3137 |
3137 |
29.19 |
|
|
|
| 2 |
A' |
3088 |
3088 |
26.23 |
|
|
|
| 3 |
A' |
3066 |
3066 |
22.45 |
|
|
|
| 4 |
A' |
2653 |
2653 |
15.84 |
|
|
|
| 5 |
A' |
1539 |
1539 |
3.05 |
|
|
|
| 6 |
A' |
1523 |
1523 |
3.38 |
|
|
|
| 7 |
A' |
1447 |
1447 |
5.58 |
|
|
|
| 8 |
A' |
1343 |
1343 |
48.43 |
|
|
|
| 9 |
A' |
1134 |
1134 |
2.62 |
|
|
|
| 10 |
A' |
1012 |
1012 |
3.12 |
|
|
|
| 11 |
A' |
877 |
877 |
2.35 |
|
|
|
| 12 |
A' |
681 |
681 |
1.45 |
|
|
|
| 13 |
A' |
308 |
308 |
2.37 |
|
|
|
| 14 |
A" |
3153 |
3153 |
37.13 |
|
|
|
| 15 |
A" |
3129 |
3129 |
0.08 |
|
|
|
| 16 |
A" |
1529 |
1529 |
10.81 |
|
|
|
| 17 |
A" |
1299 |
1299 |
1.60 |
|
|
|
| 18 |
A" |
1077 |
1077 |
2.42 |
|
|
|
| 19 |
A" |
808 |
808 |
6.18 |
|
|
|
| 20 |
A" |
264 |
264 |
1.23 |
|
|
|
| 21 |
A" |
178 |
178 |
22.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16622.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16622.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.517 |
0.693 |
0.000 |
| C2 |
0.000 |
0.832 |
0.000 |
| S3 |
-0.757 |
-0.841 |
0.000 |
| H4 |
1.985 |
1.679 |
0.000 |
| H5 |
1.865 |
0.155 |
0.884 |
| H6 |
1.865 |
0.155 |
-0.884 |
| H7 |
-0.331 |
1.375 |
0.885 |
| H8 |
-0.331 |
1.375 |
-0.885 |
| H9 |
-2.042 |
-0.437 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5234 | 2.7426 | 1.0917 | 1.0912 | 1.0912 | 2.1598 | 2.1598 | 3.7343 |
C2 | 1.5234 | | 1.8358 | 2.1582 | 2.1717 | 2.1717 | 1.0899 | 1.0899 | 2.4044 | S3 | 2.7426 | 1.8358 | | 3.7238 | 2.9403 | 2.9403 | 2.4232 | 2.4232 | 1.3472 | H4 | 1.0917 | 2.1582 | 3.7238 | | 1.7656 | 1.7656 | 2.4983 | 2.4983 | 4.5494 | H5 | 1.0912 | 2.1717 | 2.9403 | 1.7656 | | 1.7671 | 2.5120 | 3.0722 | 4.0493 | H6 | 1.0912 | 2.1717 | 2.9403 | 1.7656 | 1.7671 | | 3.0722 | 2.5120 | 4.0493 | H7 | 2.1598 | 1.0899 | 2.4232 | 2.4983 | 2.5120 | 3.0722 | | 1.7703 | 2.6445 | H8 | 2.1598 | 1.0899 | 2.4232 | 2.4983 | 3.0722 | 2.5120 | 1.7703 | | 2.6445 | H9 | 3.7343 | 2.4044 | 1.3472 | 4.5494 | 4.0493 | 4.0493 | 2.6445 | 2.6445 | |
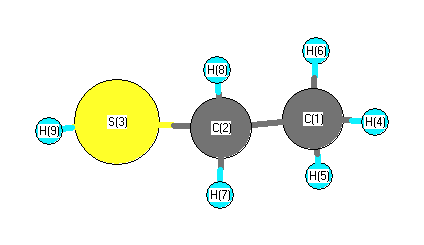 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.109 |
|
C1 |
C2 |
H7 |
110.377 |
| C1 |
C2 |
H8 |
110.377 |
|
C2 |
C1 |
H4 |
110.141 |
| C2 |
C1 |
H5 |
111.247 |
|
C2 |
C1 |
H6 |
111.247 |
| C2 |
S3 |
H9 |
96.923 |
|
S3 |
C2 |
H7 |
109.172 |
| S3 |
C2 |
H8 |
109.172 |
|
H4 |
C1 |
H5 |
107.970 |
| H4 |
C1 |
H6 |
107.970 |
|
H5 |
C1 |
H6 |
108.140 |
| H7 |
C2 |
H8 |
108.608 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/6-311G*
| | hartrees |
|---|
| Energy at 0K | -477.827429 |
| Energy at 298.15K | |
| HF Energy | -477.646482 |
| Nuclear repulsion energy | 106.981730 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3154 |
3154 |
34.88 |
10.60 |
0.75 |
0.86 |
| 2 |
A |
3133 |
3133 |
10.65 |
98.30 |
0.74 |
0.85 |
| 3 |
A |
3122 |
3122 |
27.49 |
108.16 |
0.66 |
0.80 |
| 4 |
A |
3090 |
3090 |
16.28 |
116.62 |
0.14 |
0.25 |
| 5 |
A |
3058 |
3058 |
29.06 |
139.51 |
0.05 |
0.09 |
| 6 |
A |
2648 |
2648 |
14.80 |
129.34 |
0.36 |
0.53 |
| 7 |
A |
1533 |
1533 |
4.23 |
7.82 |
0.75 |
0.86 |
| 8 |
A |
1526 |
1526 |
11.40 |
10.99 |
0.74 |
0.85 |
| 9 |
A |
1511 |
1511 |
1.85 |
15.75 |
0.72 |
0.84 |
| 10 |
A |
1445 |
1445 |
4.93 |
2.09 |
0.67 |
0.80 |
| 11 |
A |
1346 |
1346 |
22.98 |
1.61 |
0.66 |
0.79 |
| 12 |
A |
1310 |
1310 |
4.45 |
6.02 |
0.75 |
0.86 |
| 13 |
A |
1150 |
1150 |
13.65 |
6.55 |
0.73 |
0.85 |
| 14 |
A |
1090 |
1090 |
0.42 |
3.80 |
0.37 |
0.54 |
| 15 |
A |
1002 |
1002 |
7.45 |
6.40 |
0.75 |
0.86 |
| 16 |
A |
890 |
890 |
8.53 |
2.69 |
0.68 |
0.81 |
| 17 |
A |
751 |
751 |
2.66 |
2.80 |
0.35 |
0.52 |
| 18 |
A |
668 |
668 |
4.24 |
15.45 |
0.28 |
0.43 |
| 19 |
A |
333 |
333 |
1.51 |
0.90 |
0.65 |
0.79 |
| 20 |
A |
271 |
271 |
1.38 |
0.53 |
0.59 |
0.74 |
| 21 |
A |
214 |
214 |
20.35 |
8.03 |
0.73 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 16622.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16622.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.641 |
-0.351 |
-0.053 |
| C2 |
0.498 |
0.645 |
0.090 |
| S3 |
-1.166 |
-0.101 |
-0.081 |
| H4 |
2.605 |
0.161 |
0.014 |
| H5 |
1.609 |
-1.105 |
0.736 |
| H6 |
1.590 |
-0.869 |
-1.011 |
| H7 |
0.553 |
1.178 |
1.040 |
| H8 |
0.540 |
1.395 |
-0.701 |
| H9 |
-1.081 |
-0.909 |
0.994 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5223 | 2.8178 | 1.0933 | 1.0919 | 1.0903 | 2.1713 | 2.1636 | 2.9688 |
C2 | 1.5223 | | 1.8318 | 2.1627 | 2.1713 | 2.1669 | 1.0901 | 1.0909 | 2.3928 | S3 | 2.8178 | 1.8318 | | 3.7808 | 3.0623 | 3.0083 | 2.4175 | 2.3516 | 1.3477 | H4 | 1.0933 | 2.1627 | 3.7808 | | 1.7650 | 1.7714 | 2.5098 | 2.5099 | 3.9608 | H5 | 1.0919 | 2.1713 | 3.0623 | 1.7650 | | 1.7634 | 2.5339 | 3.0757 | 2.7094 | H6 | 1.0903 | 2.1669 | 3.0083 | 1.7714 | 1.7634 | | 3.0772 | 2.5145 | 3.3402 | H7 | 2.1713 | 1.0901 | 2.4175 | 2.5098 | 2.5339 | 3.0772 | | 1.7542 | 2.6504 | H8 | 2.1636 | 1.0909 | 2.3516 | 2.5099 | 3.0757 | 2.5145 | 1.7542 | | 3.2873 | H9 | 2.9688 | 2.3928 | 1.3477 | 3.9608 | 2.7094 | 3.3402 | 2.6504 | 3.2873 | |
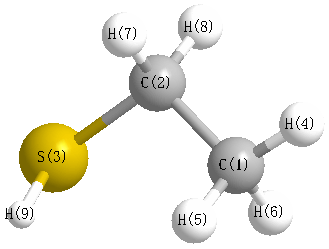 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.988 |
|
C1 |
C2 |
H7 |
111.362 |
| C1 |
C2 |
H8 |
110.695 |
|
C2 |
C1 |
H4 |
110.475 |
| C2 |
C1 |
H5 |
111.248 |
|
C2 |
C1 |
H6 |
110.994 |
| C2 |
S3 |
H9 |
96.442 |
|
S3 |
C2 |
H7 |
109.004 |
| S3 |
C2 |
H8 |
104.260 |
|
H4 |
C1 |
H5 |
107.744 |
| H4 |
C1 |
H6 |
108.434 |
|
H5 |
C1 |
H6 |
107.822 |
| H7 |
C2 |
H8 |
107.092 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
