Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.525688 |
| Energy at 298.15K | |
| HF Energy | -168.972061 |
| Nuclear repulsion energy | 71.142747 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3805 |
3603 |
43.24 |
|
|
|
| 2 |
A' |
3659 |
3465 |
44.18 |
|
|
|
| 3 |
A' |
3014 |
2854 |
113.47 |
|
|
|
| 4 |
A' |
1838 |
1740 |
366.30 |
|
|
|
| 5 |
A' |
1681 |
1592 |
58.15 |
|
|
|
| 6 |
A' |
1442 |
1366 |
4.03 |
|
|
|
| 7 |
A' |
1300 |
1231 |
107.26 |
|
|
|
| 8 |
A' |
1076 |
1019 |
6.03 |
|
|
|
| 9 |
A' |
571 |
541 |
10.99 |
|
|
|
| 10 |
A" |
1053 |
997 |
2.36 |
|
|
|
| 11 |
A" |
654 |
619 |
17.41 |
|
|
|
| 12 |
A" |
216i |
205i |
289.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9937.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 9411.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
1.200 |
0.236 |
0.000 |
| N3 |
-0.941 |
-0.565 |
0.000 |
| H4 |
-0.453 |
1.427 |
0.000 |
| H5 |
-0.641 |
-1.524 |
0.000 |
| H6 |
-1.922 |
-0.357 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2139 | 1.3611 | 1.1054 | 2.0466 | 2.0727 |
O2 | 1.2139 | | 2.2857 | 2.0374 | 2.5478 | 3.1779 | N3 | 1.3611 | 2.2857 | | 2.0509 | 1.0054 | 1.0029 | H4 | 1.1054 | 2.0374 | 2.0509 | | 2.9579 | 2.3114 | H5 | 2.0466 | 2.5478 | 1.0054 | 2.9579 | | 1.7328 | H6 | 2.0727 | 3.1779 | 1.0029 | 2.3114 | 1.7328 | |
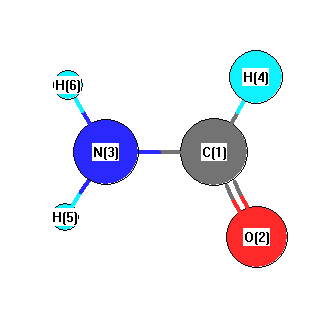 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.960 |
|
C1 |
N3 |
H6 |
121.773 |
| O2 |
C1 |
N3 |
125.060 |
|
O2 |
C1 |
H4 |
122.845 |
| N3 |
C1 |
H4 |
112.096 |
|
H5 |
N3 |
H6 |
119.267 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.525763 |
| Energy at 298.15K | -169.529563 |
| HF Energy | -168.971948 |
| Nuclear repulsion energy | 71.115471 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3790 |
3589 |
38.23 |
|
|
|
| 2 |
A |
3648 |
3455 |
38.58 |
|
|
|
| 3 |
A |
3016 |
2856 |
111.44 |
|
|
|
| 4 |
A |
1836 |
1739 |
349.48 |
|
|
|
| 5 |
A |
1683 |
1594 |
56.32 |
|
|
|
| 6 |
A |
1444 |
1368 |
3.79 |
|
|
|
| 7 |
A |
1302 |
1233 |
103.10 |
|
|
|
| 8 |
A |
1087 |
1030 |
7.99 |
|
|
|
| 9 |
A |
1050 |
994 |
1.59 |
|
|
|
| 10 |
A |
647 |
613 |
35.76 |
|
|
|
| 11 |
A |
573 |
542 |
10.69 |
|
|
|
| 12 |
A |
293 |
277 |
294.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10184.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 9645.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.162 |
0.387 |
0.002 |
| O2 |
1.197 |
-0.246 |
0.006 |
| N3 |
-1.089 |
-0.158 |
-0.041 |
| H4 |
0.136 |
1.492 |
-0.002 |
| H5 |
-1.168 |
-1.154 |
0.082 |
| H6 |
-1.896 |
0.409 |
0.146 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2137 | 1.3644 | 1.1053 | 2.0367 | 2.0626 |
O2 | 1.2137 | | 2.2879 | 2.0362 | 2.5346 | 3.1647 | N3 | 1.3644 | 2.2879 | | 2.0552 | 1.0063 | 1.0040 | H4 | 1.1053 | 2.0362 | 2.0552 | | 2.9509 | 2.3075 | H5 | 2.0367 | 2.5346 | 1.0063 | 2.9509 | | 1.7249 | H6 | 2.0626 | 3.1647 | 1.0040 | 2.3075 | 1.7249 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.631 |
|
C1 |
N3 |
H6 |
120.360 |
| O2 |
C1 |
N3 |
125.001 |
|
O2 |
C1 |
H4 |
122.747 |
| N3 |
C1 |
H4 |
112.222 |
|
H5 |
N3 |
H6 |
118.191 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
