Jump to
S1C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.759249 |
| Energy at 298.15K | -169.762954 |
| HF Energy | -169.759249 |
| Nuclear repulsion energy | 71.436444 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3769 |
3617 |
30.99 |
|
|
|
| 2 |
A' |
3629 |
3483 |
26.04 |
|
|
|
| 3 |
A' |
2958 |
2839 |
126.86 |
|
|
|
| 4 |
A' |
1857 |
1782 |
411.14 |
|
|
|
| 5 |
A' |
1659 |
1592 |
71.75 |
|
|
|
| 6 |
A' |
1433 |
1375 |
5.19 |
|
|
|
| 7 |
A' |
1293 |
1241 |
99.77 |
|
|
|
| 8 |
A' |
1074 |
1031 |
3.42 |
|
|
|
| 9 |
A' |
570 |
547 |
11.87 |
|
|
|
| 10 |
A" |
1049 |
1007 |
0.57 |
|
|
|
| 11 |
A" |
664 |
637 |
22.82 |
|
|
|
| 12 |
A" |
219 |
210 |
269.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10086.7 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 9679.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| O2 |
1.193 |
0.241 |
0.000 |
| N3 |
-0.933 |
-0.566 |
0.000 |
| H4 |
-0.459 |
1.425 |
0.000 |
| H5 |
-0.635 |
-1.527 |
0.000 |
| H6 |
-1.916 |
-0.361 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2057 | 1.3548 | 1.1082 | 2.0438 | 2.0677 |
O2 | 1.2057 | | 2.2745 | 2.0319 | 2.5430 | 3.1669 | N3 | 1.3548 | 2.2745 | | 2.0464 | 1.0063 | 1.0037 | H4 | 1.1082 | 2.0319 | 2.0464 | | 2.9567 | 2.3051 | H5 | 2.0438 | 2.5430 | 1.0063 | 2.9567 | | 1.7321 | H6 | 2.0677 | 3.1669 | 1.0037 | 2.3051 | 1.7321 | |
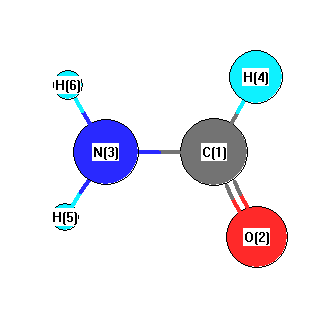 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.180 |
|
C1 |
N3 |
H6 |
121.792 |
| O2 |
C1 |
N3 |
125.227 |
|
O2 |
C1 |
H4 |
122.783 |
| N3 |
C1 |
H4 |
111.991 |
|
H5 |
N3 |
H6 |
119.028 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.225 |
|
|
|
| 2 |
O |
-0.359 |
|
|
|
| 3 |
N |
-0.744 |
|
|
|
| 4 |
H |
0.153 |
|
|
|
| 5 |
H |
0.362 |
|
|
|
| 6 |
H |
0.363 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.850 |
-0.776 |
0.000 |
3.928 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.822 |
-0.087 |
0.000 |
| y |
-0.087 |
-14.620 |
0.000 |
| z |
0.000 |
0.000 |
-18.704 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.159 |
-0.087 |
0.000 |
| y |
-0.087 |
3.642 |
0.000 |
| z |
0.000 |
0.000 |
-2.483 |
|
| Polar |
|---|
| 3z2-r2 | -4.966 |
|---|
| x2-y2 | -3.201 |
|---|
| xy | -0.087 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.191 |
0.257 |
0.000 |
| y |
0.257 |
3.358 |
0.000 |
| z |
0.000 |
0.000 |
1.718 |
<r2> (average value of r
2) Å
2
| <r2> |
40.655 |
| (<r2>)1/2 |
6.376 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.759248 |
| Energy at 298.15K | -169.762948 |
| HF Energy | -169.759248 |
| Nuclear repulsion energy | 71.436690 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3769 |
3617 |
31.01 |
|
|
|
| 2 |
A |
3629 |
3483 |
26.04 |
|
|
|
| 3 |
A |
2959 |
2840 |
126.79 |
|
|
|
| 4 |
A |
1857 |
1782 |
411.16 |
|
|
|
| 5 |
A |
1658 |
1591 |
71.62 |
|
|
|
| 6 |
A |
1433 |
1375 |
5.18 |
|
|
|
| 7 |
A |
1293 |
1241 |
99.65 |
|
|
|
| 8 |
A |
1074 |
1031 |
3.38 |
|
|
|
| 9 |
A |
1049 |
1007 |
0.58 |
|
|
|
| 10 |
A |
664 |
637 |
23.12 |
|
|
|
| 11 |
A |
570 |
547 |
11.89 |
|
|
|
| 12 |
A |
215 |
206 |
269.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10085.5 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 9678.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.162 |
0.383 |
-0.000 |
| O2 |
1.192 |
-0.244 |
-0.000 |
| N3 |
-1.080 |
-0.156 |
0.000 |
| H4 |
0.135 |
1.491 |
0.000 |
| H5 |
-1.181 |
-1.157 |
0.000 |
| H6 |
-1.905 |
0.416 |
-0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2058 | 1.3546 | 1.1081 | 2.0435 | 2.0678 |
O2 | 1.2058 | | 2.2745 | 2.0318 | 2.5426 | 3.1670 | N3 | 1.3546 | 2.2745 | | 2.0465 | 1.0063 | 1.0037 | H4 | 1.1081 | 2.0318 | 2.0465 | | 2.9567 | 2.3056 | H5 | 2.0435 | 2.5426 | 1.0063 | 2.9567 | | 1.7321 | H6 | 2.0678 | 3.1670 | 1.0037 | 2.3056 | 1.7321 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.162 |
|
C1 |
N3 |
H6 |
121.807 |
| O2 |
C1 |
N3 |
125.222 |
|
O2 |
C1 |
H4 |
122.765 |
| N3 |
C1 |
H4 |
112.013 |
|
H5 |
N3 |
H6 |
119.031 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.225 |
|
|
|
| 2 |
O |
-0.359 |
|
|
|
| 3 |
N |
-0.744 |
|
|
|
| 4 |
H |
0.153 |
|
|
|
| 5 |
H |
0.362 |
|
|
|
| 6 |
H |
0.363 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.848 |
0.790 |
0.000 |
3.929 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.395 |
1.092 |
0.001 |
| y |
1.092 |
-15.047 |
-0.001 |
| z |
0.001 |
-0.001 |
-18.704 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.519 |
1.092 |
0.001 |
| y |
1.092 |
3.003 |
-0.001 |
| z |
0.001 |
-0.001 |
-2.483 |
|
| Polar |
|---|
| 3z2-r2 | -4.966 |
|---|
| x2-y2 | -2.348 |
|---|
| xy | 1.092 |
|---|
| xz | 0.001 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.248 |
-0.121 |
0.000 |
| y |
-0.121 |
3.301 |
0.000 |
| z |
0.000 |
0.000 |
1.717 |
<r2> (average value of r
2) Å
2
| <r2> |
40.654 |
| (<r2>)1/2 |
6.376 |
