return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

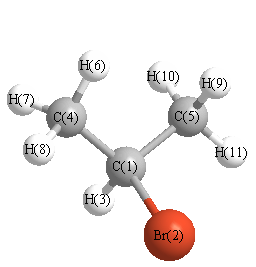
All results from a given calculation for CH3CHBrCH3 (i-propyl bromide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-2691.073594
Energy at 298.15K-2691.084202
HF Energy-2690.077081
Nuclear repulsion energy251.844731
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3193 3024 19.51      
2 A' 3170 3003 30.42      
3 A' 3142 2976 1.76      
4 A' 3080 2917 22.09      
5 A' 1545 1464 12.81      
6 A' 1533 1452 9.02      
7 A' 1458 1381 9.01      
8 A' 1309 1240 47.42      
9 A' 1217 1152 19.17      
10 A' 1091 1034 11.49      
11 A' 926 877 5.52      
12 A' 583 552 13.10      
13 A' 417 395 1.20      
14 A' 318 301 0.95      
15 A' 284 269 0.67      
16 A" 3190 3021 9.39      
17 A" 3165 2998 1.55      
18 A" 3077 2914 11.29      
19 A" 1522 1442 0.07      
20 A" 1519 1439 4.59      
21 A" 1443 1367 16.64      
22 A" 1383 1310 0.31      
23 A" 1181 1118 0.80      
24 A" 981 929 0.03      
25 A" 960 909 1.95      
26 A" 299 283 0.41      
27 A" 264 250 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 21124.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20007.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.26941 0.09757 0.07682

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.904 -0.424 0.000
Br2 -0.671 0.749 0.000
H3 1.749 0.267 0.000
C4 0.904 -1.259 1.266
C5 0.904 -1.259 -1.266
H6 0.025 -1.906 1.300
H7 1.799 -1.889 1.285
H8 0.904 -0.632 2.159
H9 0.025 -1.906 -1.300
H10 1.799 -1.889 -1.285
H11 0.904 -0.632 -2.159

Atom - Atom Distances (Å)
  C1 Br2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.96351.09161.51711.51712.15892.14462.16852.15892.14462.1685
Br21.96352.46772.84892.84893.03713.83553.00803.03713.83553.0080
H31.09162.46772.15562.15563.06362.51022.48613.06362.51022.4861
C41.51712.84892.15562.53271.09201.09471.09062.78922.77623.4819
C51.51712.84892.15562.53272.78922.77623.48191.09201.09471.0906
H62.15893.03713.06361.09202.78921.77431.77022.60083.13563.7897
H72.14463.83552.51021.09472.77621.77431.77323.13562.56983.7734
H82.16853.00802.48611.09063.48191.77021.77323.78973.77344.3171
H92.15893.03713.06362.78921.09202.60083.13563.78971.77431.7702
H102.14463.83552.51022.77621.09473.13562.56983.77341.77431.7732
H112.16853.00802.48613.48191.09063.78973.77344.31711.77021.7732

picture of i-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.619 C1 C4 H7 109.333
C1 C4 H8 111.479 C1 C5 H9 110.619
C1 C5 H10 109.333 C1 C5 H11 111.479
Br2 C1 H3 104.078 Br2 C1 C4 109.204
Br2 C1 C5 109.204 H3 C1 C4 110.383
H3 C1 C5 110.383 C4 C1 C5 113.181
H6 C4 H7 108.471 H6 C4 H8 108.397
H7 C4 H8 108.467 H9 C5 H10 108.471
H9 C5 H11 108.397 H10 C5 H11 108.467
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability