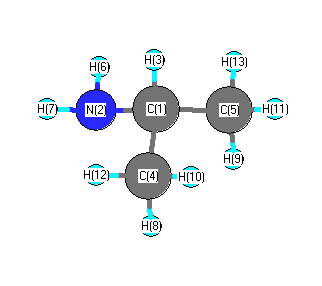
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3737 |
3379 |
1.58 |
|
|
|
| 2 |
A' |
3252 |
2941 |
109.24 |
|
|
|
| 3 |
A' |
3229 |
2920 |
93.39 |
|
|
|
| 4 |
A' |
3177 |
2873 |
4.89 |
|
|
|
| 5 |
A' |
3098 |
2802 |
94.70 |
|
|
|
| 6 |
A' |
1857 |
1679 |
47.53 |
|
|
|
| 7 |
A' |
1643 |
1486 |
6.42 |
|
|
|
| 8 |
A' |
1630 |
1474 |
3.19 |
|
|
|
| 9 |
A' |
1556 |
1407 |
14.70 |
|
|
|
| 10 |
A' |
1512 |
1367 |
18.56 |
|
|
|
| 11 |
A' |
1307 |
1182 |
5.79 |
|
|
|
| 12 |
A' |
1273 |
1151 |
29.74 |
|
|
|
| 13 |
A' |
1072 |
970 |
4.54 |
|
|
|
| 14 |
A' |
964 |
872 |
148.75 |
|
|
|
| 15 |
A' |
871 |
788 |
0.41 |
|
|
|
| 16 |
A' |
508 |
460 |
11.32 |
|
|
|
| 17 |
A' |
392 |
355 |
0.13 |
|
|
|
| 18 |
A' |
288 |
261 |
0.19 |
|
|
|
| 19 |
A" |
3807 |
3443 |
0.42 |
|
|
|
| 20 |
A" |
3243 |
2933 |
1.57 |
|
|
|
| 21 |
A" |
3224 |
2915 |
30.74 |
|
|
|
| 22 |
A" |
3170 |
2867 |
69.41 |
|
|
|
| 23 |
A" |
1623 |
1468 |
0.09 |
|
|
|
| 24 |
A" |
1620 |
1465 |
0.10 |
|
|
|
| 25 |
A" |
1562 |
1413 |
16.60 |
|
|
|
| 26 |
A" |
1515 |
1371 |
0.30 |
|
|
|
| 27 |
A" |
1375 |
1243 |
0.06 |
|
|
|
| 28 |
A" |
1118 |
1011 |
1.47 |
|
|
|
| 29 |
A" |
1037 |
938 |
0.06 |
|
|
|
| 30 |
A" |
1009 |
912 |
0.00 |
|
|
|
| 31 |
A" |
431 |
389 |
5.79 |
|
|
|
| 32 |
A" |
313 |
283 |
45.67 |
|
|
|
| 33 |
A" |
237 |
214 |
8.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28324.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 25616.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.043 |
0.739 |
|
0.707 |
| 2 |
N |
-0.825 |
-1.096 |
|
-1.072 |
| 3 |
H |
0.172 |
-0.150 |
|
-0.103 |
| 4 |
C |
-0.600 |
-0.282 |
|
-0.477 |
| 5 |
C |
-0.600 |
-0.282 |
|
-0.477 |
| 6 |
H |
0.330 |
0.372 |
|
0.373 |
| 7 |
H |
0.330 |
0.372 |
|
0.373 |
| 8 |
H |
0.217 |
0.070 |
|
0.131 |
| 9 |
H |
0.217 |
0.070 |
|
0.131 |
| 10 |
H |
0.203 |
0.037 |
|
0.097 |
| 11 |
H |
0.203 |
0.037 |
|
0.097 |
| 12 |
H |
0.198 |
0.057 |
|
0.110 |
| 13 |
H |
0.198 |
0.057 |
|
0.110 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.123 |
1.425 |
0.000 |
1.430 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.090 |
1.420 |
0.000 |
1.423 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.342 |
-2.455 |
0.000 |
| y |
-2.455 |
-28.867 |
0.000 |
| z |
0.000 |
0.000 |
-26.758 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.470 |
-2.455 |
0.000 |
| y |
-2.455 |
-2.317 |
0.000 |
| z |
0.000 |
0.000 |
0.847 |
|
| Polar |
|---|
| 3z2-r2 | 1.694 |
|---|
| x2-y2 | 2.525 |
|---|
| xy | -2.455 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.220 |
-0.038 |
0.000 |
| y |
-0.038 |
5.408 |
0.000 |
| z |
0.000 |
0.000 |
6.399 |
<r2> (average value of r
2) Å
2
| <r2> |
93.018 |
| (<r2>)1/2 |
9.645 |