return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHSHCH3 (2-Propanethiol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-516.540986
Energy at 298.15K-516.549702
HF Energy-515.814287
Nuclear repulsion energy170.483691
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3185 3017 18.87      
2 A' 3160 2993 54.42      
3 A' 3104 2939 4.33      
4 A' 3075 2912 25.92      
5 A' 2665 2524 9.32      
6 A' 1543 1462 16.64      
7 A' 1538 1456 9.59      
8 A' 1458 1381 8.13      
9 A' 1351 1280 11.61      
10 A' 1224 1160 1.86      
11 A' 1145 1084 27.49      
12 A' 939 889 0.98      
13 A' 895 847 11.03      
14 A' 659 624 3.71      
15 A' 420 398 0.83      
16 A' 354 335 0.30      
17 A' 291 276 0.20      
18 A" 3183 3015 11.51      
19 A" 3154 2987 0.18      
20 A" 3071 2909 15.54      
21 A" 1525 1444 4.26      
22 A" 1520 1440 0.11      
23 A" 1440 1364 12.07      
24 A" 1359 1287 0.22      
25 A" 1173 1111 0.63      
26 A" 983 931 0.00      
27 A" 954 904 2.38      
28 A" 343 325 2.40      
29 A" 260 246 0.00      
30 A" 201 190 20.11      

Unscaled Zero Point Vibrational Energy (zpe) 23086.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 21864.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.26453 0.14825 0.10616

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.562 0.000
S2 -0.987 -0.969 0.000
H3 -0.760 1.351 0.000
H4 0.070 -1.804 0.000
C5 0.842 0.689 1.263
C6 0.842 0.689 -1.263
H7 1.588 -0.110 1.307
H8 1.588 -0.110 -1.307
H9 1.373 1.646 1.275
H10 0.223 0.622 2.159
H11 0.223 0.622 -2.159
H12 1.373 1.646 -1.275

Atom - Atom Distances (Å)
  C1 S2 H3 H4 C5 C6 H7 H8 H9 H10 H11 H12
C11.82181.09542.36711.52351.52352.16422.16422.16422.17102.17102.1642
S21.82182.33151.34662.77302.77303.01273.01273.74572.94182.94183.7457
H31.09542.33153.26242.14512.14513.05923.05922.50202.48172.48172.5020
H42.36711.34663.26242.89912.89912.62322.62323.90153.25053.25053.9015
C51.52352.77302.14512.89912.52631.09392.79331.09411.09083.47802.7640
C61.52352.77302.14512.89912.52632.79331.09392.76403.47801.09081.0941
H72.16423.01273.05922.62321.09392.79332.61491.76961.76723.79643.1305
H82.16423.01273.05922.62322.79331.09392.61493.13053.79641.76721.7696
H92.16423.74572.50203.90151.09412.76401.76963.13051.77503.76312.5502
H102.17102.94182.48173.25051.09083.47801.76723.79641.77504.31733.7631
H112.17102.94182.48173.25053.47801.09083.79641.76723.76314.31731.7750
H122.16423.74572.50203.90152.76401.09413.13051.76962.55023.76311.7750

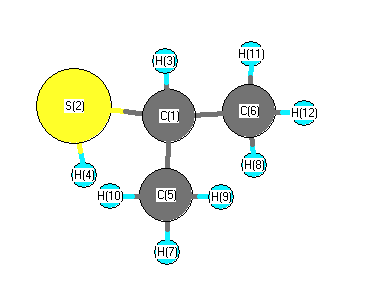
picture of 2-Propanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H4 95.510 C1 C5 H7 110.477
C1 C5 H9 110.464 C1 C5 H10 111.211
C1 C6 H8 110.477 C1 C6 H11 111.211
C1 C6 H12 110.464 S2 C1 H3 103.283
S2 C1 C5 111.667 S2 C1 C6 111.667
H3 C1 C5 108.891 H3 C1 C6 108.891
C5 C1 C6 112.016 H7 C5 H9 107.945
H7 C5 H10 107.980 H8 C6 H11 107.980
H8 C6 H12 107.945 H9 C5 H10 108.661
H11 C6 H12 108.661
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability