return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H14 (Butane, 2,2-dimethyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-236.365761
Energy at 298.15K-236.381068
HF Energy-235.404625
Nuclear repulsion energy266.727544
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3168 3001 43.02      
2 A' 3161 2994 49.48      
3 A' 3153 2986 64.69      
4 A' 3147 2981 1.71      
5 A' 3085 2921 19.40      
6 A' 3068 2906 8.84      
7 A' 3059 2897 39.24      
8 A' 3055 2893 21.73      
9 A' 1560 1478 18.63      
10 A' 1549 1467 11.19      
11 A' 1545 1464 7.86      
12 A' 1526 1445 0.05      
13 A' 1516 1436 0.26      
14 A' 1464 1386 6.28      
15 A' 1446 1370 12.14      
16 A' 1433 1357 12.18      
17 A' 1397 1323 0.84      
18 A' 1317 1247 1.69      
19 A' 1273 1205 4.60      
20 A' 1121 1062 1.54      
21 A' 1066 1009 2.67      
22 A' 1033 979 3.70      
23 A' 970 919 1.18      
24 A' 910 862 0.29      
25 A' 735 696 0.39      
26 A' 493 467 0.41      
27 A' 418 396 0.06      
28 A' 369 349 0.01      
29 A' 295 279 0.04      
30 A' 262 248 0.04      
31 A" 3179 3011 26.47      
32 A" 3159 2992 22.68      
33 A" 3151 2984 31.78      
34 A" 3147 2981 1.99      
35 A" 3104 2940 18.32      
36 A" 3062 2900 33.20      
37 A" 1554 1472 0.23      
38 A" 1543 1461 17.78      
39 A" 1521 1441 0.15      
40 A" 1515 1435 0.07      
41 A" 1434 1358 16.80      
42 A" 1371 1299 0.14      
43 A" 1279 1212 0.98      
44 A" 1127 1067 0.12      
45 A" 1024 970 0.31      
46 A" 977 926 0.11      
47 A" 971 920 0.41      
48 A" 795 753 2.72      
49 A" 429 406 0.05      
50 A" 349 331 0.01      
51 A" 305 289 0.00      
52 A" 267 253 0.00      
53 A" 211 200 0.00      
54 A" 68 65 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 42068.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 39843.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14427 0.08381 0.08353

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.036 -2.172 0.000
C2 0.826 -0.865 0.000
C3 0.000 0.432 0.000
C4 0.973 1.613 0.000
C5 -0.879 0.516 1.249
C6 -0.879 0.516 -1.249
H7 0.434 2.565 0.000
H8 0.719 -3.025 0.000
H9 -0.277 0.418 2.158
H10 -0.277 0.418 -2.158
H11 -1.642 -0.266 1.262
H12 -1.642 -0.266 -1.262
H13 -1.395 1.480 1.290
H14 -1.395 1.480 -1.290
H15 -0.598 -2.264 0.884
H16 -0.598 -2.264 -0.884
H17 1.486 -0.846 0.877
H18 1.486 -0.846 -0.877
H19 1.617 1.589 0.885
H20 1.617 1.589 -0.885

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.52722.60443.89883.10163.10164.75381.09313.38623.38622.83562.83564.12904.12901.09201.09202.15182.15184.17504.1750
C21.52721.53782.48192.52432.52433.45242.16282.74262.74262.83602.83603.47793.47792.18352.18351.09781.09782.72642.7264
C32.60441.53781.52991.52931.52932.17673.53142.17612.17612.18562.18562.16972.16972.90002.90002.14742.14742.17652.1765
C43.89882.48191.52992.48862.48861.09464.64462.76552.76553.45883.45882.70012.70014.27554.27552.66012.66011.09471.0947
C53.10162.52431.52932.48862.49762.73574.08061.09503.46131.09292.73811.09462.76432.81763.51502.75413.45912.74103.4548
C63.10162.52431.52932.48862.49762.73574.08063.46131.09502.73811.09292.76431.09463.51502.81763.45912.75413.45482.7410
H74.75383.45242.17671.09462.73572.73575.59763.12623.12623.73113.73112.48752.48755.01705.01703.67573.67571.77031.7703
H81.09312.16283.53144.64464.08064.08065.59764.18444.18443.84463.84465.14115.14111.75981.75982.47112.47114.78384.7838
H93.38622.74262.17612.76551.09503.46133.12624.18444.31671.77113.74571.76953.77732.98714.06892.51973.73102.56483.7712
H103.38622.74262.17612.76553.46131.09503.12624.18444.31673.74571.77113.77731.76954.06892.98713.73102.51973.77122.5648
H112.83562.83602.18563.45881.09292.73813.73113.84461.77113.74572.52361.76393.10192.28563.11233.20483.83373.76924.3215
H122.83562.83602.18563.45882.73811.09293.73113.84463.74571.77112.52363.10191.76393.11232.28563.83373.20484.32153.7692
H134.12903.47792.16972.70011.09462.76432.48755.14111.76953.77731.76393.10192.57993.84944.40233.72574.29023.04083.7168
H144.12903.47792.16972.70012.76431.09462.48755.14113.77731.76953.10191.76392.57994.40233.84944.29023.72573.71683.0408
H151.09202.18352.90004.27552.81763.51505.01701.75982.98714.06892.28563.11233.84944.40231.76832.52113.07534.44484.7841
H161.09202.18352.90004.27553.51502.81765.01701.75984.06892.98713.11232.28564.40233.84941.76833.07532.52114.78414.4448
H172.15181.09782.14742.66012.75413.45913.67572.47112.51973.73103.20483.83373.72574.29022.52113.07531.75402.43893.0090
H182.15181.09782.14742.66013.45912.75413.67572.47113.73102.51973.83373.20484.29023.72573.07532.52111.75403.00902.4389
H194.17502.72642.17651.09472.74103.45481.77034.78382.56483.77123.76924.32153.04083.71684.44484.78412.43893.00901.7707
H204.17502.72642.17651.09473.45482.74101.77034.78383.77122.56484.32153.76923.71683.04084.78414.44483.00902.43891.7707

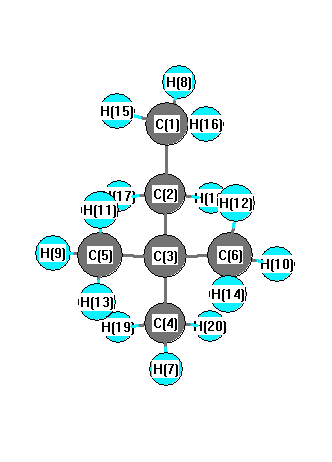
picture of Butane, 2,2-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 116.364 C1 C2 H17 109.018
C1 C2 H18 109.018 C2 C1 H8 110.161
C2 C1 H15 111.881 C2 C1 H16 111.881
C2 C3 C4 108.002 C2 C3 C5 110.773
C2 C3 C6 110.773 C3 C2 H17 107.959
C3 C2 H18 107.959 C3 C4 H7 110.985
C3 C4 H19 110.961 C3 C4 H20 110.961
C3 C5 H9 110.952 C3 C5 H11 111.846
C3 C5 H13 110.473 C3 C6 H10 110.952
C3 C6 H12 111.846 C3 C6 H14 110.473
C4 C3 C5 108.876 C4 C3 C6 108.876
C5 C3 C6 109.487 H7 C4 H19 107.926
H7 C4 H20 107.926 H8 C1 H15 107.289
H8 C1 H16 107.289 H9 C5 H11 108.090
H9 C5 H13 107.834 H10 C6 H12 108.090
H10 C6 H14 107.834 H11 C5 H13 107.485
H12 C6 H14 107.485 H15 C1 H16 108.123
H17 C2 H18 106.039 H19 C4 H20 107.947
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability