return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CF3CHO (trifluoroacetaldehyde)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-450.777698
Energy at 298.15K-450.780161
HF Energy-449.600513
Nuclear repulsion energy252.571702
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3042 2881 51.91      
2 A' 1808 1712 50.28      
3 A' 1431 1355 36.77      
4 A' 1355 1284 154.92      
5 A' 1242 1177 223.81      
6 A' 863 817 40.82      
7 A' 718 680 34.93      
8 A' 539 511 16.77      
9 A' 440 417 4.63      
10 A' 260 246 5.99      
11 A" 1223 1158 310.67      
12 A" 989 937 10.89      
13 A" 541 512 1.59      
14 A" 322 305 0.83      
15 A" 74 70 12.32      

Unscaled Zero Point Vibrational Energy (zpe) 7423.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 7031.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18258 0.09954 0.09786

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.020 0.361 0.000
C2 0.502 -1.093 0.000
O3 -0.257 -2.029 0.000
F4 -1.301 0.453 0.000
F5 0.502 0.986 1.085
F6 0.502 0.986 -1.085
H7 1.601 -1.196 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 F4 F5 F6 H7
C11.53202.40581.32451.34141.34142.2188
C21.53201.20492.37572.34512.34511.1040
O32.40581.20492.69283.29273.29272.0367
F41.32452.37572.69282.17072.17073.3382
F51.34142.34513.29272.17072.16932.6728
F61.34142.34513.29272.17072.16932.6728
H72.21881.10402.03673.33822.67282.6728

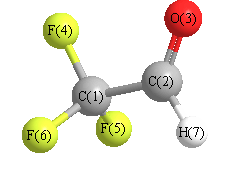
picture of trifluoroacetaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 122.602 C1 C2 H7 113.659
C2 C1 F4 112.341 C2 C1 F5 109.221
C2 C1 F6 109.221 O3 C2 H7 123.740
F4 C1 F5 109.021 F4 C1 F6 109.021
F5 C1 F6 107.914
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability