Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -637.765409 |
| Energy at 298.15K | -637.770380 |
| HF Energy | -637.045609 |
| Nuclear repulsion energy | 157.482002 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
2981 |
18.86 |
|
|
|
| 2 |
A' |
3125 |
2960 |
17.31 |
|
|
|
| 3 |
A' |
1563 |
1481 |
1.69 |
|
|
|
| 4 |
A' |
1529 |
1449 |
5.84 |
|
|
|
| 5 |
A' |
1470 |
1392 |
4.46 |
|
|
|
| 6 |
A' |
1343 |
1272 |
14.56 |
|
|
|
| 7 |
A' |
1118 |
1058 |
23.93 |
|
|
|
| 8 |
A' |
1091 |
1033 |
87.86 |
|
|
|
| 9 |
A' |
826 |
783 |
40.36 |
|
|
|
| 10 |
A' |
396 |
375 |
2.98 |
|
|
|
| 11 |
A' |
250 |
236 |
11.70 |
|
|
|
| 12 |
A" |
3224 |
3053 |
12.85 |
|
|
|
| 13 |
A" |
3191 |
3022 |
13.78 |
|
|
|
| 14 |
A" |
1347 |
1276 |
0.27 |
|
|
|
| 15 |
A" |
1263 |
1196 |
0.69 |
|
|
|
| 16 |
A" |
1113 |
1054 |
5.26 |
|
|
|
| 17 |
A" |
820 |
776 |
0.78 |
|
|
|
| 18 |
A" |
144 |
137 |
10.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13480.2 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12767.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.613 |
0.000 |
| C2 |
0.980 |
-0.539 |
0.000 |
| Cl3 |
-1.661 |
-0.028 |
0.000 |
| F4 |
2.263 |
-0.011 |
0.000 |
| H5 |
0.131 |
1.226 |
0.890 |
| H6 |
0.131 |
1.226 |
-0.890 |
| H7 |
0.860 |
-1.155 |
0.892 |
| H8 |
0.860 |
-1.155 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5124 | 1.7801 | 2.3474 | 1.0886 | 1.0886 | 2.1591 | 2.1591 |
C2 | 1.5124 | | 2.6898 | 1.3869 | 2.1514 | 2.1514 | 1.0912 | 1.0912 | Cl3 | 1.7801 | 2.6898 | | 3.9236 | 2.3612 | 2.3612 | 2.9015 | 2.9015 | F4 | 2.3474 | 1.3869 | 3.9236 | | 2.6209 | 2.6209 | 2.0183 | 2.0183 | H5 | 1.0886 | 2.1514 | 2.3612 | 2.6209 | | 1.7795 | 2.4906 | 3.0624 | H6 | 1.0886 | 2.1514 | 2.3612 | 2.6209 | 1.7795 | | 3.0624 | 2.4906 | H7 | 2.1591 | 1.0912 | 2.9015 | 2.0183 | 2.4906 | 3.0624 | | 1.7845 | H8 | 2.1591 | 1.0912 | 2.9015 | 2.0183 | 3.0624 | 2.4906 | 1.7845 | |
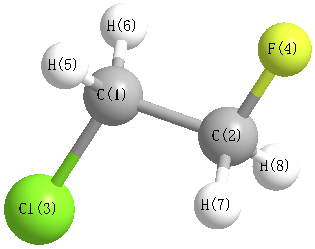 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.048 |
|
C1 |
C2 |
H7 |
111.014 |
| C1 |
C2 |
H8 |
111.014 |
|
C2 |
C1 |
Cl3 |
109.294 |
| C2 |
C1 |
H5 |
110.557 |
|
C2 |
C1 |
H6 |
110.557 |
| Cl3 |
C1 |
H5 |
108.367 |
|
Cl3 |
C1 |
H6 |
108.367 |
| F4 |
C2 |
H7 |
108.484 |
|
F4 |
C2 |
H8 |
108.484 |
| H5 |
C1 |
H6 |
109.639 |
|
H7 |
C2 |
H8 |
109.712 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -637.764599 |
| Energy at 298.15K | -637.769678 |
| HF Energy | -637.044352 |
| Nuclear repulsion energy | 161.579913 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3206 |
3037 |
7.44 |
|
|
|
| 2 |
A |
3175 |
3007 |
21.40 |
|
|
|
| 3 |
A |
3133 |
2967 |
17.02 |
|
|
|
| 4 |
A |
3102 |
2938 |
27.00 |
|
|
|
| 5 |
A |
1547 |
1466 |
2.65 |
|
|
|
| 6 |
A |
1502 |
1423 |
11.06 |
|
|
|
| 7 |
A |
1471 |
1393 |
13.33 |
|
|
|
| 8 |
A |
1393 |
1319 |
29.57 |
|
|
|
| 9 |
A |
1317 |
1248 |
0.94 |
|
|
|
| 10 |
A |
1263 |
1196 |
0.82 |
|
|
|
| 11 |
A |
1147 |
1087 |
48.21 |
|
|
|
| 12 |
A |
1106 |
1047 |
39.27 |
|
|
|
| 13 |
A |
1013 |
959 |
6.23 |
|
|
|
| 14 |
A |
881 |
834 |
9.27 |
|
|
|
| 15 |
A |
727 |
688 |
25.65 |
|
|
|
| 16 |
A |
480 |
454 |
12.22 |
|
|
|
| 17 |
A |
298 |
282 |
1.03 |
|
|
|
| 18 |
A |
145 |
137 |
2.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13452.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12740.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.092 |
0.841 |
-0.295 |
| C2 |
1.194 |
0.414 |
0.362 |
| Cl3 |
-1.423 |
-0.285 |
0.065 |
| F4 |
1.658 |
-0.758 |
-0.203 |
| H5 |
-0.383 |
1.828 |
0.067 |
| H6 |
0.034 |
0.871 |
-1.377 |
| H7 |
1.952 |
1.188 |
0.206 |
| H8 |
1.053 |
0.245 |
1.432 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5063 | 1.7804 | 2.3723 | 1.0907 | 1.0894 | 2.1328 | 2.1559 |
C2 | 1.5063 | | 2.7253 | 1.3813 | 2.1382 | 2.1405 | 1.0944 | 1.0917 | Cl3 | 1.7804 | 2.7253 | | 3.1289 | 2.3554 | 2.3533 | 3.6852 | 2.8774 | F4 | 2.3723 | 1.3813 | 3.1289 | | 3.3050 | 2.5830 | 2.0096 | 2.0111 | H5 | 1.0907 | 2.1382 | 2.3554 | 3.3050 | | 1.7816 | 2.4245 | 2.5351 | H6 | 1.0894 | 2.1405 | 2.3533 | 2.5830 | 1.7816 | | 2.5072 | 3.0529 | H7 | 2.1328 | 1.0944 | 3.6852 | 2.0096 | 2.4245 | 2.5072 | | 1.7889 | H8 | 2.1559 | 1.0917 | 2.8774 | 2.0111 | 2.5351 | 3.0529 | 1.7889 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.405 |
|
C1 |
C2 |
H7 |
109.157 |
| C1 |
C2 |
H8 |
111.163 |
|
C2 |
C1 |
Cl3 |
111.758 |
| C2 |
C1 |
H5 |
109.805 |
|
C2 |
C1 |
H6 |
110.069 |
| Cl3 |
C1 |
H5 |
107.813 |
|
Cl3 |
C1 |
H6 |
107.726 |
| F4 |
C2 |
H7 |
107.971 |
|
F4 |
C2 |
H8 |
108.255 |
| H5 |
C1 |
H6 |
109.607 |
|
H7 |
C2 |
H8 |
109.830 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
