Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -638.428809 |
| Energy at 298.15K | -638.433716 |
| HF Energy | -638.197393 |
| Nuclear repulsion energy | 156.736725 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3132 |
3132 |
20.83 |
|
|
|
| 2 |
A' |
3101 |
3101 |
20.79 |
|
|
|
| 3 |
A' |
1552 |
1552 |
1.55 |
|
|
|
| 4 |
A' |
1523 |
1523 |
4.95 |
|
|
|
| 5 |
A' |
1454 |
1454 |
4.02 |
|
|
|
| 6 |
A' |
1324 |
1324 |
16.95 |
|
|
|
| 7 |
A' |
1093 |
1093 |
11.87 |
|
|
|
| 8 |
A' |
1061 |
1061 |
107.92 |
|
|
|
| 9 |
A' |
780 |
780 |
49.23 |
|
|
|
| 10 |
A' |
385 |
385 |
2.73 |
|
|
|
| 11 |
A' |
244 |
244 |
12.20 |
|
|
|
| 12 |
A" |
3200 |
3200 |
15.25 |
|
|
|
| 13 |
A" |
3158 |
3158 |
17.85 |
|
|
|
| 14 |
A" |
1332 |
1332 |
0.23 |
|
|
|
| 15 |
A" |
1247 |
1247 |
0.83 |
|
|
|
| 16 |
A" |
1094 |
1094 |
5.01 |
|
|
|
| 17 |
A" |
813 |
813 |
0.83 |
|
|
|
| 18 |
A" |
136 |
136 |
10.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13313.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13313.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.611 |
0.000 |
| C2 |
0.996 |
-0.529 |
0.000 |
| Cl3 |
-1.673 |
-0.051 |
0.000 |
| F4 |
2.272 |
0.024 |
0.000 |
| H5 |
0.114 |
1.227 |
0.889 |
| H6 |
0.114 |
1.227 |
-0.889 |
| H7 |
0.889 |
-1.147 |
0.892 |
| H8 |
0.889 |
-1.147 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5138 | 1.7989 | 2.3471 | 1.0869 | 1.0869 | 2.1623 | 2.1623 |
C2 | 1.5138 | | 2.7110 | 1.3910 | 2.1564 | 2.1564 | 1.0902 | 1.0902 | Cl3 | 1.7989 | 2.7110 | | 3.9458 | 2.3691 | 2.3691 | 2.9251 | 2.9251 | F4 | 2.3471 | 1.3910 | 3.9458 | | 2.6262 | 2.6262 | 2.0201 | 2.0201 | H5 | 1.0869 | 2.1564 | 2.3691 | 2.6262 | | 1.7770 | 2.4969 | 3.0666 | H6 | 1.0869 | 2.1564 | 2.3691 | 2.6262 | 1.7770 | | 3.0666 | 2.4969 | H7 | 2.1623 | 1.0902 | 2.9251 | 2.0201 | 2.4969 | 3.0666 | | 1.7834 | H8 | 2.1623 | 1.0902 | 2.9251 | 2.0201 | 3.0666 | 2.4969 | 1.7834 | |
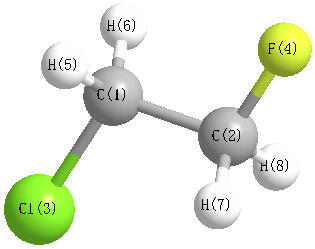 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.733 |
|
C1 |
C2 |
H7 |
111.231 |
| C1 |
C2 |
H8 |
111.231 |
|
C2 |
C1 |
Cl3 |
109.548 |
| C2 |
C1 |
H5 |
110.961 |
|
C2 |
C1 |
H6 |
110.961 |
| Cl3 |
C1 |
H5 |
107.800 |
|
Cl3 |
C1 |
H6 |
107.800 |
| F4 |
C2 |
H7 |
108.397 |
|
F4 |
C2 |
H8 |
108.397 |
| H5 |
C1 |
H6 |
109.665 |
|
H7 |
C2 |
H8 |
109.747 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -638.428012 |
| Energy at 298.15K | -638.433027 |
| HF Energy | -638.196499 |
| Nuclear repulsion energy | 160.698268 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3184 |
3184 |
9.77 |
|
|
|
| 2 |
A |
3140 |
3140 |
27.98 |
|
|
|
| 3 |
A |
3117 |
3117 |
16.18 |
|
|
|
| 4 |
A |
3073 |
3073 |
30.77 |
|
|
|
| 5 |
A |
1534 |
1534 |
2.56 |
|
|
|
| 6 |
A |
1495 |
1495 |
10.35 |
|
|
|
| 7 |
A |
1457 |
1457 |
13.63 |
|
|
|
| 8 |
A |
1373 |
1373 |
34.89 |
|
|
|
| 9 |
A |
1303 |
1303 |
0.59 |
|
|
|
| 10 |
A |
1248 |
1248 |
1.02 |
|
|
|
| 11 |
A |
1122 |
1122 |
43.38 |
|
|
|
| 12 |
A |
1085 |
1085 |
49.41 |
|
|
|
| 13 |
A |
992 |
992 |
6.71 |
|
|
|
| 14 |
A |
869 |
869 |
10.91 |
|
|
|
| 15 |
A |
686 |
686 |
32.41 |
|
|
|
| 16 |
A |
471 |
471 |
13.22 |
|
|
|
| 17 |
A |
291 |
291 |
1.06 |
|
|
|
| 18 |
A |
137 |
137 |
2.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13287.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13287.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.082 |
0.842 |
-0.296 |
| C2 |
1.203 |
0.412 |
0.364 |
| Cl3 |
-1.439 |
-0.284 |
0.064 |
| F4 |
1.673 |
-0.759 |
-0.203 |
| H5 |
-0.377 |
1.827 |
0.062 |
| H6 |
0.035 |
0.866 |
-1.377 |
| H7 |
1.958 |
1.189 |
0.210 |
| H8 |
1.063 |
0.243 |
1.433 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5072 | 1.7995 | 2.3776 | 1.0889 | 1.0876 | 2.1306 | 2.1591 |
C2 | 1.5072 | | 2.7481 | 1.3841 | 2.1421 | 2.1456 | 1.0941 | 1.0908 | Cl3 | 1.7995 | 2.7481 | | 3.1597 | 2.3629 | 2.3607 | 3.7052 | 2.9003 | F4 | 2.3776 | 1.3841 | 3.1597 | | 3.3109 | 2.5893 | 2.0110 | 2.0130 | H5 | 1.0889 | 2.1421 | 2.3629 | 3.3109 | | 1.7788 | 2.4255 | 2.5427 | H6 | 1.0876 | 2.1456 | 2.3607 | 2.5893 | 1.7788 | | 2.5146 | 3.0569 | H7 | 2.1306 | 1.0941 | 3.7052 | 2.0110 | 2.4255 | 2.5146 | | 1.7860 | H8 | 2.1591 | 1.0908 | 2.9003 | 2.0130 | 2.5427 | 3.0569 | 1.7860 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.568 |
|
C1 |
C2 |
H7 |
108.945 |
| C1 |
C2 |
H8 |
111.407 |
|
C2 |
C1 |
Cl3 |
112.117 |
| C2 |
C1 |
H5 |
110.155 |
|
C2 |
C1 |
H6 |
110.514 |
| Cl3 |
C1 |
H5 |
107.205 |
|
Cl3 |
C1 |
H6 |
107.116 |
| F4 |
C2 |
H7 |
107.914 |
|
F4 |
C2 |
H8 |
108.274 |
| H5 |
C1 |
H6 |
109.622 |
|
H7 |
C2 |
H8 |
109.666 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
