return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-235.153550
Energy at 298.15K-235.165905
HF Energy-234.216643
Nuclear repulsion energy232.381736
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3184 3015 61.40      
2 A 3173 3005 2.52      
3 A 3166 2999 47.78      
4 A 3147 2981 0.19      
5 A 3086 2923 5.08      
6 A 3077 2914 2.12      
7 A 1721 1630 0.47      
8 A 1554 1472 12.32      
9 A 1538 1456 4.29      
10 A 1535 1454 2.30      
11 A 1442 1366 0.00      
12 A 1369 1297 0.00      
13 A 1323 1253 0.14      
14 A 1294 1225 0.19      
15 A 1111 1052 1.32      
16 A 1095 1037 0.91      
17 A 1040 985 0.80      
18 A 972 921 0.62      
19 A 865 819 1.38      
20 A 790 748 0.84      
21 A 534 506 0.38      
22 A 326 309 0.00      
23 A 217 205 0.02      
24 A 194 183 0.01      
25 A 38 36 0.00      
26 B 3173 3005 66.83      
27 B 3166 2998 5.92      
28 B 3159 2992 6.05      
29 B 3135 2969 5.73      
30 B 3083 2919 43.27      
31 B 3076 2914 58.40      
32 B 1543 1462 6.88      
33 B 1537 1456 11.89      
34 B 1523 1442 2.26      
35 B 1458 1380 6.70      
36 B 1440 1363 7.56      
37 B 1354 1282 3.77      
38 B 1316 1246 0.13      
39 B 1198 1134 0.15      
40 B 1115 1056 8.56      
41 B 1072 1015 1.73      
42 B 930 881 5.86      
43 B 826 782 4.06      
44 B 760 720 44.89      
45 B 563 533 4.85      
46 B 365 346 1.40      
47 B 242 230 0.05      
48 B 37 35 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 36929.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 34976.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.25446 0.05397 0.05124

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.009 0.672 0.909
C2 -0.009 -0.672 0.909
C3 -0.009 1.577 -0.289
C4 0.009 -1.577 -0.289
C5 -1.370 2.261 -0.458
C6 1.370 -2.261 -0.458
H7 0.003 1.177 1.876
H8 -0.003 -1.177 1.876
H9 0.244 1.026 -1.198
H10 -0.244 -1.026 -1.198
H11 0.766 2.343 -0.165
H12 -0.766 -2.343 -0.165
H13 -1.364 2.957 -1.300
H14 -2.151 1.517 -0.631
H15 -1.638 2.822 0.441
H16 1.364 -2.957 -1.300
H17 2.151 -1.517 -0.631
H18 1.638 -2.822 0.441

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34431.50132.54832.50853.51001.09072.08632.14912.71792.12503.29263.46222.78452.74844.45943.42843.8835
C21.34432.54831.50133.51002.50852.08631.09072.71792.14913.29262.12504.45943.42843.88353.46222.78452.7484
C31.50132.54833.15461.53224.08172.20153.50271.09262.76781.09613.99412.18232.17032.17604.84433.78974.7540
C42.54831.50133.15464.08171.53223.50272.20152.76781.09263.99411.09614.84433.78974.75402.18232.17032.1760
C52.50853.51001.53224.08175.28682.91654.37362.16263.55252.15724.65201.09321.09231.09315.95045.16775.9741
C63.51002.50854.08171.53225.28684.37362.91653.55252.16264.65202.15725.95045.16775.97411.09321.09231.0931
H71.09072.08632.20153.50272.91654.37362.35303.08683.78972.47114.14003.88953.32292.73115.38764.26104.5520
H82.08631.09073.50272.20154.37362.91652.35303.78973.08684.14002.47115.38764.26104.55203.88953.32292.7311
H92.14912.71791.09262.76782.16263.55253.08683.78972.11021.75243.66562.51462.51033.07454.13893.22964.4091
H102.71792.14912.76781.09263.55252.16263.78973.08682.11023.66561.75244.13893.22964.40912.51462.51033.0745
H112.12503.29261.09613.99412.15724.65202.47114.14001.75243.66564.92902.49053.06722.52465.45264.12765.2728
H123.29262.12503.99411.09614.65202.15724.14002.47113.66561.75244.92905.45264.12765.27282.49053.06722.5246
H133.46224.45942.18234.84431.09325.95043.88955.38762.51464.13892.49055.45261.77221.76826.51255.72946.7410
H142.78453.42842.17033.78971.09235.16773.32294.26102.51033.22963.06724.12761.77221.76515.72945.26555.8600
H152.74843.88352.17604.75401.09315.97412.73114.55203.07454.40912.52465.27281.76821.76516.74105.86006.5260
H164.45943.46224.84432.18235.95041.09325.38763.88954.13892.51465.45262.49056.51255.72946.74101.77221.7682
H173.42842.78453.78972.17035.16771.09234.26103.32293.22962.51034.12763.06725.72945.26555.86001.77221.7651
H183.88352.74844.75402.17605.97411.09314.55202.73114.40913.07455.27282.52466.74105.86006.52601.76821.7651

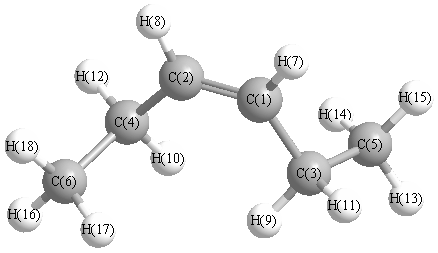
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.062 C1 C2 H8 117.538
C1 C3 C5 111.570 C1 C3 H9 110.909
C1 C3 H11 108.796 C2 C1 C3 127.062
C2 C1 H7 117.538 C2 C4 C6 111.570
C2 C4 H10 110.909 C2 C4 H12 108.796
C3 C1 H7 115.366 C3 C5 H13 111.353
C3 C5 H14 110.449 C3 C5 H15 110.860
C4 C2 H8 115.366 C4 C6 H16 111.353
C4 C6 H17 110.449 C4 C6 H18 110.860
C5 C3 H9 109.824 C5 C3 H11 109.201
C6 C4 H10 109.824 C6 C4 H12 109.201
H9 C3 H11 106.385 H10 C4 H12 106.385
H13 C5 H14 108.358 H13 C5 H15 107.953
H14 C5 H15 107.742 H16 C6 H17 108.358
H16 C6 H18 107.953 H17 C6 H18 107.742
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability