return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-235.625491
Energy at 298.15K-235.637856
HF Energy-235.319485
Nuclear repulsion energy230.853647
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3157 3157 68.77      
2 A 3134 3134 11.50      
3 A 3128 3128 73.91      
4 A 3109 3109 0.05      
5 A 3056 3056 0.92      
6 A 3047 3047 4.77      
7 A 1727 1727 0.69      
8 A 1543 1543 12.47      
9 A 1532 1532 4.42      
10 A 1526 1526 0.00      
11 A 1438 1438 0.11      
12 A 1369 1369 0.09      
13 A 1325 1325 0.05      
14 A 1296 1296 0.25      
15 A 1106 1106 0.71      
16 A 1091 1091 1.10      
17 A 1028 1028 0.30      
18 A 999 999 0.86      
19 A 855 855 1.21      
20 A 789 789 0.65      
21 A 521 521 0.24      
22 A 312 312 0.00      
23 A 210 210 0.01      
24 A 199 199 0.02      
25 A 52 52 0.01      
26 B 3135 3135 78.37      
27 B 3131 3131 24.68      
28 B 3125 3125 2.40      
29 B 3096 3096 6.33      
30 B 3056 3056 77.49      
31 B 3045 3045 54.11      
32 B 1539 1539 4.72      
33 B 1532 1532 10.27      
34 B 1515 1515 2.06      
35 B 1461 1461 5.01      
36 B 1436 1436 4.96      
37 B 1351 1351 8.99      
38 B 1312 1312 0.09      
39 B 1195 1195 0.22      
40 B 1112 1112 7.09      
41 B 1050 1050 2.90      
42 B 923 923 6.65      
43 B 823 823 4.80      
44 B 758 758 38.83      
45 B 568 568 5.30      
46 B 366 366 1.19      
47 B 241 241 0.05      
48 B 48 48 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 36681.7 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 36681.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.28336 0.05064 0.04829

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.006 0.669 0.858
C2 -0.006 -0.669 0.858
C3 -0.006 1.595 -0.326
C4 0.006 -1.595 -0.326
C5 -1.284 2.442 -0.385
C6 1.284 -2.442 -0.385
H7 0.003 1.169 1.826
H8 -0.003 -1.169 1.826
H9 0.110 1.041 -1.259
H10 -0.110 -1.041 -1.259
H11 0.859 2.265 -0.259
H12 -0.859 -2.265 -0.259
H13 -1.257 3.143 -1.222
H14 -2.165 1.807 -0.501
H15 -1.414 3.023 0.532
H16 1.257 -3.143 -1.222
H17 2.165 -1.807 -0.501
H18 1.414 -3.023 0.532

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33851.50292.55522.51993.58551.08932.07732.15172.72362.12633.25643.46942.80292.76824.51883.55593.9651
C21.33852.55521.50293.58552.51992.07731.08932.72362.15173.25642.12634.51883.55593.96513.46942.80292.7682
C31.50292.55523.19051.53454.23852.19373.50281.09172.79801.09613.95402.18222.17702.18114.98444.04024.9070
C42.55521.50293.19054.23851.53453.50282.19372.79801.09173.95401.09614.98444.04024.90702.18222.17702.1811
C52.51993.58551.53454.23855.51782.85724.42302.16153.77772.15464.72751.09241.09171.09276.19225.47456.1631
C63.58552.51994.23851.53455.51784.42302.85723.77772.16154.72752.15466.19225.47456.16311.09241.09171.0927
H71.08932.07732.19373.50282.85724.42302.33733.08953.79612.50614.10833.84343.24402.66865.42664.35304.6083
H82.07731.08933.50282.19374.42302.85722.33733.79613.08954.10832.50615.42664.35304.60833.84343.24402.6686
H92.15172.72361.09172.79802.16153.77773.08953.79612.09271.74983.58702.50752.51783.07554.33773.59324.6282
H102.72362.15172.79801.09173.77772.16153.79613.08952.09273.58701.74984.33773.59324.62822.50752.51783.0755
H112.12633.25641.09613.95402.15464.72752.50614.10831.74983.58704.84502.48503.06862.52355.50714.28375.3753
H123.25642.12633.95401.09614.72752.15464.10832.50613.58701.74984.84505.50714.28375.37532.48503.06862.5235
H133.46944.51882.18224.98441.09246.19223.84345.42662.50754.33772.48505.50711.76861.76466.76926.06086.9442
H142.80293.55592.17704.04021.09175.47453.24404.35302.51783.59323.06864.28371.76861.76296.06085.64126.1002
H152.76823.96512.18114.90701.09276.16312.66864.60833.07554.62822.52355.37531.76461.76296.94426.10026.6748
H164.51883.46944.98442.18226.19221.09245.42663.84344.33772.50755.50712.48506.76926.06086.94421.76861.7646
H173.55592.80294.04022.17705.47451.09174.35303.24403.59322.51784.28373.06866.06085.64126.10021.76861.7629
H183.96512.76824.90702.18116.16311.09274.60832.66864.62823.07555.37532.52356.94426.10026.67481.76461.7629

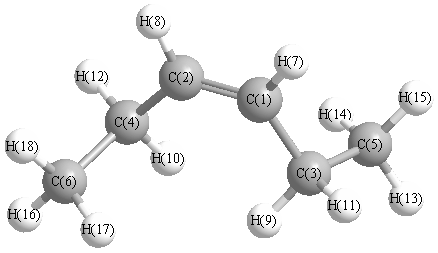
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 128.027 C1 C2 H8 117.286
C1 C3 C5 112.110 C1 C3 H9 111.062
C1 C3 H11 108.784 C2 C1 C3 128.027
C2 C1 H7 117.286 C2 C4 C6 112.110
C2 C4 H10 111.062 C2 C4 H12 108.784
C3 C1 H7 114.674 C3 C5 H13 111.228
C3 C5 H14 110.854 C3 C5 H15 111.128
C4 C2 H8 114.674 C4 C6 H16 111.228
C4 C6 H17 110.854 C4 C6 H18 111.128
C5 C3 H9 109.627 C5 C3 H11 108.837
C6 C4 H10 109.627 C6 C4 H12 108.837
H9 C3 H11 106.220 H10 C4 H12 106.220
H13 C5 H14 108.144 H13 C5 H15 107.721
H14 C5 H15 107.613 H16 C6 H17 108.144
H16 C6 H18 107.721 H17 C6 H18 107.613
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability