return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (2-Hexyne)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.945093
Energy at 298.15K-233.953756
HF Energy-233.019689
Nuclear repulsion energy206.299152
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3168 3000 29.53      
2 A 3165 2997 9.80      
3 A 3095 2931 27.92      
4 A 3082 2919 26.56      
5 A 3076 2913 29.81      
6 A 3068 2905 15.53      
7 A 2285 2164 0.10      
8 A 1546 1464 9.61      
9 A 1532 1451 1.01      
10 A 1522 1442 8.80      
11 A 1519 1439 1.27      
12 A 1451 1374 1.19      
13 A 1448 1372 4.89      
14 A 1411 1337 2.80      
15 A 1323 1253 5.62      
16 A 1194 1131 0.46      
17 A 1137 1077 3.67      
18 A 1083 1025 0.76      
19 A 1055 999 0.64      
20 A 919 870 3.91      
21 A 770 729 0.27      
22 A 450 426 2.78      
23 A 324 307 0.59      
24 A 230 217 5.43      
25 A 94 89 1.48      
26 A 3167 2999 55.27      
27 A 3166 2998 8.55      
28 A 3143 2977 3.57      
29 A 3111 2946 5.80      
30 A 1540 1459 10.54      
31 A 1523 1442 8.22      
32 A 1355 1283 0.00      
33 A 1287 1219 0.22      
34 A 1137 1077 0.42      
35 A 1055 1000 0.79      
36 A 884 837 0.50      
37 A 745 706 2.87      
38 A 268 254 0.03      
39 A 236 224 0.18      
40 A 210 199 9.33      
41 A 82 77 0.33      
42 A 21 20 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 31435.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29772.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.58997 0.04002 0.03855

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.274 -2.797 0.000
C2 -1.806 2.811 0.000
C3 0.955 -0.280 0.000
C4 0.000 0.829 0.000
C5 0.267 -1.651 0.000
C6 -0.832 1.721 0.000
H7 0.771 -3.767 0.000
H8 -2.827 2.422 0.000
H9 1.917 -2.754 0.884
H10 1.917 -2.754 -0.884
H11 -0.383 -1.717 -0.878
H12 -0.383 -1.717 0.878
H13 1.607 -0.200 -0.878
H14 1.607 -0.200 0.878
H15 -1.685 3.442 0.884
H16 -1.685 3.442 -0.884

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C16.39802.53723.84291.52544.98421.09256.63661.09351.09352.16332.16332.76152.76156.96106.9610
C26.39804.14422.68164.92001.46217.06471.09266.75386.75384.82734.82734.63524.63521.09241.0924
C32.53724.14421.46291.53382.68213.49194.64702.79832.79832.15082.15081.09661.09664.64754.6475
C43.84292.68161.46292.49391.21974.65983.24454.15864.15862.72032.72032.10032.10033.23243.2324
C51.52544.92001.53382.49393.54612.17535.11422.17252.17251.09421.09422.16132.16135.52495.5249
C64.98421.46212.68211.21973.54615.71692.11425.32575.32573.57673.57673.22613.22612.11452.1145
H71.09257.06473.49194.65982.17535.71697.15811.76581.76582.51042.51043.76733.76737.66657.6665
H86.63661.09264.64703.24455.11422.11427.15817.07607.07604.88604.88605.22495.22491.76761.7676
H91.09356.75382.79834.15862.17255.32571.76587.07601.76723.07662.52253.11852.57347.16697.3817
H101.09356.75382.79834.15862.17255.32571.76587.07601.76722.52253.07662.57343.11857.38177.1669
H112.16334.82732.15082.72031.09423.57672.51044.88603.07662.52251.75562.50253.05695.60495.3208
H122.16334.82732.15082.72031.09423.57672.51044.88602.52253.07661.75563.05692.50255.32085.6049
H132.76154.63521.09662.10032.16133.22613.76735.22493.11852.57342.50253.05691.75585.21574.9091
H142.76154.63521.09662.10032.16133.22613.76735.22492.57343.11853.05692.50251.75584.90915.2157
H156.96101.09244.64753.23245.52492.11457.66651.76767.16697.38175.60495.32085.21574.90911.7681
H166.96101.09244.64753.23245.52492.11457.66651.76767.38177.16695.32085.60494.90915.21571.7681

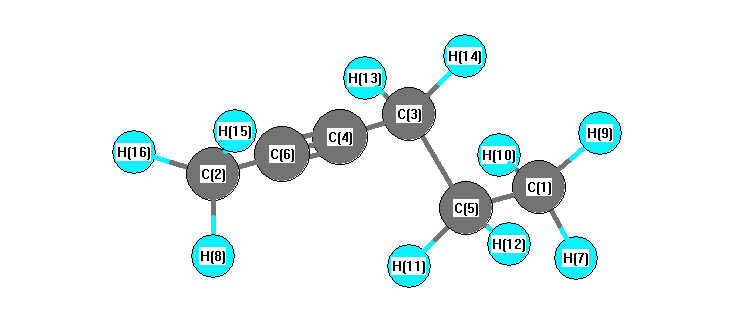
picture of 2-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 C3 112.072 C1 C5 H11 110.257
C1 C5 H12 110.257 C2 C6 C4 178.758
C3 C4 C6 177.725 C3 C5 H11 108.704
C3 C5 H12 108.704 C4 C3 C5 112.632
C4 C3 H13 109.450 C4 C3 H14 109.450
C5 C1 H7 111.308 C5 C1 H9 111.030
C5 C1 H10 111.030 C5 C3 H13 109.377
C5 C3 H14 109.377 C6 C2 H8 110.875
C6 C2 H15 110.910 C6 C2 H16 110.910
H7 C1 H9 107.750 H7 C1 H10 107.750
H8 C2 H15 107.987 H8 C2 H16 107.987
H9 C1 H10 107.811 H11 C5 H12 106.679
H13 C3 H14 106.359 H15 C2 H16 108.048
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability