return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7N (Cyclopropylamine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-172.775365
Energy at 298.15K-172.783868
HF Energy-172.115384
Nuclear repulsion energy125.340420
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3546 3358 0.86      
2 A 3295 3121 13.99      
3 A 3190 3021 4.82      
4 A 3131 2965 46.27      
5 A 1739 1647 27.91      
6 A 1543 1461 17.71      
7 A 1445 1369 14.69      
8 A 1271 1204 0.32      
9 A 1206 1142 7.43      
10 A 1073 1016 21.33      
11 A 1043 988 10.78      
12 A 898 851 136.96      
13 A 842 797 25.11      
14 A 794 752 2.72      
15 A 418 396 6.46      
16 A 3642 3449 0.91      
17 A 3283 3110 0.30      
18 A 3184 3015 14.62      
19 A 1486 1407 4.53      
20 A 1311 1242 0.87      
21 A 1222 1157 0.20      
22 A 1154 1093 1.04      
23 A 1077 1020 2.42      
24 A 929 880 10.60      
25 A 867 821 3.68      
26 A 405 384 6.93      
27 A 278 263 42.29      

Unscaled Zero Point Vibrational Energy (zpe) 22135.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20964.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.54542 0.22544 0.19448

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.532 0.000
H2 -0.415 1.541 0.000
N3 1.431 0.369 0.000
C4 -0.789 -0.501 0.755
C5 -0.789 -0.501 -0.755
H6 1.831 0.811 0.819
H7 1.831 0.811 -0.819
H8 -1.690 -0.188 1.270
H9 -1.690 -0.188 -1.270
H10 -0.210 -1.274 1.244
H11 -0.210 -1.274 -1.244

Atom - Atom Distances (Å)
  C1 H2 N3 C4 C5 H6 H7 H8 H9 H10 H11
C11.09131.44031.50321.50322.02522.02522.23292.23292.20342.2034
H21.09132.18672.20932.20932.49972.49972.49562.49563.08513.0851
N31.44032.18672.50112.50111.01301.01303.41503.41502.63462.6346
C41.50322.20932.50111.51012.93083.32621.08362.23791.08302.2206
C51.50322.20932.50111.51013.32622.93082.23791.08362.22061.0830
H62.02522.49971.01302.93083.32621.63813.68754.21392.94853.5736
H72.02522.49971.01303.32622.93081.63814.21393.68753.57362.9485
H82.23292.49563.41501.08362.23793.68754.21392.53901.83603.1129
H92.23292.49563.41502.23791.08364.21393.68752.53903.11291.8360
H102.20343.08512.63461.08302.22062.94853.57361.83603.11292.4888
H112.20343.08512.63462.22061.08303.57362.94853.11291.83602.4888

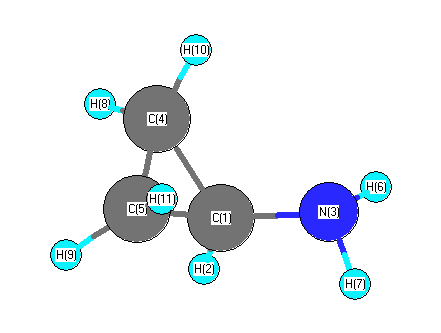
picture of Cyclopropylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 110.059 C1 N3 H7 110.059
C1 C4 C5 59.848 C1 C4 H8 118.453
C1 C4 H10 115.904 C1 C5 C4 59.848
C1 C5 H9 118.453 C1 C5 H11 115.904
H2 C1 N3 118.842 H2 C1 C4 115.851
H2 C1 C5 115.851 N3 C1 C4 116.343
N3 C1 C5 116.343 C4 C1 C5 60.304
C4 C5 H9 118.346 C4 C5 H11 116.863
C5 C4 H8 118.346 C5 C4 H10 116.863
H6 N3 H7 107.910 H8 C4 H10 115.867
H9 C5 H11 115.867
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability