return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Methyl cyclopropyl ketone)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-270.343094
Energy at 298.15K-270.351761
HF Energy-270.020265
Nuclear repulsion energy230.389012
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3265 3265 9.89      
2 A' 3193 3193 11.59      
3 A' 3176 3176 12.78      
4 A' 3164 3164 11.85      
5 A' 3059 3059 4.25      
6 A' 1765 1765 113.40      
7 A' 1519 1519 9.52      
8 A' 1499 1499 17.67      
9 A' 1435 1435 112.46      
10 A' 1407 1407 9.78      
11 A' 1238 1238 12.13      
12 A' 1208 1208 77.19      
13 A' 1126 1126 13.97      
14 A' 1073 1073 10.47      
15 A' 982 982 51.12      
16 A' 927 927 20.80      
17 A' 823 823 2.33      
18 A' 760 760 0.33      
19 A' 595 595 10.49      
20 A' 376 376 4.06      
21 A' 243 243 3.93      
22 A" 3251 3251 0.39      
23 A" 3161 3161 17.54      
24 A" 3122 3122 11.88      
25 A" 1508 1508 11.25      
26 A" 1482 1482 5.11      
27 A" 1232 1232 0.31      
28 A" 1141 1141 2.39      
29 A" 1103 1103 1.97      
30 A" 1066 1066 4.45      
31 A" 889 889 0.20      
32 A" 841 841 5.78      
33 A" 609 609 1.57      
34 A" 260 260 0.29      
35 A" 144 144 0.06      
36 A" 72 72 1.86      

Unscaled Zero Point Vibrational Energy (zpe) 26355.4 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 26355.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.24048 0.08761 0.07659

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.481 -0.555 0.000
C2 -0.280 -0.758 0.000
C3 0.299 -2.156 0.000
C4 0.700 0.367 0.000
C5 0.299 1.633 0.744
C6 0.299 1.633 -0.744
H7 -0.504 -2.890 0.000
H8 0.932 -2.303 -0.879
H9 0.932 -2.303 0.879
H10 1.751 0.103 0.000
H11 -0.661 1.595 1.242
H12 1.082 2.165 1.268
H13 -0.661 1.595 -1.242
H14 1.082 2.165 -1.268

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.21752.39382.36722.91612.91612.53143.10653.10653.29752.61423.94622.61423.9462
C21.21751.51311.49212.56982.56982.14382.15162.15162.20582.68773.46522.68773.4652
C32.39381.51312.55483.86083.86081.08801.09341.09342.68524.06604.57074.06604.5707
C42.36721.49212.55481.52161.52163.47272.82082.82081.08362.21402.23292.21402.2329
C52.91612.56983.86081.52161.48844.65324.30403.98842.23621.08221.08242.20652.2240
C62.91612.56983.86081.52161.48844.65323.98844.30402.23622.20652.22401.08221.0824
H72.53142.14381.08803.47274.65324.65321.78331.78333.74734.65645.44784.65645.4478
H83.10652.15161.09342.82084.30403.98841.78331.75832.68924.71494.95944.22644.4873
H93.10652.15161.09342.82083.98844.30401.78331.75832.68924.22644.48734.71494.9594
H103.29752.20582.68521.08362.23622.23623.74732.68922.68923.09602.51123.09602.5112
H112.61422.68774.06602.21401.08222.20654.65644.71494.22643.09601.83462.48423.1089
H123.94623.46524.57072.23291.08242.22405.44784.95944.48732.51121.83463.10892.5360
H132.61422.68774.06602.21402.20651.08224.65644.22644.71493.09602.48423.10891.8346
H143.94623.46524.57072.23292.22401.08245.44784.48734.95942.51123.10892.53601.8346

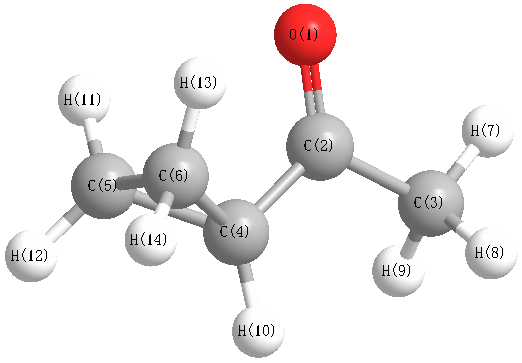
picture of Methyl cyclopropyl ketone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 122.115 O1 C2 C4 121.435
C2 C3 H7 109.942 C2 C3 H8 110.235
C2 C3 H9 110.235 C2 C4 C5 117.004
C2 C4 C6 117.004 C2 C4 H10 116.937
C3 C2 C4 116.450 C4 C5 C6 60.719
C4 C5 H11 115.453 C4 C5 H12 117.067
C4 C6 C5 60.719 C4 C6 H13 115.453
C4 C6 H14 117.067 C5 C4 C6 58.561
C5 C4 H10 117.274 C5 C6 H13 117.392
C5 C6 H14 118.941 C6 C4 H10 117.274
C6 C5 H11 117.392 C6 C5 H12 118.941
H7 C3 H8 109.671 H7 C3 H9 109.671
H8 C3 H9 107.040 H11 C5 H12 115.890
H13 C6 H14 115.890
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability