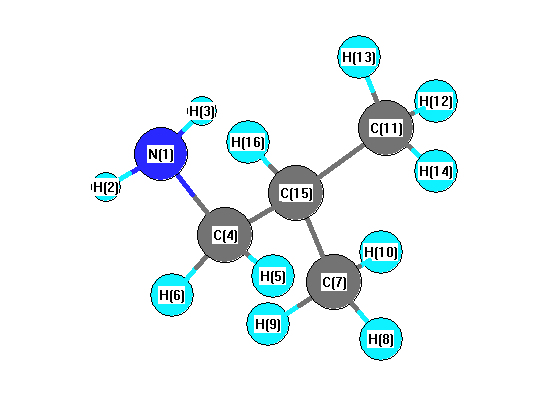
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3820 |
3454 |
0.08 |
|
|
|
| 2 |
A |
3751 |
3393 |
1.01 |
|
|
|
| 3 |
A |
3241 |
2931 |
61.96 |
|
|
|
| 4 |
A |
3233 |
2924 |
129.49 |
|
|
|
| 5 |
A |
3229 |
2920 |
97.38 |
|
|
|
| 6 |
A |
3217 |
2910 |
19.10 |
|
|
|
| 7 |
A |
3208 |
2902 |
26.71 |
|
|
|
| 8 |
A |
3182 |
2878 |
16.45 |
|
|
|
| 9 |
A |
3170 |
2867 |
38.30 |
|
|
|
| 10 |
A |
3163 |
2860 |
38.93 |
|
|
|
| 11 |
A |
3103 |
2807 |
79.08 |
|
|
|
| 12 |
A |
1862 |
1684 |
45.03 |
|
|
|
| 13 |
A |
1653 |
1495 |
12.59 |
|
|
|
| 14 |
A |
1643 |
1486 |
1.30 |
|
|
|
| 15 |
A |
1638 |
1481 |
4.97 |
|
|
|
| 16 |
A |
1628 |
1473 |
1.84 |
|
|
|
| 17 |
A |
1621 |
1466 |
0.97 |
|
|
|
| 18 |
A |
1576 |
1425 |
9.86 |
|
|
|
| 19 |
A |
1555 |
1406 |
2.13 |
|
|
|
| 20 |
A |
1543 |
1396 |
6.49 |
|
|
|
| 21 |
A |
1501 |
1358 |
1.48 |
|
|
|
| 22 |
A |
1473 |
1332 |
0.05 |
|
|
|
| 23 |
A |
1434 |
1297 |
7.19 |
|
|
|
| 24 |
A |
1340 |
1212 |
0.09 |
|
|
|
| 25 |
A |
1295 |
1171 |
1.97 |
|
|
|
| 26 |
A |
1272 |
1151 |
10.24 |
|
|
|
| 27 |
A |
1174 |
1062 |
1.93 |
|
|
|
| 28 |
A |
1163 |
1052 |
24.54 |
|
|
|
| 29 |
A |
1041 |
941 |
0.21 |
|
|
|
| 30 |
A |
1020 |
922 |
81.22 |
|
|
|
| 31 |
A |
1009 |
912 |
0.68 |
|
|
|
| 32 |
A |
1002 |
906 |
0.19 |
|
|
|
| 33 |
A |
931 |
842 |
75.00 |
|
|
|
| 34 |
A |
846 |
765 |
2.55 |
|
|
|
| 35 |
A |
515 |
466 |
7.11 |
|
|
|
| 36 |
A |
447 |
404 |
0.63 |
|
|
|
| 37 |
A |
389 |
352 |
0.66 |
|
|
|
| 38 |
A |
300 |
272 |
20.95 |
|
|
|
| 39 |
A |
286 |
258 |
5.40 |
|
|
|
| 40 |
A |
252 |
228 |
18.89 |
|
|
|
| 41 |
A |
234 |
212 |
15.70 |
|
|
|
| 42 |
A |
129 |
117 |
4.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35042.2 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 31692.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.840 |
-1.093 |
|
-1.095 |
| 2 |
H |
0.333 |
0.399 |
|
0.406 |
| 3 |
H |
0.332 |
0.379 |
|
0.388 |
| 4 |
C |
-0.193 |
0.346 |
|
0.243 |
| 5 |
H |
0.176 |
-0.090 |
|
-0.050 |
| 6 |
H |
0.203 |
-0.007 |
|
0.029 |
| 7 |
C |
-0.612 |
-0.458 |
|
-0.622 |
| 8 |
H |
0.199 |
0.081 |
|
0.129 |
| 9 |
H |
0.208 |
0.067 |
|
0.108 |
| 10 |
H |
0.211 |
0.110 |
|
0.155 |
| 11 |
C |
-0.629 |
-0.244 |
|
-0.428 |
| 12 |
H |
0.212 |
0.037 |
|
0.088 |
| 13 |
H |
0.217 |
0.045 |
|
0.095 |
| 14 |
H |
0.197 |
0.041 |
|
0.096 |
| 15 |
C |
-0.220 |
0.473 |
|
0.505 |
| 16 |
H |
0.208 |
-0.085 |
|
-0.049 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.262 |
-0.245 |
1.230 |
1.281 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.269 |
-0.227 |
1.265 |
1.313 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.822 |
0.820 |
-3.388 |
| y |
0.820 |
-33.543 |
0.598 |
| z |
-3.388 |
0.598 |
-35.836 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.868 |
0.820 |
-3.388 |
| y |
0.820 |
0.785 |
0.598 |
| z |
-3.388 |
0.598 |
-2.653 |
|
| Polar |
|---|
| 3z2-r2 | -5.306 |
|---|
| x2-y2 | 0.722 |
|---|
| xy | 0.820 |
|---|
| xz | -3.388 |
|---|
| yz | 0.598 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.157 |
-0.017 |
-0.011 |
| y |
-0.017 |
7.665 |
0.019 |
| z |
-0.011 |
0.019 |
6.732 |
<r2> (average value of r
2) Å
2
| <r2> |
150.206 |
| (<r2>)1/2 |
12.256 |