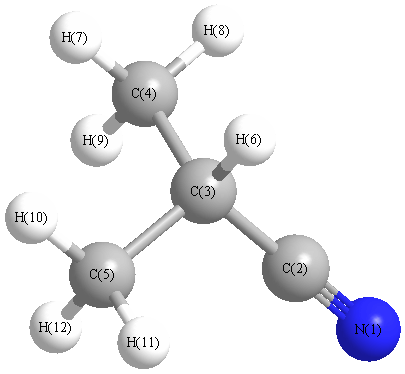
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3270 |
2957 |
57.45 |
|
|
|
| 2 |
A' |
3264 |
2952 |
66.50 |
|
|
|
| 3 |
A' |
3216 |
2908 |
3.25 |
|
|
|
| 4 |
A' |
3196 |
2890 |
23.89 |
|
|
|
| 5 |
A' |
2588 |
2341 |
14.70 |
|
|
|
| 6 |
A' |
1644 |
1487 |
10.68 |
|
|
|
| 7 |
A' |
1634 |
1478 |
6.68 |
|
|
|
| 8 |
A' |
1564 |
1414 |
3.14 |
|
|
|
| 9 |
A' |
1483 |
1342 |
8.92 |
|
|
|
| 10 |
A' |
1299 |
1174 |
3.26 |
|
|
|
| 11 |
A' |
1230 |
1112 |
5.82 |
|
|
|
| 12 |
A' |
989 |
894 |
0.45 |
|
|
|
| 13 |
A' |
813 |
735 |
3.14 |
|
|
|
| 14 |
A' |
602 |
545 |
1.61 |
|
|
|
| 15 |
A' |
380 |
343 |
0.45 |
|
|
|
| 16 |
A' |
314 |
284 |
1.77 |
|
|
|
| 17 |
A' |
244 |
221 |
2.47 |
|
|
|
| 18 |
A" |
3265 |
2953 |
26.21 |
|
|
|
| 19 |
A" |
3256 |
2944 |
0.80 |
|
|
|
| 20 |
A" |
3191 |
2886 |
27.34 |
|
|
|
| 21 |
A" |
1623 |
1467 |
3.63 |
|
|
|
| 22 |
A" |
1621 |
1466 |
0.02 |
|
|
|
| 23 |
A" |
1543 |
1395 |
2.10 |
|
|
|
| 24 |
A" |
1455 |
1316 |
0.21 |
|
|
|
| 25 |
A" |
1225 |
1108 |
1.84 |
|
|
|
| 26 |
A" |
1047 |
947 |
0.26 |
|
|
|
| 27 |
A" |
1018 |
921 |
1.27 |
|
|
|
| 28 |
A" |
633 |
572 |
0.02 |
|
|
|
| 29 |
A" |
245 |
222 |
0.01 |
|
|
|
| 30 |
A" |
212 |
192 |
6.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24030.3 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 21733.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.339 |
|
|
|
| 2 |
C |
0.184 |
|
|
|
| 3 |
C |
-0.275 |
|
|
|
| 4 |
C |
-0.597 |
|
|
|
| 5 |
C |
-0.597 |
|
|
|
| 6 |
H |
0.255 |
|
|
|
| 7 |
H |
0.223 |
|
|
|
| 8 |
H |
0.234 |
|
|
|
| 9 |
H |
0.227 |
|
|
|
| 10 |
H |
0.223 |
|
|
|
| 11 |
H |
0.234 |
|
|
|
| 12 |
H |
0.227 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.216 |
4.040 |
0.000 |
4.219 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.905 |
1.781 |
0.000 |
| y |
1.781 |
-40.106 |
0.000 |
| z |
0.000 |
0.000 |
-30.528 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.412 |
1.781 |
0.000 |
| y |
1.781 |
-9.389 |
0.000 |
| z |
0.000 |
0.000 |
4.977 |
|
| Polar |
|---|
| 3z2-r2 | 9.954 |
|---|
| x2-y2 | 9.201 |
|---|
| xy | 1.781 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.619 |
-0.563 |
0.000 |
| y |
-0.563 |
7.840 |
0.000 |
| z |
0.000 |
0.000 |
6.168 |
<r2> (average value of r
2) Å
2
| <r2> |
128.405 |
| (<r2>)1/2 |
11.332 |