return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Propanol, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.041835
Energy at 298.15K-233.053200
Nuclear repulsion energy193.540062
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3863 3658 19.71      
2 A 3167 2999 29.35      
3 A 3164 2997 26.14      
4 A 3157 2990 67.07      
5 A 3151 2984 11.87      
6 A 3147 2980 16.83      
7 A 3089 2926 18.16      
8 A 3069 2907 19.61      
9 A 3065 2902 23.34      
10 A 3031 2871 47.44      
11 A 1562 1479 18.20      
12 A 1544 1462 8.14      
13 A 1542 1460 0.57      
14 A 1535 1454 5.62      
15 A 1524 1444 1.36      
16 A 1461 1384 24.21      
17 A 1445 1369 4.52      
18 A 1439 1363 27.93      
19 A 1422 1347 2.39      
20 A 1399 1325 4.36      
21 A 1350 1278 0.02      
22 A 1282 1214 15.61      
23 A 1226 1161 0.45      
24 A 1187 1124 2.76      
25 A 1146 1085 11.76      
26 A 1110 1051 113.58      
27 A 997 944 0.20      
28 A 978 926 6.19      
29 A 948 897 0.86      
30 A 919 870 9.06      
31 A 842 797 7.09      
32 A 502 475 6.41      
33 A 432 409 1.45      
34 A 373 353 28.74      
35 A 325 308 87.55      
36 A 267 253 28.98      
37 A 255 242 14.89      
38 A 229 217 2.01      
39 A 127 120 4.09      

Unscaled Zero Point Vibrational Energy (zpe) 30632.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29011.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.25415 0.11804 0.08882

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.454 0.004 -0.355
C2 -0.764 -0.714 0.218
C3 1.726 -0.740 0.040
C4 0.500 1.459 0.102
O5 -1.998 -0.116 -0.148
H6 0.352 -0.015 -1.447
H7 1.695 -1.784 -0.285
H8 2.609 -0.277 -0.407
H9 1.860 -0.731 1.127
H10 -0.362 2.022 -0.264
H11 0.517 1.520 1.196
H12 1.396 1.961 -0.272
H13 -0.808 -1.736 -0.166
H14 -0.666 -0.775 1.313
H15 -2.097 0.690 0.364

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52541.52581.52552.46331.09692.17792.17392.17082.17862.16992.17382.15762.15422.7377
C21.52542.49652.51661.41942.12302.72913.45812.77712.80742.75523.47311.09271.10121.9417
C31.52582.49652.51803.78072.15041.09371.09291.09493.47562.81162.73892.73062.70914.0947
C41.52552.51662.51802.96352.14293.47742.77772.77351.09261.09611.09293.46272.79552.7216
O52.46331.41943.78072.96352.68734.05524.61694.10952.69473.28783.98102.01052.08460.9597
H61.09692.12302.15042.14292.68732.50652.49833.06752.46173.06092.52502.43903.03813.1268
H72.17792.72911.09373.47744.05522.50651.76651.76924.32643.80773.75672.50703.02434.5746
H82.17393.45811.09292.77774.61692.49831.76651.76623.75863.18932.54853.72343.73194.8658
H92.17082.77711.09492.77354.10953.06751.76921.76623.80092.62173.06843.13092.53264.2732
H102.17862.80743.47561.09262.69472.46174.32643.75863.80091.77631.75883.78603.22522.2763
H112.16992.75522.81161.09613.28783.06093.80773.18932.62171.77631.76643.77052.58462.8668
H122.17383.47312.73891.09293.98102.52503.75672.54853.06841.75881.76644.30563.77433.7714
H132.15761.09272.73063.46272.01052.43902.50703.72343.13093.78603.77054.30561.77002.7981
H142.15421.10122.70912.79552.08463.03813.02433.73192.53263.22522.58463.77431.77002.2577
H152.73771.94174.09472.72160.95973.12684.57464.86584.27322.27632.86683.77142.79812.2577

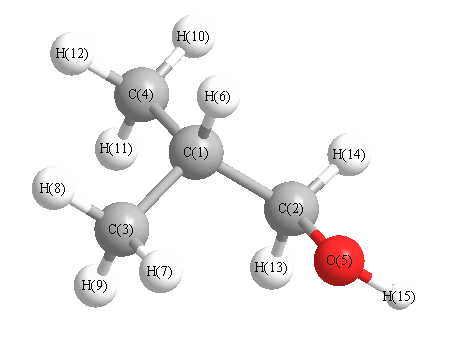
picture of 1-Propanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 113.500 C1 C2 H13 109.897
C1 C2 H14 109.132 C1 C3 H7 111.417
C1 C3 H8 111.149 C1 C3 H9 110.785
C1 C4 H10 111.570 C1 C4 H11 110.660
C1 C4 H12 111.165 C2 C1 C3 109.813
C2 C1 C4 111.156 C2 C1 H6 106.974
C2 O5 H15 107.841 C3 C1 C4 111.225
C3 C1 H6 109.052 C4 C1 H6 108.497
O5 C2 H13 105.586 O5 C2 H14 110.956
H7 C3 H8 107.781 H7 C3 H9 107.876
H8 C3 H9 107.671 H10 C4 H11 108.498
H10 C4 H12 107.180 H11 C4 H12 107.602
H13 C2 H14 107.564
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability