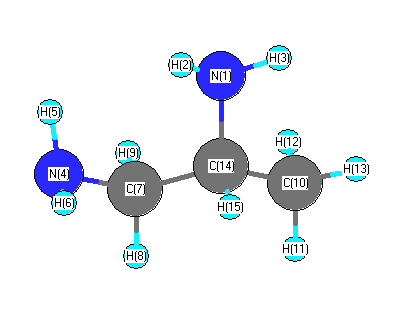
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3828 |
3462 |
0.02 |
|
|
|
| 2 |
A |
3815 |
3450 |
0.18 |
|
|
|
| 3 |
A |
3749 |
3390 |
1.04 |
|
|
|
| 4 |
A |
3744 |
3386 |
0.79 |
|
|
|
| 5 |
A |
3250 |
2939 |
89.60 |
|
|
|
| 6 |
A |
3235 |
2925 |
16.33 |
|
|
|
| 7 |
A |
3229 |
2921 |
63.83 |
|
|
|
| 8 |
A |
3193 |
2888 |
57.34 |
|
|
|
| 9 |
A |
3175 |
2871 |
29.95 |
|
|
|
| 10 |
A |
3080 |
2785 |
100.23 |
|
|
|
| 11 |
A |
1861 |
1683 |
25.10 |
|
|
|
| 12 |
A |
1849 |
1672 |
58.85 |
|
|
|
| 13 |
A |
1636 |
1479 |
2.73 |
|
|
|
| 14 |
A |
1628 |
1472 |
5.08 |
|
|
|
| 15 |
A |
1616 |
1462 |
0.35 |
|
|
|
| 16 |
A |
1568 |
1418 |
0.52 |
|
|
|
| 17 |
A |
1547 |
1399 |
13.68 |
|
|
|
| 18 |
A |
1529 |
1383 |
7.25 |
|
|
|
| 19 |
A |
1508 |
1364 |
7.58 |
|
|
|
| 20 |
A |
1469 |
1329 |
13.58 |
|
|
|
| 21 |
A |
1375 |
1243 |
0.65 |
|
|
|
| 22 |
A |
1318 |
1192 |
7.83 |
|
|
|
| 23 |
A |
1264 |
1143 |
10.85 |
|
|
|
| 24 |
A |
1187 |
1074 |
19.36 |
|
|
|
| 25 |
A |
1140 |
1031 |
17.88 |
|
|
|
| 26 |
A |
1098 |
993 |
2.58 |
|
|
|
| 27 |
A |
1029 |
930 |
11.44 |
|
|
|
| 28 |
A |
995 |
900 |
116.94 |
|
|
|
| 29 |
A |
963 |
870 |
142.01 |
|
|
|
| 30 |
A |
931 |
842 |
73.74 |
|
|
|
| 31 |
A |
857 |
775 |
56.96 |
|
|
|
| 32 |
A |
525 |
475 |
0.57 |
|
|
|
| 33 |
A |
503 |
454 |
20.19 |
|
|
|
| 34 |
A |
397 |
359 |
0.82 |
|
|
|
| 35 |
A |
373 |
338 |
58.64 |
|
|
|
| 36 |
A |
279 |
252 |
10.29 |
|
|
|
| 37 |
A |
264 |
238 |
22.37 |
|
|
|
| 38 |
A |
238 |
215 |
35.63 |
|
|
|
| 39 |
A |
132 |
119 |
5.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32685.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 29560.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.839 |
-1.033 |
|
-1.015 |
| 2 |
H |
0.323 |
0.331 |
|
0.328 |
| 3 |
H |
0.336 |
0.386 |
|
0.386 |
| 4 |
N |
-0.837 |
-1.083 |
|
-1.078 |
| 5 |
H |
0.332 |
0.403 |
|
0.407 |
| 6 |
H |
0.320 |
0.368 |
|
0.372 |
| 7 |
C |
-0.171 |
0.480 |
|
0.406 |
| 8 |
H |
0.203 |
-0.052 |
|
-0.023 |
| 9 |
H |
0.212 |
-0.031 |
|
-0.006 |
| 10 |
C |
-0.618 |
-0.442 |
|
-0.507 |
| 11 |
H |
0.209 |
0.067 |
|
0.088 |
| 12 |
H |
0.219 |
0.109 |
|
0.131 |
| 13 |
H |
0.202 |
0.125 |
|
0.142 |
| 14 |
C |
-0.056 |
0.443 |
|
0.421 |
| 15 |
H |
0.164 |
-0.072 |
|
-0.052 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.074 |
1.152 |
1.822 |
2.408 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.065 |
1.153 |
1.788 |
2.380 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.438 |
-0.230 |
-0.106 |
| y |
-0.230 |
6.935 |
-0.051 |
| z |
-0.106 |
-0.051 |
6.249 |
<r2> (average value of r
2) Å
2
| <r2> |
143.436 |
| (<r2>)1/2 |
11.976 |