Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-311G*
Geometric Data calculated at B3PW91/6-311G*
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -232.435414 |
| Energy at 298.15K | -232.443654 |
| Nuclear repulsion energy | 176.233380 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3040 |
13.02 |
|
|
|
| 2 |
A |
3135 |
3018 |
25.34 |
|
|
|
| 3 |
A |
3126 |
3009 |
26.87 |
|
|
|
| 4 |
A |
3106 |
2990 |
13.74 |
|
|
|
| 5 |
A |
3055 |
2942 |
24.16 |
|
|
|
| 6 |
A |
3051 |
2937 |
13.52 |
|
|
|
| 7 |
A |
3042 |
2929 |
7.74 |
|
|
|
| 8 |
A |
3023 |
2910 |
18.75 |
|
|
|
| 9 |
A |
1817 |
1749 |
152.51 |
|
|
|
| 10 |
A |
1513 |
1456 |
10.91 |
|
|
|
| 11 |
A |
1506 |
1450 |
10.35 |
|
|
|
| 12 |
A |
1489 |
1433 |
12.35 |
|
|
|
| 13 |
A |
1478 |
1423 |
22.34 |
|
|
|
| 14 |
A |
1462 |
1408 |
7.87 |
|
|
|
| 15 |
A |
1424 |
1371 |
8.22 |
|
|
|
| 16 |
A |
1397 |
1345 |
63.81 |
|
|
|
| 17 |
A |
1377 |
1326 |
11.02 |
|
|
|
| 18 |
A |
1292 |
1244 |
0.00 |
|
|
|
| 19 |
A |
1192 |
1148 |
70.40 |
|
|
|
| 20 |
A |
1134 |
1091 |
0.61 |
|
|
|
| 21 |
A |
1114 |
1072 |
1.30 |
|
|
|
| 22 |
A |
1011 |
974 |
4.79 |
|
|
|
| 23 |
A |
958 |
922 |
4.14 |
|
|
|
| 24 |
A |
946 |
910 |
10.95 |
|
|
|
| 25 |
A |
770 |
741 |
3.49 |
|
|
|
| 26 |
A |
753 |
725 |
3.20 |
|
|
|
| 27 |
A |
594 |
572 |
7.86 |
|
|
|
| 28 |
A |
476 |
458 |
0.23 |
|
|
|
| 29 |
A |
399 |
385 |
3.97 |
|
|
|
| 30 |
A |
247 |
237 |
5.16 |
|
|
|
| 31 |
A |
199 |
192 |
0.16 |
|
|
|
| 32 |
A |
106 |
102 |
0.10 |
|
|
|
| 33 |
A |
55 |
53 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24701.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 23780.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.880 |
-0.504 |
0.033 |
| C2 |
0.526 |
0.170 |
-0.020 |
| C3 |
-0.682 |
-0.750 |
-0.058 |
| C4 |
-2.011 |
-0.022 |
0.048 |
| O5 |
0.415 |
1.374 |
-0.022 |
| H6 |
2.667 |
0.226 |
-0.154 |
| H7 |
2.035 |
-0.938 |
1.028 |
| H8 |
1.953 |
-1.325 |
-0.686 |
| H9 |
-2.847 |
-0.725 |
-0.011 |
| H10 |
-2.089 |
0.522 |
0.992 |
| H11 |
-2.122 |
0.710 |
-0.754 |
| H12 |
-0.623 |
-1.326 |
-0.992 |
| H13 |
-0.570 |
-1.500 |
0.737 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5137 | 2.5757 | 3.9212 | 2.3824 | 1.0898 | 1.0959 | 1.0941 | 4.7325 | 4.2100 | 4.2557 | 2.8278 | 2.7369 |
C2 | 1.5137 | | 1.5189 | 2.5454 | 1.2085 | 2.1464 | 2.1454 | 2.1716 | 3.4896 | 2.8256 | 2.8003 | 2.1226 | 2.1359 | C3 | 2.5757 | 1.5189 | | 1.5190 | 2.3901 | 3.4900 | 2.9320 | 2.7693 | 2.1656 | 2.1676 | 2.1655 | 1.0992 | 1.0983 | C4 | 3.9212 | 2.5454 | 1.5190 | | 2.7997 | 4.6896 | 4.2628 | 4.2369 | 1.0932 | 1.0924 | 1.0917 | 2.1696 | 2.1758 | O5 | 2.3824 | 1.2085 | 2.3901 | 2.7997 | | 2.5315 | 3.0119 | 3.1763 | 3.8782 | 2.8320 | 2.7220 | 3.0506 | 3.1306 | H6 | 1.0898 | 2.1464 | 3.4900 | 4.6896 | 2.5315 | | 1.7754 | 1.7885 | 5.5975 | 4.9012 | 4.8508 | 3.7338 | 3.7755 | H7 | 1.0959 | 2.1454 | 2.9320 | 4.2628 | 3.0119 | 1.7754 | | 1.7587 | 4.9957 | 4.3749 | 4.8137 | 3.3612 | 2.6809 | H8 | 1.0941 | 2.1716 | 2.7693 | 4.2369 | 3.1763 | 1.7885 | 1.7587 | | 4.8843 | 4.7500 | 4.5553 | 2.5947 | 2.9020 | H9 | 4.7325 | 3.4896 | 2.1656 | 1.0932 | 3.8782 | 5.5975 | 4.9957 | 4.8843 | | 1.7701 | 1.7713 | 2.5038 | 2.5185 | H10 | 4.2100 | 2.8256 | 2.1676 | 1.0924 | 2.8320 | 4.9012 | 4.3749 | 4.7500 | 1.7701 | | 1.7563 | 3.0820 | 2.5412 | H11 | 4.2557 | 2.8003 | 2.1655 | 1.0917 | 2.7220 | 4.8508 | 4.8137 | 4.5553 | 1.7713 | 1.7563 | | 2.5394 | 3.0845 | H12 | 2.8278 | 2.1226 | 1.0992 | 2.1696 | 3.0506 | 3.7338 | 3.3612 | 2.5947 | 2.5038 | 3.0820 | 2.5394 | | 1.7385 | H13 | 2.7369 | 2.1359 | 1.0983 | 2.1758 | 3.1306 | 3.7755 | 2.6809 | 2.9020 | 2.5185 | 2.5412 | 3.0845 | 1.7385 | |
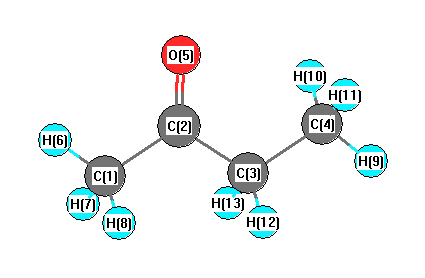 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
116.277 |
|
C1 |
C2 |
O5 |
121.727 |
| C2 |
C1 |
H6 |
109.989 |
|
C2 |
C1 |
H7 |
109.558 |
| C2 |
C1 |
H8 |
111.752 |
|
C2 |
C3 |
C4 |
113.832 |
| C2 |
C3 |
H12 |
107.248 |
|
C2 |
C3 |
H13 |
108.319 |
| C3 |
C2 |
O5 |
121.994 |
|
C3 |
C4 |
H9 |
110.948 |
| C3 |
C4 |
H10 |
111.155 |
|
C3 |
C4 |
H11 |
111.031 |
| C4 |
C3 |
H12 |
110.908 |
|
C4 |
C3 |
H13 |
111.449 |
| H6 |
C1 |
H7 |
108.640 |
|
H6 |
C1 |
H8 |
109.961 |
| H7 |
C1 |
H8 |
106.845 |
|
H9 |
C4 |
H10 |
108.173 |
| H9 |
C4 |
H11 |
108.334 |
|
H10 |
C4 |
H11 |
107.050 |
| H12 |
C3 |
H13 |
104.584 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.732 |
|
|
|
| 2 |
C |
0.329 |
|
|
|
| 3 |
C |
-0.535 |
|
|
|
| 4 |
C |
-0.648 |
|
|
|
| 5 |
O |
-0.305 |
|
|
|
| 6 |
H |
0.251 |
|
|
|
| 7 |
H |
0.245 |
|
|
|
| 8 |
H |
0.234 |
|
|
|
| 9 |
H |
0.219 |
|
|
|
| 10 |
H |
0.231 |
|
|
|
| 11 |
H |
0.235 |
|
|
|
| 12 |
H |
0.240 |
|
|
|
| 13 |
H |
0.234 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.225 |
-2.751 |
0.033 |
2.760 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.179 |
-1.056 |
0.163 |
| y |
-1.056 |
-35.014 |
0.030 |
| z |
0.163 |
0.030 |
-30.501 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.578 |
-1.056 |
0.163 |
| y |
-1.056 |
-4.674 |
0.030 |
| z |
0.163 |
0.030 |
2.095 |
|
| Polar |
|---|
| 3z2-r2 | 4.191 |
|---|
| x2-y2 | 4.835 |
|---|
| xy | -1.056 |
|---|
| xz | 0.163 |
|---|
| yz | 0.030 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.953 |
-0.198 |
0.036 |
| y |
-0.198 |
7.316 |
0.008 |
| z |
0.036 |
0.008 |
5.666 |
<r2> (average value of r
2) Å
2
| <r2> |
136.825 |
| (<r2>)1/2 |
11.697 |
